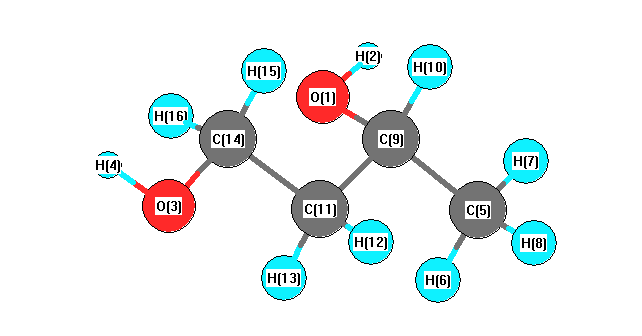
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3646 |
3586 |
6.06 |
|
|
|
| 2 |
A |
3629 |
3570 |
3.87 |
|
|
|
| 3 |
A |
3068 |
3018 |
29.11 |
|
|
|
| 4 |
A |
3047 |
2996 |
33.12 |
|
|
|
| 5 |
A |
3037 |
2987 |
27.07 |
|
|
|
| 6 |
A |
2988 |
2939 |
23.66 |
|
|
|
| 7 |
A |
2975 |
2926 |
21.25 |
|
|
|
| 8 |
A |
2968 |
2920 |
34.47 |
|
|
|
| 9 |
A |
2893 |
2846 |
103.65 |
|
|
|
| 10 |
A |
2879 |
2832 |
33.39 |
|
|
|
| 11 |
A |
1508 |
1483 |
3.77 |
|
|
|
| 12 |
A |
1480 |
1455 |
4.19 |
|
|
|
| 13 |
A |
1475 |
1451 |
4.93 |
|
|
|
| 14 |
A |
1460 |
1436 |
1.74 |
|
|
|
| 15 |
A |
1429 |
1405 |
11.21 |
|
|
|
| 16 |
A |
1403 |
1380 |
19.52 |
|
|
|
| 17 |
A |
1381 |
1359 |
9.85 |
|
|
|
| 18 |
A |
1349 |
1327 |
8.61 |
|
|
|
| 19 |
A |
1298 |
1276 |
36.13 |
|
|
|
| 20 |
A |
1291 |
1270 |
3.28 |
|
|
|
| 21 |
A |
1266 |
1245 |
29.46 |
|
|
|
| 22 |
A |
1227 |
1207 |
44.39 |
|
|
|
| 23 |
A |
1186 |
1167 |
2.85 |
|
|
|
| 24 |
A |
1131 |
1112 |
17.51 |
|
|
|
| 25 |
A |
1106 |
1088 |
51.50 |
|
|
|
| 26 |
A |
1071 |
1053 |
8.66 |
|
|
|
| 27 |
A |
1040 |
1023 |
87.71 |
|
|
|
| 28 |
A |
1014 |
997 |
7.83 |
|
|
|
| 29 |
A |
965 |
949 |
8.78 |
|
|
|
| 30 |
A |
925 |
910 |
9.29 |
|
|
|
| 31 |
A |
842 |
828 |
16.08 |
|
|
|
| 32 |
A |
786 |
773 |
4.58 |
|
|
|
| 33 |
A |
491 |
483 |
17.62 |
|
|
|
| 34 |
A |
467 |
460 |
5.94 |
|
|
|
| 35 |
A |
383 |
376 |
7.10 |
|
|
|
| 36 |
A |
330 |
325 |
12.46 |
|
|
|
| 37 |
A |
304 |
299 |
114.43 |
|
|
|
| 38 |
A |
273 |
268 |
97.73 |
|
|
|
| 39 |
A |
249 |
245 |
0.35 |
|
|
|
| 40 |
A |
174 |
171 |
3.56 |
|
|
|
| 41 |
A |
114 |
113 |
7.18 |
|
|
|
| 42 |
A |
92 |
91 |
5.95 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30320.9 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 29820.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.588 |
|
|
|
| 2 |
H |
0.382 |
|
|
|
| 3 |
O |
-0.589 |
|
|
|
| 4 |
H |
0.385 |
|
|
|
| 5 |
C |
-0.499 |
|
|
|
| 6 |
H |
0.170 |
|
|
|
| 7 |
H |
0.145 |
|
|
|
| 8 |
H |
0.159 |
|
|
|
| 9 |
C |
0.091 |
|
|
|
| 10 |
H |
0.122 |
|
|
|
| 11 |
C |
-0.291 |
|
|
|
| 12 |
H |
0.157 |
|
|
|
| 13 |
H |
0.164 |
|
|
|
| 14 |
C |
-0.083 |
|
|
|
| 15 |
H |
0.121 |
|
|
|
| 16 |
H |
0.154 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.094 |
0.837 |
1.296 |
1.892 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.153 |
-2.631 |
-0.225 |
| y |
-2.631 |
-38.260 |
0.874 |
| z |
-0.225 |
0.874 |
-38.589 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
7.271 |
-2.631 |
-0.225 |
| y |
-2.631 |
-3.388 |
0.874 |
| z |
-0.225 |
0.874 |
-3.883 |
|
| Polar |
|---|
| 3z2-r2 | -7.766 |
|---|
| x2-y2 | 7.107 |
|---|
| xy | -2.631 |
|---|
| xz | -0.225 |
|---|
| yz | 0.874 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.289 |
-0.092 |
0.101 |
| y |
-0.092 |
7.477 |
0.057 |
| z |
0.101 |
0.057 |
6.730 |
<r2> (average value of r
2) Å
2
| <r2> |
225.320 |
| (<r2>)1/2 |
15.011 |