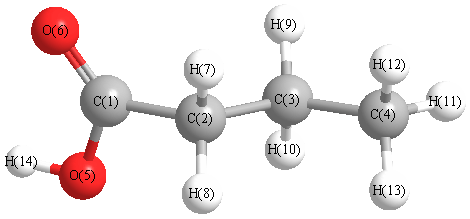Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3721 |
3560 |
50.71 |
|
|
|
| 2 |
A |
3148 |
3011 |
32.08 |
|
|
|
| 3 |
A |
3136 |
3000 |
29.66 |
|
|
|
| 4 |
A |
3131 |
2995 |
26.73 |
|
|
|
| 5 |
A |
3104 |
2970 |
0.45 |
|
|
|
| 6 |
A |
3073 |
2940 |
25.84 |
|
|
|
| 7 |
A |
3060 |
2928 |
7.12 |
|
|
|
| 8 |
A |
3055 |
2922 |
23.67 |
|
|
|
| 9 |
A |
1866 |
1785 |
304.55 |
|
|
|
| 10 |
A |
1533 |
1467 |
6.32 |
|
|
|
| 11 |
A |
1525 |
1459 |
8.51 |
|
|
|
| 12 |
A |
1520 |
1454 |
1.33 |
|
|
|
| 13 |
A |
1498 |
1434 |
6.53 |
|
|
|
| 14 |
A |
1439 |
1376 |
3.29 |
|
|
|
| 15 |
A |
1404 |
1344 |
44.10 |
|
|
|
| 16 |
A |
1392 |
1332 |
32.15 |
|
|
|
| 17 |
A |
1339 |
1281 |
1.52 |
|
|
|
| 18 |
A |
1305 |
1249 |
1.51 |
|
|
|
| 19 |
A |
1273 |
1218 |
22.97 |
|
|
|
| 20 |
A |
1223 |
1170 |
144.91 |
|
|
|
| 21 |
A |
1127 |
1078 |
7.84 |
|
|
|
| 22 |
A |
1095 |
1047 |
74.11 |
|
|
|
| 23 |
A |
1065 |
1019 |
5.55 |
|
|
|
| 24 |
A |
934 |
893 |
1.11 |
|
|
|
| 25 |
A |
900 |
861 |
2.37 |
|
|
|
| 26 |
A |
883 |
844 |
7.73 |
|
|
|
| 27 |
A |
762 |
729 |
17.98 |
|
|
|
| 28 |
A |
734 |
702 |
46.55 |
|
|
|
| 29 |
A |
634 |
607 |
74.73 |
|
|
|
| 30 |
A |
580 |
554 |
45.28 |
|
|
|
| 31 |
A |
430 |
411 |
3.43 |
|
|
|
| 32 |
A |
332 |
318 |
1.06 |
|
|
|
| 33 |
A |
251 |
240 |
0.04 |
|
|
|
| 34 |
A |
188 |
180 |
0.03 |
|
|
|
| 35 |
A |
100 |
95 |
0.23 |
|
|
|
| 36 |
A |
40 |
39 |
0.26 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26397.9 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 25254.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.569 |
|
|
|
| 2 |
C |
-0.388 |
|
|
|
| 3 |
C |
-0.322 |
|
|
|
| 4 |
C |
-0.511 |
|
|
|
| 5 |
O |
-0.573 |
|
|
|
| 6 |
O |
-0.466 |
|
|
|
| 7 |
H |
0.197 |
|
|
|
| 8 |
H |
0.192 |
|
|
|
| 9 |
H |
0.176 |
|
|
|
| 10 |
H |
0.187 |
|
|
|
| 11 |
H |
0.175 |
|
|
|
| 12 |
H |
0.171 |
|
|
|
| 13 |
H |
0.170 |
|
|
|
| 14 |
H |
0.421 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.128 |
1.310 |
0.286 |
1.753 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.419 |
5.421 |
0.271 |
| y |
5.421 |
-38.402 |
-1.457 |
| z |
0.271 |
-1.457 |
-35.623 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.593 |
5.421 |
0.271 |
| y |
5.421 |
-2.881 |
-1.457 |
| z |
0.271 |
-1.457 |
1.287 |
|
| Polar |
|---|
| 3z2-r2 | 2.574 |
|---|
| x2-y2 | 2.983 |
|---|
| xy | 5.421 |
|---|
| xz | 0.271 |
|---|
| yz | -1.457 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.456 |
-0.089 |
-0.318 |
| y |
-0.089 |
6.980 |
0.105 |
| z |
-0.318 |
0.105 |
5.777 |
<r2> (average value of r
2) Å
2
| <r2> |
208.615 |
| (<r2>)1/2 |
14.444 |