return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for NH2CH2CH2COOH (β–alanine)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-323.407459
Energy at 298.15K-323.417327
HF Energy-323.106198
Nuclear repulsion energy246.887772
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3602 3419 5.30      
2 A 3511 3333 1.00      
3 A 3261 3096 663.21      
4 A 3167 3006 7.43      
5 A 3133 2974 21.77      
6 A 3073 2917 8.71      
7 A 3061 2905 62.92      
8 A 1859 1765 329.95      
9 A 1707 1620 32.75      
10 A 1560 1481 5.01      
11 A 1521 1444 288.67      
12 A 1514 1437 5.35      
13 A 1449 1375 11.06      
14 A 1398 1327 6.12      
15 A 1345 1277 2.58      
16 A 1319 1252 14.86      
17 A 1269 1205 76.53      
18 A 1170 1111 6.57      
19 A 1097 1042 5.33      
20 A 1051 997 13.02      
21 A 1005 953 82.17      
22 A 964 915 41.34      
23 A 931 884 25.76      
24 A 885 840 50.54      
25 A 823 781 13.22      
26 A 699 663 10.47      
27 A 578 548 3.50      
28 A 495 470 10.18      
29 A 410 389 14.32      
30 A 335 318 8.04      
31 A 292 277 7.69      
32 A 205 195 4.05      
33 A 89 84 0.37      

Unscaled Zero Point Vibrational Energy (zpe) 24389.2 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 23150.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.23888 0.08335 0.06602

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 1.860 0.576 0.128
C2 1.340 -0.710 -0.367
C3 0.002 -1.010 0.312
C4 -1.073 0.042 0.028
O5 -0.609 1.301 -0.033
O6 -2.242 -0.232 -0.113
H7 2.624 0.912 -0.451
H8 2.218 0.476 1.074
H9 2.026 -1.556 -0.223
H10 1.184 -0.602 -1.445
H11 -0.390 -1.982 0.009
H12 0.152 -1.045 1.399
H13 0.370 1.285 0.076

Atom - Atom Distances (Å)
  N1 C2 C3 C4 O5 O6 H7 H8 H9 H10 H11 H12 H13
N11.47272.44962.98252.57854.18761.01591.01692.16612.07713.40862.67561.6509
C21.47271.53002.55752.82043.62242.07052.06341.09851.09392.17982.15442.2622
C32.44961.53001.52962.41492.41253.33922.77562.16372.15641.09101.09822.3365
C42.98252.55751.52961.34341.20923.82773.48053.49522.77102.13582.13511.9056
O52.57852.82042.41491.34342.24103.28313.14653.89112.97183.29072.85170.9856
O64.18763.62242.41251.20922.24105.00984.66974.47013.69472.55122.94613.0268
H71.01592.07053.33923.82773.28315.00981.63762.54912.31274.20363.65542.3441
H81.01692.06342.77563.48053.14654.66971.63762.41822.92843.73932.58642.2510
H92.16611.09852.16373.49523.89114.47012.54912.41821.76382.46452.53033.3017
H102.07711.09392.15642.77102.97183.69472.31272.92841.76382.54913.05742.5563
H113.40862.17981.09102.13583.29072.55124.20363.73932.46452.54911.76173.3554
H122.67562.15441.09822.13512.85172.94613.65542.58642.53033.05741.76172.6886
H131.65092.26222.33651.90560.98563.02682.34412.25103.30172.55633.35542.6886

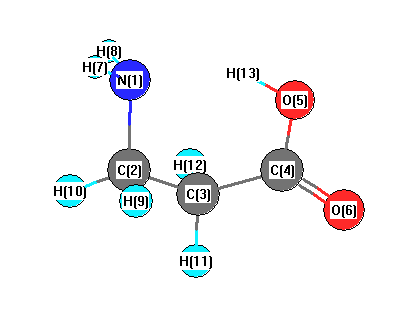
picture of β–alanine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C3 109.319 N1 C2 H9 114.009
N1 C2 H10 107.132 C2 N1 H7 111.285
C2 N1 H8 110.616 C2 C3 C4 113.417
C2 C3 H11 111.438 C2 C3 H12 109.015
C3 C2 H9 109.718 C3 C2 H10 109.415
C3 C4 O5 114.234 C3 C4 O6 123.061
C4 C3 H11 108.001 C4 C3 H12 107.547
C4 O5 H13 108.840 O5 C4 O6 122.699
H7 N1 H8 107.339 H9 C2 H10 107.125
H11 C3 H12 107.169
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 N -0.823      
2 C -0.168      
3 C -0.404      
4 C 0.672      
5 O -0.662      
6 O -0.507      
7 H 0.346      
8 H 0.338      
9 H 0.163      
10 H 0.183      
11 H 0.200      
12 H 0.181      
13 H 0.483      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 7.829 0.496 0.147
y 0.496 6.310 -0.013
z 0.147 -0.013 5.108


<r2> (average value of r2) Å2
<r2> 172.254
(<r2>)1/2 13.125