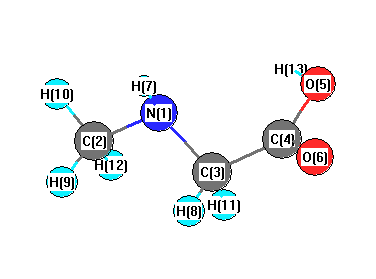
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3527 |
3375 |
3.00 |
|
|
|
| 2 |
A |
3384 |
3237 |
329.51 |
|
|
|
| 3 |
A |
3152 |
3015 |
21.43 |
|
|
|
| 4 |
A |
3111 |
2976 |
3.88 |
|
|
|
| 5 |
A |
3108 |
2974 |
34.00 |
|
|
|
| 6 |
A |
3059 |
2926 |
19.26 |
|
|
|
| 7 |
A |
3017 |
2886 |
72.50 |
|
|
|
| 8 |
A |
1903 |
1820 |
348.41 |
|
|
|
| 9 |
A |
1541 |
1475 |
11.83 |
|
|
|
| 10 |
A |
1522 |
1456 |
30.02 |
|
|
|
| 11 |
A |
1511 |
1445 |
19.66 |
|
|
|
| 12 |
A |
1496 |
1432 |
27.37 |
|
|
|
| 13 |
A |
1478 |
1414 |
38.08 |
|
|
|
| 14 |
A |
1460 |
1397 |
355.51 |
|
|
|
| 15 |
A |
1357 |
1298 |
6.11 |
|
|
|
| 16 |
A |
1300 |
1244 |
4.19 |
|
|
|
| 17 |
A |
1251 |
1197 |
15.89 |
|
|
|
| 18 |
A |
1193 |
1142 |
26.91 |
|
|
|
| 19 |
A |
1167 |
1116 |
30.35 |
|
|
|
| 20 |
A |
1146 |
1097 |
9.78 |
|
|
|
| 21 |
A |
1019 |
974 |
25.31 |
|
|
|
| 22 |
A |
987 |
944 |
22.02 |
|
|
|
| 23 |
A |
930 |
890 |
60.23 |
|
|
|
| 24 |
A |
889 |
851 |
15.94 |
|
|
|
| 25 |
A |
792 |
757 |
76.73 |
|
|
|
| 26 |
A |
650 |
622 |
3.35 |
|
|
|
| 27 |
A |
577 |
552 |
8.70 |
|
|
|
| 28 |
A |
477 |
457 |
9.89 |
|
|
|
| 29 |
A |
383 |
367 |
4.54 |
|
|
|
| 30 |
A |
290 |
278 |
4.90 |
|
|
|
| 31 |
A |
207 |
198 |
1.93 |
|
|
|
| 32 |
A |
141 |
135 |
2.62 |
|
|
|
| 33 |
A |
77 |
73 |
6.42 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24051.1 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 23009.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.626 |
|
|
|
| 2 |
C |
-0.370 |
|
|
|
| 3 |
C |
-0.299 |
|
|
|
| 4 |
C |
0.603 |
|
|
|
| 5 |
O |
-0.584 |
|
|
|
| 6 |
O |
-0.465 |
|
|
|
| 7 |
H |
0.343 |
|
|
|
| 8 |
H |
0.200 |
|
|
|
| 9 |
H |
0.167 |
|
|
|
| 10 |
H |
0.187 |
|
|
|
| 11 |
H |
0.213 |
|
|
|
| 12 |
H |
0.190 |
|
|
|
| 13 |
H |
0.440 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
5.483 |
-0.277 |
-0.623 |
5.525 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.617 |
-0.701 |
-0.546 |
| y |
-0.701 |
-36.920 |
-0.622 |
| z |
-0.546 |
-0.622 |
-32.742 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.787 |
-0.701 |
-0.546 |
| y |
-0.701 |
0.760 |
-0.622 |
| z |
-0.546 |
-0.622 |
7.027 |
|
| Polar |
|---|
| 3z2-r2 | 14.053 |
|---|
| x2-y2 | -5.698 |
|---|
| xy | -0.701 |
|---|
| xz | -0.546 |
|---|
| yz | -0.622 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.119 |
0.460 |
0.023 |
| y |
0.460 |
6.441 |
0.019 |
| z |
0.023 |
0.019 |
5.017 |
<r2> (average value of r
2) Å
2
| <r2> |
198.279 |
| (<r2>)1/2 |
14.081 |