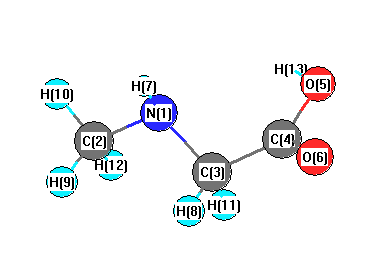
Vibrational Frequencies calculated at PBE1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3550 |
3373 |
3.94 |
|
|
|
| 2 |
A |
3423 |
3253 |
329.47 |
|
|
|
| 3 |
A |
3168 |
3011 |
19.55 |
|
|
|
| 4 |
A |
3126 |
2971 |
8.53 |
|
|
|
| 5 |
A |
3123 |
2968 |
26.79 |
|
|
|
| 6 |
A |
3073 |
2920 |
16.91 |
|
|
|
| 7 |
A |
3032 |
2882 |
67.50 |
|
|
|
| 8 |
A |
1924 |
1828 |
355.40 |
|
|
|
| 9 |
A |
1545 |
1468 |
13.41 |
|
|
|
| 10 |
A |
1525 |
1449 |
37.06 |
|
|
|
| 11 |
A |
1515 |
1440 |
18.46 |
|
|
|
| 12 |
A |
1500 |
1426 |
46.70 |
|
|
|
| 13 |
A |
1483 |
1409 |
114.65 |
|
|
|
| 14 |
A |
1470 |
1397 |
263.50 |
|
|
|
| 15 |
A |
1362 |
1294 |
6.43 |
|
|
|
| 16 |
A |
1306 |
1241 |
4.31 |
|
|
|
| 17 |
A |
1267 |
1204 |
13.34 |
|
|
|
| 18 |
A |
1206 |
1146 |
33.27 |
|
|
|
| 19 |
A |
1176 |
1117 |
21.39 |
|
|
|
| 20 |
A |
1151 |
1094 |
10.18 |
|
|
|
| 21 |
A |
1022 |
972 |
21.92 |
|
|
|
| 22 |
A |
1000 |
950 |
25.80 |
|
|
|
| 23 |
A |
932 |
886 |
60.39 |
|
|
|
| 24 |
A |
902 |
857 |
15.49 |
|
|
|
| 25 |
A |
795 |
756 |
78.81 |
|
|
|
| 26 |
A |
656 |
624 |
3.28 |
|
|
|
| 27 |
A |
581 |
553 |
9.04 |
|
|
|
| 28 |
A |
481 |
457 |
10.01 |
|
|
|
| 29 |
A |
387 |
368 |
4.55 |
|
|
|
| 30 |
A |
292 |
278 |
4.63 |
|
|
|
| 31 |
A |
209 |
199 |
1.96 |
|
|
|
| 32 |
A |
142 |
135 |
2.65 |
|
|
|
| 33 |
A |
78 |
74 |
6.55 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24200.8 cm
-1
Scaled (by 0.9503) Zero Point Vibrational Energy (zpe) 22998.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.636 |
|
|
|
| 2 |
C |
-0.379 |
|
|
|
| 3 |
C |
-0.308 |
|
|
|
| 4 |
C |
0.605 |
|
|
|
| 5 |
O |
-0.589 |
|
|
|
| 6 |
O |
-0.464 |
|
|
|
| 7 |
H |
0.348 |
|
|
|
| 8 |
H |
0.205 |
|
|
|
| 9 |
H |
0.170 |
|
|
|
| 10 |
H |
0.192 |
|
|
|
| 11 |
H |
0.218 |
|
|
|
| 12 |
H |
0.194 |
|
|
|
| 13 |
H |
0.445 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
5.473 |
-0.253 |
-0.609 |
5.513 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.698 |
-0.726 |
-0.576 |
| y |
-0.726 |
-36.987 |
-0.641 |
| z |
-0.576 |
-0.641 |
-32.818 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.796 |
-0.726 |
-0.576 |
| y |
-0.726 |
0.771 |
-0.641 |
| z |
-0.576 |
-0.641 |
7.025 |
|
| Polar |
|---|
| 3z2-r2 | 14.049 |
|---|
| x2-y2 | -5.712 |
|---|
| xy | -0.726 |
|---|
| xz | -0.576 |
|---|
| yz | -0.641 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.038 |
0.463 |
0.028 |
| y |
0.463 |
6.397 |
0.020 |
| z |
0.028 |
0.020 |
4.996 |
<r2> (average value of r
2) Å
2
| <r2> |
191.996 |
| (<r2>)1/2 |
13.856 |