return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5Br (bromobenzene)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-2802.607145
Energy at 298.15K-2802.615105
HF Energy-2802.316419
Nuclear repulsion energy429.990800
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3251 3086 2.31      
2 A1 3239 3074 16.66      
3 A1 3217 3054 0.06      
4 A1 1648 1564 22.23      
5 A1 1532 1454 36.86      
6 A1 1221 1159 0.00      
7 A1 1101 1045 17.36      
8 A1 1050 997 17.82      
9 A1 1016 965 14.94      
10 A1 682 647 19.88      
11 A1 318 302 2.12      
12 A2 956 908 0.00      
13 A2 850 807 0.00      
14 A2 416 395 0.00      
15 B1 965 916 0.07      
16 B1 913 867 2.05      
17 B1 752 714 61.89      
18 B1 674 639 4.97      
19 B1 470 446 5.29      
20 B1 173 164 0.25      
21 B2 3249 3084 7.27      
22 B2 3226 3062 10.78      
23 B2 1651 1568 2.10      
24 B2 1498 1422 7.85      
25 B2 1376 1306 0.72      
26 B2 1353 1284 0.43      
27 B2 1205 1143 0.09      
28 B2 1113 1056 3.21      
29 B2 629 597 0.23      
30 B2 254 241 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 19998.4 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 18982.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.18912 0.03286 0.02800

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
Br1 0.000 0.000 1.809
C2 0.000 0.000 -0.099
C3 0.000 1.214 -0.784
C4 0.000 -1.214 -0.784
C5 0.000 1.207 -2.179
C6 0.000 -1.207 -2.179
C7 0.000 0.000 -2.880
H8 0.000 2.147 -0.234
H9 0.000 -2.147 -0.234
H10 0.000 2.149 -2.716
H11 0.000 -2.149 -2.716
H12 0.000 0.000 -3.964

Atom - Atom Distances (Å)
  Br1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12
Br11.90782.86272.86274.16644.16644.68842.96372.96375.00915.00915.7728
C21.90781.39391.39392.40482.40482.78052.15162.15163.38633.38633.8650
C32.86271.39392.42831.39532.79452.42211.08323.40622.14653.87893.4042
C42.86271.39392.42832.79451.39532.42213.40621.08323.87892.14653.4042
C54.16642.40481.39532.79452.41421.39572.16053.87761.08443.39912.1548
C64.16642.40482.79451.39532.41421.39573.87762.16053.39911.08442.1548
C74.68842.78052.42212.42211.39571.39573.40753.40752.15562.15561.0844
H82.96372.15161.08323.40622.16053.87763.40754.29482.48204.96214.3042
H92.96372.15163.40621.08323.87762.16053.40754.29484.96212.48204.3042
H105.00913.38632.14653.87891.08443.39912.15562.48204.96214.29872.4855
H115.00913.38633.87892.14653.39911.08442.15564.96212.48204.29872.4855
H125.77283.86503.40423.40422.15482.15481.08444.30424.30422.48552.4855

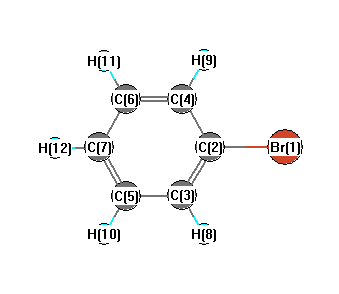
picture of bromobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
Br1 C2 C3 119.419 Br1 C2 C4 119.419
C2 C3 C5 119.128 C2 C3 H8 120.070
C2 C4 C6 119.128 C2 C4 H9 120.070
C3 C2 C4 121.161 C3 C5 C7 120.422
C3 C5 H10 119.379 C4 C6 C7 120.422
C4 C6 H11 119.379 C5 C3 H8 120.802
C5 C7 C6 119.738 C5 C7 H12 120.131
C6 C4 H9 120.802 C6 C7 H12 120.131
C7 C5 H10 120.198 C7 C6 H11 120.198
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 Br -0.130      
2 C 0.045      
3 C -0.164      
4 C -0.164      
5 C -0.159      
6 C -0.159      
7 C -0.156      
8 H 0.187      
9 H 0.187      
10 H 0.172      
11 H 0.172      
12 H 0.169      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 4.563 0.000 0.000
y 0.000 11.507 0.000
z 0.000 0.000 16.015


<r2> (average value of r2) Å2
<r2> 344.634
(<r2>)1/2 18.564