return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5CH3 (toluene)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-271.240764
Energy at 298.15K-271.248739
HF Energy-270.945032
Nuclear repulsion energy268.809921
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3235 3071 18.31      
2 A' 3214 3051 6.98      
3 A' 3199 3037 9.81      
4 A' 3138 2979 20.42      
5 A' 3075 2918 27.55      
6 A' 1677 1592 5.41      
7 A' 1557 1478 12.64      
8 A' 1538 1459 5.48      
9 A' 1459 1385 0.05      
10 A' 1252 1188 0.60      
11 A' 1225 1163 0.18      
12 A' 1083 1028 8.13      
13 A' 1065 1011 1.86      
14 A' 1022 971 0.05      
15 A' 957 908 0.04      
16 A' 898 853 0.40      
17 A' 806 765 0.46      
18 A' 740 703 55.60      
19 A' 673 639 1.94      
20 A' 530 503 0.64      
21 A' 471 447 5.37      
22 A' 211 200 1.81      
23 A" 3222 3059 42.01      
24 A" 3201 3038 7.16      
25 A" 3160 3000 15.82      
26 A" 1655 1571 0.27      
27 A" 1547 1468 10.81      
28 A" 1502 1425 1.36      
29 A" 1383 1313 0.00      
30 A" 1368 1298 0.15      
31 A" 1203 1142 0.06      
32 A" 1130 1073 4.05      
33 A" 1022 970 0.00      
34 A" 945 897 0.00      
35 A" 856 813 0.02      
36 A" 639 606 0.07      
37 A" 413 392 0.00      
38 A" 343 326 0.23      
39 A" 41 39 0.24      

Unscaled Zero Point Vibrational Energy (zpe) 28328.6 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 26889.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.18463 0.08383 0.05828

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.005 0.914 0.000
C2 0.007 0.196 1.204
C3 0.007 -1.201 1.206
C4 0.007 -1.905 0.000
C5 0.007 -1.201 -1.206
C6 0.007 0.196 -1.204
C7 -0.029 2.425 0.000
H8 0.011 0.735 2.148
H9 0.012 -1.738 2.151
H10 0.010 -2.991 0.000
H11 0.012 -1.738 -2.151
H12 0.011 0.735 -2.148
H13 -1.060 2.796 0.000
H14 0.467 2.832 0.885
H15 0.467 2.832 -0.885

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12 H13 H14 H15
C11.40152.43472.81902.43471.40151.51112.15513.41433.90493.41432.15512.16282.16252.1625
C21.40151.39672.42122.78552.40712.53331.08742.15323.40653.87173.39443.05772.69523.3948
C32.43471.39671.39702.41292.78553.82112.15271.08632.15873.39993.87284.30954.07194.5664
C42.81902.42121.39701.39702.42124.32993.40352.15731.08592.15733.40354.82094.84124.8412
C52.43472.78552.41291.39701.39673.82113.87283.39992.15871.08632.15274.30954.56644.0719
C61.40152.40712.78552.42121.39672.53333.39443.87173.40652.15321.08743.05773.39482.6952
C71.51112.53333.82114.32993.82112.53332.73284.68555.41584.68552.73281.09581.09341.0934
H82.15511.08742.15273.40353.87283.39442.73282.47314.30084.95914.29533.16342.48993.7152
H93.41432.15321.08632.15733.39993.87174.68552.47312.48924.30154.95915.13154.76395.5054
H103.90493.40652.15871.08592.15873.40655.41584.30082.48922.48924.30085.88545.90785.9078
H113.41433.87173.39992.15731.08632.15324.68554.95914.30152.48922.47315.13155.50544.7639
H122.15513.39443.87283.40352.15271.08742.73284.29534.95914.30082.47313.16343.71522.4899
H132.16283.05774.30954.82094.30953.05771.09583.16345.13155.88545.13153.16341.76521.7652
H142.16252.69524.07194.84124.56643.39481.09342.48994.76395.90785.50543.71521.76521.7701
H152.16253.39484.56644.84124.07192.69521.09343.71525.50545.90784.76392.48991.76521.7701

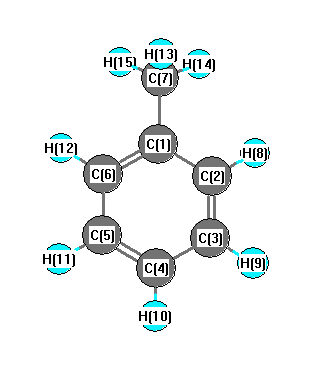
picture of toluene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.943 C1 C2 H8 119.436
C1 C6 C5 120.943 C1 C6 H12 119.436
C1 C7 H13 111.129 C1 C7 H14 111.246
C1 C7 H15 111.246 C2 C1 C6 118.353
C2 C1 C7 120.817 C2 C3 C4 120.153
C2 C3 H9 119.744 C3 C2 H8 119.621
C3 C4 C5 119.455 C3 C4 H10 120.272
C4 C3 H9 120.102 C4 C5 C6 120.153
C4 C5 H11 120.102 C5 C4 H10 120.272
C5 C6 H12 119.621 C6 C1 C7 120.817
C6 C5 H11 119.744 H13 C7 H14 107.480
H13 C7 H15 107.480 H14 C7 H15 108.082
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C 0.118      
2 C -0.198      
3 C -0.156      
4 C -0.166      
5 C -0.156      
6 C -0.198      
7 C -0.523      
8 H 0.152      
9 H 0.158      
10 H 0.157      
11 H 0.158      
12 H 0.152      
13 H 0.173      
14 H 0.164      
15 H 0.164      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 4.525 -0.092 0.000
y -0.092 13.237 0.000
z 0.000 0.000 11.567


<r2> (average value of r2) Å2
<r2> 201.248
(<r2>)1/2 14.186