return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5Cl (chlorobenzene)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-690.534399
Energy at 298.15K-690.539561
HF Energy-689.601785
Nuclear repulsion energy314.777411
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3233 3078 2.48      
2 A1 3221 3067 16.61      
3 A1 3199 3047 0.04      
4 A1 1681 1601 20.44      
5 A1 1548 1475 41.49      
6 A1 1228 1169 0.02      
7 A1 1142 1088 33.51      
8 A1 1064 1013 11.05      
9 A1 1022 973 7.21      
10 A1 725 690 19.22      
11 A1 424 404 2.51      
12 A2 925 881 0.00      
13 A2 839 799 0.00      
14 A2 408 388 0.00      
15 B1 919 875 0.16      
16 B1 879 837 1.16      
17 B1 741 706 80.22      
18 B1 573 545 0.26      
19 B1 467 445 5.56      
20 B1 190 181 0.07      
21 B2 3230 3076 10.64      
22 B2 3209 3056 9.00      
23 B2 1677 1597 2.36      
24 B2 1509 1437 7.13      
25 B2 1360 1295 0.28      
26 B2 1303 1240 0.11      
27 B2 1202 1145 0.01      
28 B2 1122 1069 3.07      
29 B2 631 601 0.20      
30 B2 301 287 0.17      

Unscaled Zero Point Vibrational Energy (zpe) 19984.6 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 19031.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.18902 0.05212 0.04085

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
Cl1 0.000 0.000 2.258
C2 0.000 0.000 0.509
C3 0.000 1.216 -0.175
C4 0.000 -1.216 -0.175
C5 0.000 1.208 -1.572
C6 0.000 -1.208 -1.572
C7 0.000 0.000 -2.273
H8 0.000 2.151 0.379
H9 0.000 -2.151 0.379
H10 0.000 2.154 -2.111
H11 0.000 -2.154 -2.111
H12 0.000 0.000 -3.362

Atom - Atom Distances (Å)
  Cl1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12
Cl11.74892.72012.72014.01644.01644.53152.85632.85634.87164.87165.6200
C21.74891.39501.39502.40672.40672.78262.15512.15513.39233.39233.8711
C32.72011.39502.43111.39692.79762.42481.08763.41212.15153.88633.4105
C42.72011.39502.43112.79761.39692.42483.41211.08763.88632.15153.4105
C54.01642.40671.39692.79762.41671.39712.16713.88511.08883.40552.1594
C64.01642.40672.79761.39692.41671.39713.88512.16713.40551.08882.1594
C74.53152.78262.42482.42481.39711.39713.41523.41522.16042.16041.0885
H82.85632.15511.08763.41212.16713.88513.41524.30242.49044.97384.3154
H92.85632.15513.41211.08763.88512.16713.41524.30244.97382.49044.3154
H104.87163.39232.15153.88631.08883.40552.16042.49044.97384.30852.4910
H114.87163.39233.88632.15153.40551.08882.16044.97382.49044.30852.4910
H125.62003.87113.41053.41052.15942.15941.08854.31544.31542.49102.4910

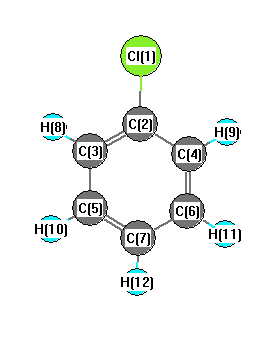
picture of chlorobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
Cl1 C2 C3 119.385 Cl1 C2 C4 119.385
C2 C3 C5 119.088 C2 C3 H8 119.967
C2 C4 C6 119.088 C2 C4 H9 119.967
C3 C2 C4 121.231 C3 C5 C7 120.424
C3 C5 H10 119.383 C4 C6 C7 120.424
C4 C6 H11 119.383 C5 C3 H8 120.945
C5 C7 C6 119.746 C5 C7 H12 120.127
C6 C4 H9 120.945 C6 C7 H12 120.127
C7 C5 H10 120.194 C7 C6 H11 120.194
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability