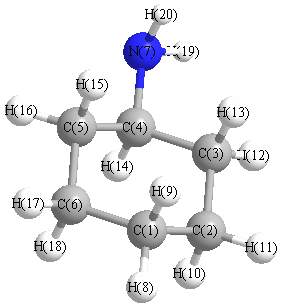
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3557 |
3403 |
0.89 |
|
|
|
| 2 |
A |
3467 |
3317 |
2.86 |
|
|
|
| 3 |
A |
3104 |
2970 |
42.29 |
|
|
|
| 4 |
A |
3093 |
2959 |
72.38 |
|
|
|
| 5 |
A |
3087 |
2953 |
42.34 |
|
|
|
| 6 |
A |
3085 |
2952 |
54.49 |
|
|
|
| 7 |
A |
3070 |
2937 |
64.08 |
|
|
|
| 8 |
A |
3039 |
2907 |
59.42 |
|
|
|
| 9 |
A |
3035 |
2903 |
13.14 |
|
|
|
| 10 |
A |
3032 |
2901 |
20.18 |
|
|
|
| 11 |
A |
3029 |
2897 |
38.52 |
|
|
|
| 12 |
A |
3015 |
2885 |
18.40 |
|
|
|
| 13 |
A |
3014 |
2884 |
6.23 |
|
|
|
| 14 |
A |
1695 |
1622 |
28.17 |
|
|
|
| 15 |
A |
1530 |
1464 |
1.86 |
|
|
|
| 16 |
A |
1515 |
1449 |
12.93 |
|
|
|
| 17 |
A |
1511 |
1445 |
4.04 |
|
|
|
| 18 |
A |
1508 |
1443 |
0.83 |
|
|
|
| 19 |
A |
1501 |
1436 |
0.12 |
|
|
|
| 20 |
A |
1440 |
1377 |
4.49 |
|
|
|
| 21 |
A |
1401 |
1340 |
2.87 |
|
|
|
| 22 |
A |
1397 |
1337 |
0.98 |
|
|
|
| 23 |
A |
1392 |
1332 |
3.58 |
|
|
|
| 24 |
A |
1383 |
1323 |
0.71 |
|
|
|
| 25 |
A |
1376 |
1316 |
3.15 |
|
|
|
| 26 |
A |
1343 |
1285 |
1.09 |
|
|
|
| 27 |
A |
1314 |
1257 |
1.39 |
|
|
|
| 28 |
A |
1298 |
1242 |
3.39 |
|
|
|
| 29 |
A |
1294 |
1238 |
1.59 |
|
|
|
| 30 |
A |
1237 |
1184 |
5.58 |
|
|
|
| 31 |
A |
1211 |
1158 |
1.95 |
|
|
|
| 32 |
A |
1147 |
1097 |
7.84 |
|
|
|
| 33 |
A |
1131 |
1082 |
7.09 |
|
|
|
| 34 |
A |
1104 |
1056 |
0.68 |
|
|
|
| 35 |
A |
1082 |
1035 |
0.31 |
|
|
|
| 36 |
A |
1059 |
1013 |
7.38 |
|
|
|
| 37 |
A |
1049 |
1003 |
0.63 |
|
|
|
| 38 |
A |
995 |
952 |
1.72 |
|
|
|
| 39 |
A |
947 |
906 |
46.20 |
|
|
|
| 40 |
A |
920 |
881 |
69.02 |
|
|
|
| 41 |
A |
898 |
859 |
27.87 |
|
|
|
| 42 |
A |
882 |
843 |
48.95 |
|
|
|
| 43 |
A |
862 |
825 |
2.74 |
|
|
|
| 44 |
A |
797 |
762 |
0.09 |
|
|
|
| 45 |
A |
794 |
759 |
2.68 |
|
|
|
| 46 |
A |
551 |
528 |
1.43 |
|
|
|
| 47 |
A |
459 |
439 |
1.29 |
|
|
|
| 48 |
A |
449 |
430 |
2.10 |
|
|
|
| 49 |
A |
405 |
387 |
0.12 |
|
|
|
| 50 |
A |
344 |
329 |
15.72 |
|
|
|
| 51 |
A |
332 |
317 |
1.58 |
|
|
|
| 52 |
A |
267 |
255 |
36.29 |
|
|
|
| 53 |
A |
228 |
218 |
7.31 |
|
|
|
| 54 |
A |
159 |
152 |
0.32 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 41415.4 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 39622.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.311 |
|
|
|
| 2 |
C |
-0.317 |
|
|
|
| 3 |
C |
-0.301 |
|
|
|
| 4 |
C |
-0.017 |
|
|
|
| 5 |
C |
-0.316 |
|
|
|
| 6 |
C |
-0.315 |
|
|
|
| 7 |
N |
-0.737 |
|
|
|
| 8 |
H |
0.159 |
|
|
|
| 9 |
H |
0.153 |
|
|
|
| 10 |
H |
0.155 |
|
|
|
| 11 |
H |
0.160 |
|
|
|
| 12 |
H |
0.175 |
|
|
|
| 13 |
H |
0.142 |
|
|
|
| 14 |
H |
0.153 |
|
|
|
| 15 |
H |
0.143 |
|
|
|
| 16 |
H |
0.148 |
|
|
|
| 17 |
H |
0.159 |
|
|
|
| 18 |
H |
0.155 |
|
|
|
| 19 |
H |
0.308 |
|
|
|
| 20 |
H |
0.303 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.545 |
1.152 |
0.517 |
1.375 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.206 |
-4.124 |
-1.731 |
| y |
-4.124 |
-45.551 |
-0.629 |
| z |
-1.731 |
-0.629 |
-43.689 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.586 |
-4.124 |
-1.731 |
| y |
-4.124 |
-0.103 |
-0.629 |
| z |
-1.731 |
-0.629 |
2.689 |
|
| Polar |
|---|
| 3z2-r2 | 5.379 |
|---|
| x2-y2 | -1.655 |
|---|
| xy | -4.124 |
|---|
| xz | -1.731 |
|---|
| yz | -0.629 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.090 |
-0.197 |
-0.116 |
| y |
-0.197 |
10.070 |
-0.127 |
| z |
-0.116 |
-0.127 |
9.196 |
<r2> (average value of r
2) Å
2
| <r2> |
232.476 |
| (<r2>)1/2 |
15.247 |