return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H13N (cyclohexanamine)

using model chemistry: MP3/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP3/6-31G*
 hartrees
Energy at 0K-290.239154
Energy at 298.15K-290.255735
HF Energy-289.228595
Nuclear repulsion energy329.985460
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3587 3367 0.52      
2 A 3499 3285 1.08      
3 A 3150 2957 34.19      
4 A 3139 2946 59.81      
5 A 3133 2940 33.41      
6 A 3132 2939 46.10      
7 A 3117 2926 53.20      
8 A 3084 2895 55.64      
9 A 3081 2892 11.58      
10 A 3080 2891 16.82      
11 A 3077 2888 25.83      
12 A 3063 2875 10.49      
13 A 3063 2875 11.65      
14 A 1730 1624 32.39      
15 A 1571 1475 1.64      
16 A 1558 1462 13.47      
17 A 1553 1458 4.45      
18 A 1550 1455 0.92      
19 A 1544 1450 0.20      
20 A 1480 1389 4.46      
21 A 1440 1351 5.12      
22 A 1437 1348 0.97      
23 A 1432 1344 1.85      
24 A 1416 1329 0.41      
25 A 1408 1321 2.54      
26 A 1377 1293 0.94      
27 A 1343 1261 1.59      
28 A 1326 1245 3.09      
29 A 1324 1242 0.91      
30 A 1268 1191 5.39      
31 A 1245 1168 2.91      
32 A 1179 1106 5.56      
33 A 1169 1097 8.66      
34 A 1130 1061 1.16      
35 A 1107 1039 0.24      
36 A 1087 1021 6.85      
37 A 1074 1008 0.64      
38 A 1019 957 2.51      
39 A 982 922 97.12      
40 A 952 894 51.80      
41 A 923 866 14.00      
42 A 910 854 20.25      
43 A 881 827 2.64      
44 A 817 767 0.12      
45 A 814 764 2.70      
46 A 567 532 1.26      
47 A 473 444 1.44      
48 A 461 433 2.03      
49 A 416 391 0.12      
50 A 353 331 16.40      
51 A 342 321 1.53      
52 A 279 262 40.94      
53 A 233 219 4.85      
54 A 165 155 0.43      

Unscaled Zero Point Vibrational Energy (zpe) 42269.8 cm-1
Scaled (by 0.9386) Zero Point Vibrational Energy (zpe) 39674.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP3/6-31G*
ABC
0.14198 0.07353 0.05340

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP3/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.878 0.014 0.291
C2 1.182 -1.252 -0.218
C3 -0.303 -1.267 0.160
C4 -1.029 -0.013 -0.331
C5 -0.323 1.251 0.177
C6 1.162 1.274 -0.205
N7 -2.449 -0.105 0.033
H8 2.928 0.024 -0.025
H9 1.879 0.009 1.390
H10 1.275 -1.299 -1.312
H11 1.679 -2.146 0.177
H12 -0.807 -2.151 -0.243
H13 -0.399 -1.318 1.256
H14 -0.988 -0.009 -1.430
H15 -0.419 1.284 1.273
H16 -0.834 2.142 -0.211
H17 1.644 2.173 0.196
H18 1.251 1.332 -1.299
H19 -2.938 0.723 -0.305
H20 -2.526 -0.075 1.050

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 N7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18 H19 H20
C11.53242.53312.97372.52741.53214.33631.09651.09912.15832.17233.49072.80923.34342.80243.48412.17362.15864.90424.4704
C21.53241.53232.53722.94762.52643.81562.17202.15941.09901.09642.18252.16212.77903.34913.94803.48112.80234.56944.0918
C32.53311.53231.52972.51822.95532.44293.48482.81102.15842.16881.09501.10102.14022.78503.46983.95253.36113.33402.6746
C42.97372.53721.52971.53542.54421.46823.96973.37952.81533.48482.15122.14901.10012.15172.16753.49332.81882.04562.0379
C52.52742.94762.51821.53541.53292.52603.48072.80353.35813.94333.46252.78752.14861.10071.09822.17162.15952.71112.7161
C61.53212.52642.95532.54421.53293.87202.17162.15862.80333.48003.95063.35982.78672.16442.17651.09641.09914.13724.1227
N74.33633.81562.44291.46822.52603.87205.37884.53654.13534.60702.63772.67692.07042.75442.77774.68654.18651.01961.0204
H81.09652.17203.48483.96973.48072.17165.37881.76182.47832.51224.32823.80964.16083.80464.32172.51322.47955.91415.5604
H91.09912.15942.81103.37952.80352.15864.53651.76183.06232.48123.81442.63974.02152.62993.80442.48243.06225.15564.4188
H102.15831.09902.15842.81533.35812.80334.13532.47833.06231.75992.49033.06522.60754.02804.18393.80362.63134.78024.6398
H112.17231.09642.16883.48483.94333.48004.60702.51222.48121.75992.52152.48363.77674.16724.98564.31923.80255.45714.7683
H123.49072.18251.09502.15123.46253.95062.63774.32823.81442.49032.52151.76312.45573.77484.29344.98974.18093.57832.9899
H132.80922.16211.10102.14902.78753.35982.67693.80962.63973.06522.48361.76313.04572.60173.78324.18114.03393.61192.4721
H143.34342.77902.14021.10012.14862.78672.07044.16084.02152.60753.77672.45573.04573.05062.47753.78562.61262.36762.9199
H152.80243.34912.78502.15171.10072.16442.75443.80462.62994.02804.16723.77482.60173.05061.76452.49103.06753.02492.5174
H163.48413.94803.46982.16751.09822.17652.77774.32173.80444.18394.98564.29343.78322.47751.76452.51142.48782.53903.0609
H172.17363.48113.95253.49332.17161.09644.68652.51322.48243.80364.31924.98974.18113.78562.49102.51141.76044.83134.8136
H182.15862.80233.36112.81882.15951.09914.18652.47953.06222.63133.80254.18094.03392.61263.06752.48781.76044.34774.6654
H194.90424.56943.33402.04562.71114.13721.01965.91415.15564.78025.45713.57833.61192.36763.02492.53904.83134.34771.6258
H204.47044.09182.67462.03792.71614.12271.02045.56044.41884.63984.76832.98992.47212.91992.51743.06094.81364.66541.6258

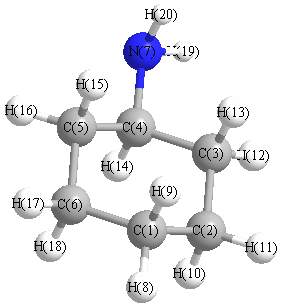
picture of cyclohexanamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 111.489 C1 C2 H10 109.105
C1 C2 H11 110.352 C1 C6 C5 111.096
C1 C6 H17 110.474 C1 C6 H18 109.139
C2 C1 C6 111.058 C2 C1 H8 110.323
C2 C1 H9 109.182 C2 C3 C4 111.911
C2 C3 H12 111.249 C2 C3 H13 109.284
C3 C2 H10 109.110 C3 C2 H11 110.077
C3 C4 C5 110.483 C3 C4 N7 109.137
C3 C4 H14 107.830 C4 C3 H12 108.965
C4 C3 H13 108.454 C4 C5 C6 112.024
C4 C5 H15 108.288 C4 C5 H16 109.655
C4 N7 H19 109.294 C4 N7 H20 108.611
C5 C4 N7 114.472 C5 C4 H14 108.084
C5 C6 H17 110.258 C5 C6 H18 109.149
C6 C1 H8 110.304 C6 C1 H9 109.139
C6 C5 H15 109.443 C6 C5 H16 110.539
N7 C4 H14 106.562 H8 C1 H9 106.723
H10 C2 H11 106.571 H12 C3 H13 106.808
H15 C5 H16 106.724 H17 C6 H18 106.606
H19 N7 H20 105.683
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability