return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H10O (cyclohexanone)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-309.530960
Energy at 298.15K 
HF Energy-309.216064
Nuclear repulsion energy309.169152
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3152 2992 16.98      
2 A' 3123 2964 48.64      
3 A' 3114 2956 43.56      
4 A' 3072 2916 39.61      
5 A' 3061 2906 11.00      
6 A' 3059 2904 17.70      
7 A' 1799 1708 156.53      
8 A' 1554 1475 0.82      
9 A' 1539 1461 8.63      
10 A' 1519 1441 7.26      
11 A' 1412 1340 6.41      
12 A' 1378 1308 2.13      
13 A' 1299 1233 0.14      
14 A' 1274 1209 0.62      
15 A' 1160 1101 0.65      
16 A' 1047 994 2.38      
17 A' 1020 968 0.82      
18 A' 891 846 3.22      
19 A' 859 815 0.13      
20 A' 764 726 2.76      
21 A' 664 630 1.44      
22 A' 484 460 0.31      
23 A' 407 387 1.08      
24 A' 309 294 0.20      
25 A' 94 89 2.65      
26 A" 3151 2991 18.53      
27 A" 3118 2960 34.94      
28 A" 3071 2915 19.10      
29 A" 3055 2900 2.86      
30 A" 1539 1460 4.25      
31 A" 1511 1434 6.08      
32 A" 1416 1344 0.62      
33 A" 1398 1327 8.78      
34 A" 1372 1303 12.18      
35 A" 1316 1249 0.07      
36 A" 1264 1200 28.20      
37 A" 1162 1103 19.38      
38 A" 1103 1047 0.78      
39 A" 1084 1029 6.35      
40 A" 938 890 3.63      
41 A" 918 871 2.85      
42 A" 780 741 4.00      
43 A" 493 468 10.34      
44 A" 417 396 2.31      
45 A" 184 175 0.84      

Unscaled Zero Point Vibrational Energy (zpe) 33672.2 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 31961.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.418 -1.077 0.000
C2 -0.165 -0.506 1.283
C3 -0.165 -0.506 -1.283
C4 -0.165 1.034 1.264
C5 -0.165 1.034 -1.264
C6 -0.838 1.579 0.000
O7 1.296 -1.926 0.000
H8 0.398 -0.907 2.128
H9 -1.201 -0.861 1.368
H10 0.398 -0.907 -2.128
H11 -1.201 -0.861 -1.368
H12 0.871 1.393 1.304
H13 -0.663 1.414 2.162
H14 0.871 1.393 -1.304
H15 -0.663 1.414 -2.162
H16 -0.807 2.674 0.000
H17 -1.899 1.295 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 O7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17
C11.52031.52032.52842.52842.93811.22102.13502.13082.13502.13082.82943.47022.82943.47023.94553.3159
C21.52032.56541.54002.97562.53872.40731.09241.09893.48032.86842.16292.16883.37203.97423.48802.8099
C31.52032.56542.97561.54002.53872.40733.48032.86841.09241.09893.37203.97422.16292.16883.48802.8099
C42.52841.54002.97562.52711.53213.53432.19872.16293.94853.40491.09701.09482.79213.48192.16722.1617
C52.52842.97561.54002.52711.53213.53433.94853.40492.19872.16292.79213.48191.09701.09482.16722.1617
C62.93812.53872.53871.53211.53214.10353.49872.82133.49872.82132.15852.17502.15852.17501.09521.0981
O71.22102.40732.40733.53433.53434.10352.52433.03942.52433.03943.59104.43403.59104.43405.05744.5365
H82.13501.09243.48032.19873.94853.49872.52431.77134.25643.84512.48852.55214.15894.99144.33653.8285
H92.13081.09892.86842.16293.40492.82133.03941.77133.84512.73653.06262.46854.06434.23373.81122.6475
H102.13503.48031.09243.94852.19873.49872.52434.25643.84511.77134.15894.99142.48852.55214.33653.8285
H112.13082.86841.09893.40492.16292.82133.03943.84512.73651.77134.06434.23373.06262.46853.81122.6475
H122.82942.16293.37201.09702.79212.15853.59102.48853.06264.15894.06431.75772.60873.79042.48153.0637
H133.47022.16883.97421.09483.48192.17504.43402.55212.46854.99144.23371.75773.79044.32322.50632.4929
H142.82943.37202.16292.79211.09702.15853.59104.15894.06432.48853.06262.60873.79041.75772.48153.0637
H153.47023.97422.16883.48191.09482.17504.43404.99144.23372.55212.46853.79044.32321.75772.50632.4929
H163.94553.48803.48802.16722.16721.09525.05744.33653.81124.33653.81122.48152.50632.48152.50631.7593
H173.31592.80992.80992.16172.16171.09814.53653.82852.64753.82852.64753.06372.49293.06372.49291.7593

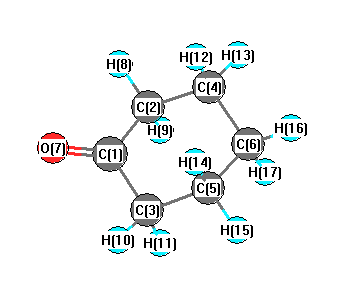
picture of cyclohexanone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 111.414 C1 C2 H8 108.497
C1 C2 H9 107.800 C1 C3 C5 111.414
C1 C3 H10 108.497 C1 C3 H11 107.800
C2 C1 C3 115.065 C2 C1 O7 122.467
C2 C4 C6 111.458 C2 C4 H12 109.052
C2 C4 H13 109.639 C3 C1 O7 122.467
C3 C5 C6 111.458 C3 C5 H14 109.052
C3 C5 H15 109.639 C4 C2 H8 112.163
C4 C2 H9 108.949 C4 C6 C5 111.119
C4 C6 H16 110.041 C4 C6 H17 109.436
C5 C3 H10 112.163 C5 C3 H11 108.949
C5 C6 H16 110.041 C5 C6 H17 109.436
C6 C4 H12 109.253 C6 C4 H13 110.681
C6 C5 H14 109.253 C6 C5 H15 110.681
H8 C2 H9 107.864 H10 C3 H11 107.864
H12 C4 H13 106.625 H14 C5 H15 106.625
H16 C6 H17 106.662
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C 0.488      
2 C -0.370      
3 C -0.370      
4 C -0.298      
5 C -0.298      
6 C -0.288      
7 O -0.491      
8 H 0.183      
9 H 0.165      
10 H 0.183      
11 H 0.165      
12 H 0.158      
13 H 0.159      
14 H 0.158      
15 H 0.159      
16 H 0.156      
17 H 0.144      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 8.467 -1.087 0.000
y -1.087 9.402 0.000
z 0.000 0.000 9.240


<r2> (average value of r2) Å2
<r2> 209.160
(<r2>)1/2 14.462