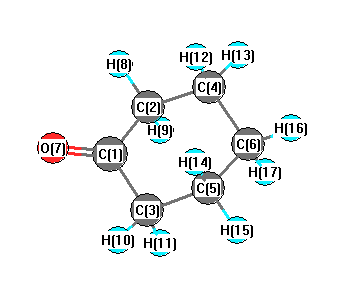
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3055 |
3005 |
15.62 |
|
|
|
| 2 |
A' |
3025 |
2975 |
47.54 |
|
|
|
| 3 |
A' |
3018 |
2968 |
41.15 |
|
|
|
| 4 |
A' |
2974 |
2925 |
39.76 |
|
|
|
| 5 |
A' |
2959 |
2911 |
17.81 |
|
|
|
| 6 |
A' |
2958 |
2909 |
12.36 |
|
|
|
| 7 |
A' |
1760 |
1731 |
169.85 |
|
|
|
| 8 |
A' |
1483 |
1459 |
1.34 |
|
|
|
| 9 |
A' |
1467 |
1443 |
9.78 |
|
|
|
| 10 |
A' |
1445 |
1421 |
7.73 |
|
|
|
| 11 |
A' |
1347 |
1324 |
7.29 |
|
|
|
| 12 |
A' |
1312 |
1290 |
3.42 |
|
|
|
| 13 |
A' |
1243 |
1222 |
0.03 |
|
|
|
| 14 |
A' |
1221 |
1201 |
0.54 |
|
|
|
| 15 |
A' |
1105 |
1087 |
0.51 |
|
|
|
| 16 |
A' |
1009 |
993 |
3.12 |
|
|
|
| 17 |
A' |
981 |
965 |
0.78 |
|
|
|
| 18 |
A' |
854 |
840 |
4.71 |
|
|
|
| 19 |
A' |
835 |
821 |
0.17 |
|
|
|
| 20 |
A' |
737 |
725 |
3.88 |
|
|
|
| 21 |
A' |
640 |
629 |
1.46 |
|
|
|
| 22 |
A' |
465 |
457 |
0.20 |
|
|
|
| 23 |
A' |
395 |
388 |
1.08 |
|
|
|
| 24 |
A' |
301 |
296 |
0.15 |
|
|
|
| 25 |
A' |
90 |
88 |
1.94 |
|
|
|
| 26 |
A" |
3054 |
3004 |
20.67 |
|
|
|
| 27 |
A" |
3020 |
2970 |
33.33 |
|
|
|
| 28 |
A" |
2972 |
2923 |
22.04 |
|
|
|
| 29 |
A" |
2952 |
2903 |
1.93 |
|
|
|
| 30 |
A" |
1468 |
1444 |
4.84 |
|
|
|
| 31 |
A" |
1437 |
1413 |
6.67 |
|
|
|
| 32 |
A" |
1350 |
1328 |
0.57 |
|
|
|
| 33 |
A" |
1333 |
1311 |
8.94 |
|
|
|
| 34 |
A" |
1303 |
1281 |
14.40 |
|
|
|
| 35 |
A" |
1261 |
1240 |
0.15 |
|
|
|
| 36 |
A" |
1207 |
1187 |
28.58 |
|
|
|
| 37 |
A" |
1108 |
1090 |
22.50 |
|
|
|
| 38 |
A" |
1067 |
1049 |
2.01 |
|
|
|
| 39 |
A" |
1038 |
1021 |
7.12 |
|
|
|
| 40 |
A" |
902 |
887 |
6.36 |
|
|
|
| 41 |
A" |
882 |
868 |
0.41 |
|
|
|
| 42 |
A" |
741 |
729 |
5.39 |
|
|
|
| 43 |
A" |
475 |
468 |
7.79 |
|
|
|
| 44 |
A" |
401 |
394 |
2.56 |
|
|
|
| 45 |
A" |
178 |
175 |
0.85 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32412.7 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 31877.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.431 |
|
|
|
| 2 |
C |
-0.373 |
|
|
|
| 3 |
C |
-0.373 |
|
|
|
| 4 |
C |
-0.299 |
|
|
|
| 5 |
C |
-0.299 |
|
|
|
| 6 |
C |
-0.292 |
|
|
|
| 7 |
O |
-0.415 |
|
|
|
| 8 |
H |
0.175 |
|
|
|
| 9 |
H |
0.167 |
|
|
|
| 10 |
H |
0.175 |
|
|
|
| 11 |
H |
0.167 |
|
|
|
| 12 |
H |
0.160 |
|
|
|
| 13 |
H |
0.157 |
|
|
|
| 14 |
H |
0.160 |
|
|
|
| 15 |
H |
0.157 |
|
|
|
| 16 |
H |
0.156 |
|
|
|
| 17 |
H |
0.147 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.958 |
-2.246 |
0.000 |
2.980 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.684 |
4.458 |
0.001 |
| y |
4.458 |
-47.597 |
-0.003 |
| z |
0.001 |
-0.003 |
-41.204 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.284 |
4.458 |
0.001 |
| y |
4.458 |
-4.653 |
-0.003 |
| z |
0.001 |
-0.003 |
4.937 |
|
| Polar |
|---|
| 3z2-r2 | 9.873 |
|---|
| x2-y2 | 2.913 |
|---|
| xy | 4.458 |
|---|
| xz | 0.001 |
|---|
| yz | -0.003 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.726 |
-1.164 |
-0.000 |
| y |
-1.164 |
9.911 |
0.001 |
| z |
-0.000 |
0.001 |
9.785 |
<r2> (average value of r
2) Å
2
| <r2> |
210.942 |
| (<r2>)1/2 |
14.524 |