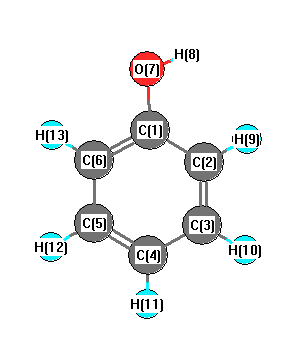
Vibrational Frequencies calculated at PBE1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3818 |
3628 |
49.09 |
|
|
|
| 2 |
A' |
3245 |
3084 |
4.83 |
|
|
|
| 3 |
A' |
3238 |
3077 |
18.21 |
|
|
|
| 4 |
A' |
3223 |
3063 |
18.42 |
|
|
|
| 5 |
A' |
3215 |
3055 |
0.15 |
|
|
|
| 6 |
A' |
3192 |
3033 |
15.23 |
|
|
|
| 7 |
A' |
1698 |
1614 |
52.19 |
|
|
|
| 8 |
A' |
1684 |
1600 |
36.77 |
|
|
|
| 9 |
A' |
1564 |
1486 |
60.96 |
|
|
|
| 10 |
A' |
1530 |
1454 |
32.31 |
|
|
|
| 11 |
A' |
1406 |
1336 |
27.44 |
|
|
|
| 12 |
A' |
1377 |
1308 |
17.59 |
|
|
|
| 13 |
A' |
1332 |
1266 |
78.01 |
|
|
|
| 14 |
A' |
1221 |
1161 |
139.68 |
|
|
|
| 15 |
A' |
1205 |
1145 |
5.33 |
|
|
|
| 16 |
A' |
1189 |
1130 |
5.15 |
|
|
|
| 17 |
A' |
1112 |
1057 |
11.85 |
|
|
|
| 18 |
A' |
1064 |
1011 |
4.69 |
|
|
|
| 19 |
A' |
1019 |
968 |
0.61 |
|
|
|
| 20 |
A' |
843 |
801 |
16.76 |
|
|
|
| 21 |
A' |
631 |
600 |
0.37 |
|
|
|
| 22 |
A' |
536 |
510 |
0.95 |
|
|
|
| 23 |
A' |
408 |
388 |
10.63 |
|
|
|
| 24 |
A" |
988 |
939 |
0.19 |
|
|
|
| 25 |
A" |
961 |
913 |
0.10 |
|
|
|
| 26 |
A" |
891 |
846 |
7.27 |
|
|
|
| 27 |
A" |
828 |
787 |
0.01 |
|
|
|
| 28 |
A" |
767 |
729 |
64.14 |
|
|
|
| 29 |
A" |
704 |
669 |
15.09 |
|
|
|
| 30 |
A" |
520 |
494 |
8.00 |
|
|
|
| 31 |
A" |
422 |
401 |
0.19 |
|
|
|
| 32 |
A" |
370 |
351 |
126.35 |
|
|
|
| 33 |
A" |
234 |
223 |
0.86 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23215.7 cm
-1
Scaled (by 0.9503) Zero Point Vibrational Energy (zpe) 22061.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.345 |
|
|
|
| 2 |
C |
-0.237 |
|
|
|
| 3 |
C |
-0.173 |
|
|
|
| 4 |
C |
-0.177 |
|
|
|
| 5 |
C |
-0.174 |
|
|
|
| 6 |
C |
-0.198 |
|
|
|
| 7 |
O |
-0.654 |
|
|
|
| 8 |
H |
0.423 |
|
|
|
| 9 |
H |
0.158 |
|
|
|
| 10 |
H |
0.170 |
|
|
|
| 11 |
H |
0.166 |
|
|
|
| 12 |
H |
0.171 |
|
|
|
| 13 |
H |
0.180 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.373 |
0.061 |
0.000 |
1.374 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.543 |
-4.409 |
0.000 |
| y |
-4.409 |
-36.335 |
0.000 |
| z |
0.000 |
0.000 |
-44.013 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.631 |
-4.409 |
0.000 |
| y |
-4.409 |
2.942 |
0.000 |
| z |
0.000 |
0.000 |
-8.574 |
|
| Polar |
|---|
| 3z2-r2 | -17.147 |
|---|
| x2-y2 | 1.792 |
|---|
| xy | -4.409 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.809 |
-0.246 |
0.000 |
| y |
-0.246 |
11.934 |
0.000 |
| z |
0.000 |
0.000 |
3.420 |
<r2> (average value of r
2) Å
2
| <r2> |
185.190 |
| (<r2>)1/2 |
13.608 |