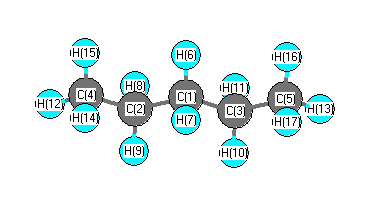
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3145 |
2991 |
45.28 |
|
|
|
| 2 |
A1 |
3064 |
2913 |
32.04 |
|
|
|
| 3 |
A1 |
3055 |
2905 |
75.25 |
|
|
|
| 4 |
A1 |
3036 |
2888 |
6.23 |
|
|
|
| 5 |
A1 |
1540 |
1465 |
7.03 |
|
|
|
| 6 |
A1 |
1524 |
1450 |
0.17 |
|
|
|
| 7 |
A1 |
1513 |
1439 |
0.00 |
|
|
|
| 8 |
A1 |
1441 |
1370 |
3.47 |
|
|
|
| 9 |
A1 |
1386 |
1318 |
0.02 |
|
|
|
| 10 |
A1 |
1182 |
1124 |
1.16 |
|
|
|
| 11 |
A1 |
1079 |
1026 |
0.62 |
|
|
|
| 12 |
A1 |
890 |
846 |
1.50 |
|
|
|
| 13 |
A1 |
403 |
383 |
0.01 |
|
|
|
| 14 |
A1 |
177 |
168 |
0.02 |
|
|
|
| 15 |
A2 |
3138 |
2984 |
0.00 |
|
|
|
| 16 |
A2 |
3085 |
2934 |
0.00 |
|
|
|
| 17 |
A2 |
1527 |
1452 |
0.00 |
|
|
|
| 18 |
A2 |
1349 |
1283 |
0.00 |
|
|
|
| 19 |
A2 |
1283 |
1220 |
0.00 |
|
|
|
| 20 |
A2 |
1004 |
955 |
0.00 |
|
|
|
| 21 |
A2 |
772 |
734 |
0.00 |
|
|
|
| 22 |
A2 |
253 |
241 |
0.00 |
|
|
|
| 23 |
A2 |
115 |
109 |
0.00 |
|
|
|
| 24 |
B1 |
3139 |
2985 |
109.64 |
|
|
|
| 25 |
B1 |
3101 |
2949 |
51.64 |
|
|
|
| 26 |
B1 |
3066 |
2915 |
0.41 |
|
|
|
| 27 |
B1 |
1526 |
1451 |
15.15 |
|
|
|
| 28 |
B1 |
1344 |
1278 |
0.62 |
|
|
|
| 29 |
B1 |
1222 |
1162 |
0.17 |
|
|
|
| 30 |
B1 |
876 |
833 |
1.81 |
|
|
|
| 31 |
B1 |
740 |
704 |
5.13 |
|
|
|
| 32 |
B1 |
244 |
232 |
0.00 |
|
|
|
| 33 |
B1 |
119 |
113 |
0.00 |
|
|
|
| 34 |
B2 |
3145 |
2990 |
29.84 |
|
|
|
| 35 |
B2 |
3063 |
2912 |
42.68 |
|
|
|
| 36 |
B2 |
3048 |
2899 |
0.02 |
|
|
|
| 37 |
B2 |
1534 |
1459 |
1.72 |
|
|
|
| 38 |
B2 |
1515 |
1441 |
0.83 |
|
|
|
| 39 |
B2 |
1436 |
1366 |
3.01 |
|
|
|
| 40 |
B2 |
1431 |
1361 |
0.74 |
|
|
|
| 41 |
B2 |
1302 |
1238 |
2.91 |
|
|
|
| 42 |
B2 |
1104 |
1050 |
0.66 |
|
|
|
| 43 |
B2 |
1061 |
1009 |
2.65 |
|
|
|
| 44 |
B2 |
949 |
903 |
3.41 |
|
|
|
| 45 |
B2 |
396 |
377 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35659.6 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 33912.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.298 |
|
|
|
| 2 |
C |
-0.309 |
|
|
|
| 3 |
C |
-0.309 |
|
|
|
| 4 |
C |
-0.513 |
|
|
|
| 5 |
C |
-0.513 |
|
|
|
| 6 |
H |
0.154 |
|
|
|
| 7 |
H |
0.154 |
|
|
|
| 8 |
H |
0.159 |
|
|
|
| 9 |
H |
0.159 |
|
|
|
| 10 |
H |
0.159 |
|
|
|
| 11 |
H |
0.159 |
|
|
|
| 12 |
H |
0.168 |
|
|
|
| 13 |
H |
0.168 |
|
|
|
| 14 |
H |
0.166 |
|
|
|
| 15 |
H |
0.166 |
|
|
|
| 16 |
H |
0.166 |
|
|
|
| 17 |
H |
0.166 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.055 |
0.055 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.056 |
0.000 |
0.000 |
| y |
0.000 |
-35.277 |
0.000 |
| z |
0.000 |
0.000 |
-34.837 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.001 |
0.000 |
0.000 |
| y |
0.000 |
-0.831 |
0.000 |
| z |
0.000 |
0.000 |
-0.170 |
|
| Polar |
|---|
| 3z2-r2 | -0.340 |
|---|
| x2-y2 | 1.222 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.364 |
0.000 |
0.000 |
| y |
0.000 |
9.398 |
0.000 |
| z |
0.000 |
0.000 |
7.563 |
<r2> (average value of r
2) Å
2
| <r2> |
200.189 |
| (<r2>)1/2 |
14.149 |