Jump to
S1C2
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -196.461941 |
| Energy at 298.15K | -196.472098 |
| Nuclear repulsion energy | 169.699572 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3247 |
3106 |
20.95 |
|
|
|
| 2 |
A |
3165 |
3028 |
6.17 |
|
|
|
| 3 |
A |
3146 |
3010 |
32.87 |
|
|
|
| 4 |
A |
3131 |
2996 |
36.34 |
|
|
|
| 5 |
A |
3126 |
2991 |
53.37 |
|
|
|
| 6 |
A |
3093 |
2959 |
20.82 |
|
|
|
| 7 |
A |
3073 |
2940 |
7.82 |
|
|
|
| 8 |
A |
3051 |
2919 |
36.90 |
|
|
|
| 9 |
A |
3047 |
2915 |
27.51 |
|
|
|
| 10 |
A |
3022 |
2891 |
21.47 |
|
|
|
| 11 |
A |
1740 |
1664 |
10.23 |
|
|
|
| 12 |
A |
1533 |
1466 |
5.53 |
|
|
|
| 13 |
A |
1524 |
1458 |
7.23 |
|
|
|
| 14 |
A |
1517 |
1452 |
0.68 |
|
|
|
| 15 |
A |
1504 |
1439 |
2.73 |
|
|
|
| 16 |
A |
1470 |
1406 |
1.07 |
|
|
|
| 17 |
A |
1434 |
1372 |
2.23 |
|
|
|
| 18 |
A |
1399 |
1338 |
0.92 |
|
|
|
| 19 |
A |
1343 |
1285 |
0.56 |
|
|
|
| 20 |
A |
1333 |
1275 |
0.18 |
|
|
|
| 21 |
A |
1305 |
1249 |
3.72 |
|
|
|
| 22 |
A |
1279 |
1223 |
0.18 |
|
|
|
| 23 |
A |
1207 |
1155 |
0.52 |
|
|
|
| 24 |
A |
1126 |
1077 |
1.26 |
|
|
|
| 25 |
A |
1072 |
1026 |
2.19 |
|
|
|
| 26 |
A |
1045 |
1000 |
7.73 |
|
|
|
| 27 |
A |
1032 |
988 |
8.01 |
|
|
|
| 28 |
A |
965 |
924 |
0.78 |
|
|
|
| 29 |
A |
936 |
896 |
40.18 |
|
|
|
| 30 |
A |
899 |
860 |
6.97 |
|
|
|
| 31 |
A |
881 |
843 |
0.92 |
|
|
|
| 32 |
A |
752 |
720 |
3.18 |
|
|
|
| 33 |
A |
645 |
617 |
8.69 |
|
|
|
| 34 |
A |
434 |
415 |
1.28 |
|
|
|
| 35 |
A |
383 |
367 |
0.13 |
|
|
|
| 36 |
A |
251 |
240 |
0.02 |
|
|
|
| 37 |
A |
229 |
219 |
0.09 |
|
|
|
| 38 |
A |
104 |
100 |
0.03 |
|
|
|
| 39 |
A |
97 |
93 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30269.1 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 28958.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
2.479 |
-0.217 |
-0.303 |
| C2 |
1.414 |
0.342 |
0.270 |
| C3 |
0.073 |
-0.311 |
0.433 |
| C4 |
-1.054 |
0.447 |
-0.283 |
| C5 |
-2.424 |
-0.196 |
-0.079 |
| H6 |
3.424 |
0.312 |
-0.397 |
| H7 |
2.448 |
-1.232 |
-0.697 |
| H8 |
1.494 |
1.363 |
0.649 |
| H9 |
0.117 |
-1.344 |
0.063 |
| H10 |
-0.174 |
-0.374 |
1.505 |
| H11 |
-1.078 |
1.485 |
0.077 |
| H12 |
-0.822 |
0.501 |
-1.355 |
| H13 |
-3.209 |
0.361 |
-0.602 |
| H14 |
-2.439 |
-1.226 |
-0.458 |
| H15 |
-2.692 |
-0.232 |
0.984 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
| C1 | | 1.3329 | 2.5182 | 3.5957 | 4.9084 | 1.0871 | 1.0888 | 2.0919 | 2.6432 | 3.2144 | 3.9618 | 3.5387 | 5.7256 | 5.0229 | 5.3288 |
C2 | 1.3329 | | 1.5001 | 2.5315 | 3.8909 | 2.1186 | 2.1172 | 1.0923 | 2.1373 | 2.1348 | 2.7481 | 2.7686 | 4.7043 | 4.2226 | 4.2064 | C3 | 2.5182 | 1.5001 | | 1.5356 | 2.5515 | 3.5084 | 2.7871 | 2.2062 | 1.0983 | 1.1015 | 2.1628 | 2.1579 | 3.5064 | 2.8179 | 2.8202 | C4 | 3.5957 | 2.5315 | 1.5356 | | 1.5268 | 4.4821 | 3.9061 | 2.8637 | 2.1676 | 2.1550 | 1.0991 | 1.0979 | 2.1798 | 2.1783 | 2.1788 | C5 | 4.9084 | 3.8909 | 2.5515 | 1.5268 | | 5.8788 | 5.0191 | 4.2790 | 2.7913 | 2.7572 | 2.1598 | 2.1630 | 1.0958 | 1.0968 | 1.0970 | H6 | 1.0871 | 2.1186 | 3.5084 | 4.4821 | 5.8788 | | 1.8508 | 2.4347 | 3.7276 | 4.1274 | 4.6764 | 4.3571 | 6.6366 | 6.0616 | 6.2936 | H7 | 1.0888 | 2.1172 | 2.7871 | 3.9061 | 5.0191 | 1.8508 | | 3.0754 | 2.4552 | 3.5302 | 4.5181 | 3.7588 | 5.8780 | 4.8929 | 5.4996 | H8 | 2.0919 | 1.0923 | 2.2062 | 2.8637 | 4.2790 | 2.4347 | 3.0754 | | 3.0934 | 2.5552 | 2.6372 | 3.1817 | 4.9683 | 4.8366 | 4.4916 | H9 | 2.6432 | 2.1373 | 1.0983 | 2.1676 | 2.7913 | 3.7276 | 2.4552 | 3.0934 | | 1.7620 | 3.0711 | 2.5090 | 3.7961 | 2.6107 | 3.1577 | H10 | 3.2144 | 2.1348 | 1.1015 | 2.1550 | 2.7572 | 4.1274 | 3.5302 | 2.5552 | 1.7620 | | 2.5123 | 3.0596 | 3.7668 | 3.1158 | 2.5749 | H11 | 3.9618 | 2.7481 | 2.1628 | 1.0991 | 2.1598 | 4.6764 | 4.5181 | 2.6372 | 3.0711 | 2.5123 | | 1.7559 | 2.5031 | 3.0802 | 2.5254 | H12 | 3.5387 | 2.7686 | 2.1579 | 1.0979 | 2.1630 | 4.3571 | 3.7588 | 3.1817 | 2.5090 | 3.0596 | 1.7559 | | 2.5066 | 2.5294 | 3.0824 | H13 | 5.7256 | 4.7043 | 3.5064 | 2.1798 | 1.0958 | 6.6366 | 5.8780 | 4.9683 | 3.7961 | 3.7668 | 2.5031 | 2.5066 | | 1.7700 | 1.7704 | H14 | 5.0229 | 4.2226 | 2.8179 | 2.1783 | 1.0968 | 6.0616 | 4.8929 | 4.8366 | 2.6107 | 3.1158 | 3.0802 | 2.5294 | 1.7700 | | 1.7693 | H15 | 5.3288 | 4.2064 | 2.8202 | 2.1788 | 1.0970 | 6.2936 | 5.4996 | 4.4916 | 3.1577 | 2.5749 | 2.5254 | 3.0824 | 1.7704 | 1.7693 | |
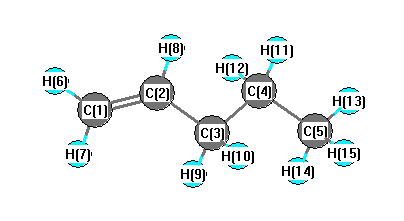 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
125.356 |
|
C1 |
C2 |
H8 |
118.885 |
| C2 |
C1 |
H6 |
121.865 |
|
C2 |
C1 |
H7 |
121.588 |
| C2 |
C3 |
C4 |
112.998 |
|
C2 |
C3 |
H9 |
109.712 |
| C2 |
C3 |
H10 |
109.323 |
|
C3 |
C2 |
H8 |
115.755 |
| C3 |
C4 |
C5 |
112.850 |
|
C3 |
C4 |
H11 |
109.227 |
| C3 |
C4 |
H12 |
108.916 |
|
C4 |
C3 |
H9 |
109.649 |
| C4 |
C3 |
H10 |
108.489 |
|
C4 |
C5 |
H13 |
111.377 |
| C4 |
C5 |
H14 |
111.196 |
|
C4 |
C5 |
H15 |
111.224 |
| C5 |
C4 |
H11 |
109.595 |
|
C5 |
C4 |
H12 |
109.911 |
| H6 |
C1 |
H7 |
116.546 |
|
H9 |
C3 |
H10 |
106.450 |
| H11 |
C4 |
H12 |
106.115 |
|
H13 |
C5 |
H14 |
107.655 |
| H13 |
C5 |
H15 |
107.684 |
|
H14 |
C5 |
H15 |
107.516 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.390 |
|
|
|
| 2 |
C |
-0.070 |
|
|
|
| 3 |
C |
-0.352 |
|
|
|
| 4 |
C |
-0.307 |
|
|
|
| 5 |
C |
-0.509 |
|
|
|
| 6 |
H |
0.162 |
|
|
|
| 7 |
H |
0.158 |
|
|
|
| 8 |
H |
0.152 |
|
|
|
| 9 |
H |
0.163 |
|
|
|
| 10 |
H |
0.166 |
|
|
|
| 11 |
H |
0.159 |
|
|
|
| 12 |
H |
0.168 |
|
|
|
| 13 |
H |
0.169 |
|
|
|
| 14 |
H |
0.166 |
|
|
|
| 15 |
H |
0.165 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.355 |
0.080 |
0.064 |
0.370 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.037 |
0.166 |
-0.576 |
| y |
0.166 |
-32.006 |
1.051 |
| z |
-0.576 |
1.051 |
-33.589 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.239 |
0.166 |
-0.576 |
| y |
0.166 |
1.307 |
1.051 |
| z |
-0.576 |
1.051 |
-1.068 |
|
| Polar |
|---|
| 3z2-r2 | -2.136 |
|---|
| x2-y2 | -1.030 |
|---|
| xy | 0.166 |
|---|
| xz | -0.576 |
|---|
| yz | 1.051 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.342 |
-0.334 |
-0.949 |
| y |
-0.334 |
7.197 |
0.631 |
| z |
-0.949 |
0.631 |
6.247 |
<r2> (average value of r
2) Å
2
| <r2> |
178.592 |
| (<r2>)1/2 |
13.364 |
Jump to
S1C1
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -196.461568 |
| Energy at 298.15K | -196.471806 |
| HF Energy | -196.461568 |
| Nuclear repulsion energy | 173.068646 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3246 |
3106 |
20.01 |
|
|
|
| 2 |
A |
3165 |
3028 |
6.07 |
|
|
|
| 3 |
A |
3142 |
3006 |
33.89 |
|
|
|
| 4 |
A |
3139 |
3003 |
33.91 |
|
|
|
| 5 |
A |
3127 |
2992 |
38.37 |
|
|
|
| 6 |
A |
3087 |
2953 |
31.74 |
|
|
|
| 7 |
A |
3077 |
2943 |
22.67 |
|
|
|
| 8 |
A |
3055 |
2922 |
22.72 |
|
|
|
| 9 |
A |
3043 |
2912 |
33.95 |
|
|
|
| 10 |
A |
3029 |
2898 |
26.46 |
|
|
|
| 11 |
A |
1738 |
1663 |
8.18 |
|
|
|
| 12 |
A |
1532 |
1466 |
3.34 |
|
|
|
| 13 |
A |
1524 |
1458 |
9.43 |
|
|
|
| 14 |
A |
1514 |
1449 |
0.49 |
|
|
|
| 15 |
A |
1503 |
1438 |
6.13 |
|
|
|
| 16 |
A |
1470 |
1407 |
1.11 |
|
|
|
| 17 |
A |
1437 |
1375 |
6.47 |
|
|
|
| 18 |
A |
1389 |
1329 |
3.48 |
|
|
|
| 19 |
A |
1370 |
1311 |
1.52 |
|
|
|
| 20 |
A |
1333 |
1275 |
1.23 |
|
|
|
| 21 |
A |
1301 |
1244 |
0.11 |
|
|
|
| 22 |
A |
1270 |
1215 |
1.97 |
|
|
|
| 23 |
A |
1203 |
1151 |
0.46 |
|
|
|
| 24 |
A |
1108 |
1060 |
3.45 |
|
|
|
| 25 |
A |
1087 |
1040 |
4.20 |
|
|
|
| 26 |
A |
1039 |
994 |
10.90 |
|
|
|
| 27 |
A |
1022 |
977 |
1.96 |
|
|
|
| 28 |
A |
945 |
904 |
2.51 |
|
|
|
| 29 |
A |
937 |
896 |
42.37 |
|
|
|
| 30 |
A |
892 |
854 |
0.41 |
|
|
|
| 31 |
A |
875 |
837 |
2.22 |
|
|
|
| 32 |
A |
778 |
745 |
4.45 |
|
|
|
| 33 |
A |
645 |
617 |
8.07 |
|
|
|
| 34 |
A |
434 |
415 |
0.44 |
|
|
|
| 35 |
A |
388 |
372 |
1.78 |
|
|
|
| 36 |
A |
299 |
286 |
0.20 |
|
|
|
| 37 |
A |
222 |
213 |
0.13 |
|
|
|
| 38 |
A |
123 |
118 |
0.02 |
|
|
|
| 39 |
A |
93 |
89 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30289.9 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 28978.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.107 |
0.568 |
0.119 |
| C2 |
-1.186 |
-0.245 |
-0.399 |
| C3 |
-0.069 |
-0.896 |
0.364 |
| C4 |
1.328 |
-0.521 |
-0.159 |
| C5 |
1.671 |
0.956 |
0.024 |
| H6 |
-2.894 |
1.006 |
-0.491 |
| H7 |
-2.114 |
0.823 |
1.177 |
| H8 |
-1.221 |
-0.469 |
-1.467 |
| H9 |
-0.183 |
-1.989 |
0.299 |
| H10 |
-0.148 |
-0.635 |
1.428 |
| H11 |
2.074 |
-1.139 |
0.357 |
| H12 |
1.396 |
-0.786 |
-1.224 |
| H13 |
2.669 |
1.183 |
-0.368 |
| H14 |
0.949 |
1.598 |
-0.493 |
| H15 |
1.657 |
1.234 |
1.085 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
| C1 | | 1.3332 | 2.5213 | 3.6140 | 3.7990 | 1.0872 | 1.0887 | 2.0921 | 3.2050 | 2.6460 | 4.5224 | 3.9884 | 4.8399 | 3.2830 | 3.9435 |
C2 | 1.3332 | | 1.5008 | 2.5397 | 3.1270 | 2.1188 | 2.1175 | 1.0926 | 2.1294 | 2.1370 | 3.4633 | 2.7636 | 4.1102 | 2.8218 | 3.5315 | C3 | 2.5213 | 1.5008 | | 1.5382 | 2.5634 | 3.5108 | 2.7920 | 2.2052 | 1.1005 | 1.0985 | 2.1567 | 2.1632 | 3.5145 | 2.8270 | 2.8351 | C4 | 3.6140 | 2.5397 | 1.5382 | | 1.5265 | 4.5011 | 3.9284 | 2.8650 | 2.1559 | 2.1703 | 1.0981 | 1.0993 | 2.1777 | 2.1778 | 2.1759 | C5 | 3.7990 | 3.1270 | 2.5634 | 1.5265 | | 4.5937 | 3.9582 | 3.5515 | 3.4901 | 2.7946 | 2.1591 | 2.1601 | 1.0959 | 1.0955 | 1.0971 | H6 | 1.0872 | 2.1188 | 3.5108 | 4.5011 | 4.5937 | | 1.8508 | 2.4344 | 4.1158 | 3.7304 | 5.4768 | 4.7061 | 5.5666 | 3.8884 | 4.8219 | H7 | 1.0887 | 2.1175 | 2.7920 | 3.9284 | 3.9582 | 1.8508 | | 3.0757 | 3.5218 | 2.4603 | 4.6964 | 4.5463 | 5.0383 | 3.5735 | 3.7944 | H8 | 2.0921 | 1.0926 | 2.2052 | 2.8650 | 3.5515 | 2.4344 | 3.0757 | | 2.5512 | 3.0925 | 3.8250 | 2.6469 | 4.3662 | 3.1513 | 4.2070 | H9 | 3.2050 | 2.1294 | 1.1005 | 2.1559 | 3.4901 | 4.1158 | 3.5218 | 2.5512 | | 1.7628 | 2.4119 | 2.5017 | 4.3167 | 3.8437 | 3.7934 | H10 | 2.6460 | 2.1370 | 1.0985 | 2.1703 | 2.7946 | 3.7304 | 2.4603 | 3.0925 | 1.7628 | | 2.5174 | 3.0723 | 3.8030 | 3.1435 | 2.6212 | H11 | 4.5224 | 3.4633 | 2.1567 | 1.0981 | 2.1591 | 5.4768 | 4.6964 | 3.8250 | 2.4119 | 2.5174 | | 1.7558 | 2.5041 | 3.0788 | 2.5169 | H12 | 3.9884 | 2.7636 | 2.1632 | 1.0993 | 2.1601 | 4.7061 | 4.5463 | 2.6469 | 2.5017 | 3.0723 | 1.7558 | | 2.4958 | 2.5333 | 3.0789 | H13 | 4.8399 | 4.1102 | 3.5145 | 2.1777 | 1.0959 | 5.5666 | 5.0383 | 4.3662 | 4.3167 | 3.8030 | 2.5041 | 2.4958 | | 1.7732 | 1.7708 | H14 | 3.2830 | 2.8218 | 2.8270 | 2.1778 | 1.0955 | 3.8884 | 3.5735 | 3.1513 | 3.8437 | 3.1435 | 3.0788 | 2.5333 | 1.7732 | | 1.7675 | H15 | 3.9435 | 3.5315 | 2.8351 | 2.1759 | 1.0971 | 4.8219 | 3.7944 | 4.2070 | 3.7934 | 2.6212 | 2.5169 | 3.0789 | 1.7708 | 1.7675 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
125.561 |
|
C1 |
C2 |
H8 |
118.848 |
| C2 |
C1 |
H6 |
121.855 |
|
C2 |
C1 |
H7 |
121.601 |
| C2 |
C3 |
C4 |
113.376 |
|
C2 |
C3 |
H9 |
108.915 |
| C2 |
C3 |
H10 |
109.633 |
|
C3 |
C2 |
H8 |
115.590 |
| C3 |
C4 |
C5 |
113.534 |
|
C3 |
C4 |
H11 |
108.637 |
| C3 |
C4 |
H12 |
109.067 |
|
C4 |
C3 |
H9 |
108.437 |
| C4 |
C3 |
H10 |
109.671 |
|
C4 |
C5 |
H13 |
111.227 |
| C4 |
C5 |
H14 |
111.255 |
|
C4 |
C5 |
H15 |
111.015 |
| C5 |
C4 |
H11 |
109.624 |
|
C5 |
C4 |
H12 |
109.627 |
| H6 |
C1 |
H7 |
116.542 |
|
H9 |
C3 |
H10 |
106.574 |
| H11 |
C4 |
H12 |
106.077 |
|
H13 |
C5 |
H14 |
108.033 |
| H13 |
C5 |
H15 |
107.703 |
|
H14 |
C5 |
H15 |
107.438 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.389 |
|
|
|
| 2 |
C |
-0.068 |
|
|
|
| 3 |
C |
-0.355 |
|
|
|
| 4 |
C |
-0.312 |
|
|
|
| 5 |
C |
-0.505 |
|
|
|
| 6 |
H |
0.162 |
|
|
|
| 7 |
H |
0.158 |
|
|
|
| 8 |
H |
0.151 |
|
|
|
| 9 |
H |
0.169 |
|
|
|
| 10 |
H |
0.163 |
|
|
|
| 11 |
H |
0.161 |
|
|
|
| 12 |
H |
0.159 |
|
|
|
| 13 |
H |
0.165 |
|
|
|
| 14 |
H |
0.177 |
|
|
|
| 15 |
H |
0.163 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.258 |
-0.213 |
-0.051 |
0.339 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.294 |
-1.217 |
0.348 |
| y |
-1.217 |
-33.408 |
0.421 |
| z |
0.348 |
0.421 |
-31.630 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.775 |
-1.217 |
0.348 |
| y |
-1.217 |
-0.946 |
0.421 |
| z |
0.348 |
0.421 |
1.721 |
|
| Polar |
|---|
| 3z2-r2 | 3.442 |
|---|
| x2-y2 | 0.114 |
|---|
| xy | -1.217 |
|---|
| xz | 0.348 |
|---|
| yz | 0.421 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.065 |
-1.421 |
-0.201 |
| y |
-1.421 |
7.220 |
0.452 |
| z |
-0.201 |
0.452 |
7.215 |
<r2> (average value of r
2) Å
2
| <r2> |
153.260 |
| (<r2>)1/2 |
12.380 |
