Jump to
S1C2
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -195.849983 |
| Energy at 298.15K | -195.860203 |
| HF Energy | -195.139317 |
| Nuclear repulsion energy | 169.500582 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3251 |
3096 |
21.60 |
|
|
|
| 2 |
A |
3172 |
3021 |
5.29 |
|
|
|
| 3 |
A |
3154 |
3004 |
28.17 |
|
|
|
| 4 |
A |
3137 |
2987 |
37.20 |
|
|
|
| 5 |
A |
3132 |
2983 |
52.55 |
|
|
|
| 6 |
A |
3098 |
2950 |
24.82 |
|
|
|
| 7 |
A |
3078 |
2931 |
4.77 |
|
|
|
| 8 |
A |
3059 |
2913 |
26.72 |
|
|
|
| 9 |
A |
3054 |
2908 |
35.81 |
|
|
|
| 10 |
A |
3037 |
2892 |
18.09 |
|
|
|
| 11 |
A |
1743 |
1660 |
5.67 |
|
|
|
| 12 |
A |
1560 |
1486 |
4.80 |
|
|
|
| 13 |
A |
1549 |
1475 |
6.14 |
|
|
|
| 14 |
A |
1545 |
1471 |
0.65 |
|
|
|
| 15 |
A |
1538 |
1464 |
2.17 |
|
|
|
| 16 |
A |
1493 |
1422 |
1.17 |
|
|
|
| 17 |
A |
1466 |
1396 |
2.22 |
|
|
|
| 18 |
A |
1432 |
1364 |
0.38 |
|
|
|
| 19 |
A |
1362 |
1297 |
0.50 |
|
|
|
| 20 |
A |
1349 |
1285 |
0.08 |
|
|
|
| 21 |
A |
1330 |
1267 |
3.09 |
|
|
|
| 22 |
A |
1302 |
1240 |
0.14 |
|
|
|
| 23 |
A |
1231 |
1173 |
0.37 |
|
|
|
| 24 |
A |
1149 |
1094 |
0.77 |
|
|
|
| 25 |
A |
1094 |
1042 |
1.13 |
|
|
|
| 26 |
A |
1060 |
1010 |
3.24 |
|
|
|
| 27 |
A |
1026 |
977 |
14.40 |
|
|
|
| 28 |
A |
979 |
933 |
1.00 |
|
|
|
| 29 |
A |
930 |
885 |
35.43 |
|
|
|
| 30 |
A |
914 |
870 |
4.15 |
|
|
|
| 31 |
A |
898 |
855 |
0.55 |
|
|
|
| 32 |
A |
761 |
725 |
2.04 |
|
|
|
| 33 |
A |
645 |
615 |
7.72 |
|
|
|
| 34 |
A |
443 |
422 |
1.02 |
|
|
|
| 35 |
A |
392 |
373 |
0.14 |
|
|
|
| 36 |
A |
255 |
242 |
0.01 |
|
|
|
| 37 |
A |
237 |
225 |
0.09 |
|
|
|
| 38 |
A |
106 |
101 |
0.02 |
|
|
|
| 39 |
A |
94 |
89 |
0.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30527.2 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 29071.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
2.477 |
-0.227 |
-0.305 |
| C2 |
1.414 |
0.352 |
0.269 |
| C3 |
0.070 |
-0.303 |
0.446 |
| C4 |
-1.052 |
0.446 |
-0.288 |
| C5 |
-2.423 |
-0.200 |
-0.082 |
| H6 |
3.425 |
0.295 |
-0.417 |
| H7 |
2.437 |
-1.249 |
-0.681 |
| H8 |
1.499 |
1.380 |
0.630 |
| H9 |
0.116 |
-1.342 |
0.087 |
| H10 |
-0.178 |
-0.351 |
1.519 |
| H11 |
-1.079 |
1.490 |
0.059 |
| H12 |
-0.812 |
0.482 |
-1.360 |
| H13 |
-3.208 |
0.348 |
-0.617 |
| H14 |
-2.427 |
-1.236 |
-0.446 |
| H15 |
-2.692 |
-0.220 |
0.983 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
| C1 | | 1.3390 | 2.5226 | 3.5922 | 4.9048 | 1.0882 | 1.0898 | 2.1011 | 2.6395 | 3.2232 | 3.9649 | 3.5266 | 5.7223 | 5.0089 | 5.3264 |
C2 | 1.3390 | | 1.5056 | 2.5298 | 3.8924 | 2.1251 | 2.1241 | 1.0935 | 2.1410 | 2.1429 | 2.7483 | 2.7620 | 4.7063 | 4.2178 | 4.2065 | C3 | 2.5226 | 1.5056 | | 1.5351 | 2.5499 | 3.5155 | 2.7880 | 2.2151 | 1.1002 | 1.1021 | 2.1635 | 2.1582 | 3.5067 | 2.8114 | 2.8143 | C4 | 3.5922 | 2.5298 | 1.5351 | | 1.5296 | 4.4812 | 3.8989 | 2.8672 | 2.1678 | 2.1596 | 1.1004 | 1.0994 | 2.1833 | 2.1784 | 2.1787 | C5 | 4.9048 | 3.8924 | 2.5499 | 1.5296 | | 5.8783 | 5.0081 | 4.2875 | 2.7890 | 2.7613 | 2.1640 | 2.1667 | 1.0972 | 1.0980 | 1.0981 | H6 | 1.0882 | 2.1251 | 3.5155 | 4.4812 | 5.8783 | | 1.8518 | 2.4465 | 3.7254 | 4.1407 | 4.6835 | 4.3452 | 6.6362 | 6.0493 | 6.2958 | H7 | 1.0898 | 2.1241 | 2.7880 | 3.8989 | 5.0081 | 1.8518 | | 3.0845 | 2.4467 | 3.5336 | 4.5178 | 3.7444 | 5.8672 | 4.8704 | 5.4895 | H8 | 2.1011 | 1.0935 | 2.2151 | 2.8672 | 4.2875 | 2.4465 | 3.0845 | | 3.1004 | 2.5686 | 2.6421 | 3.1795 | 4.9773 | 4.8391 | 4.4991 | H9 | 2.6395 | 2.1410 | 1.1002 | 2.1678 | 2.7890 | 3.7254 | 2.4467 | 3.1004 | | 1.7653 | 3.0732 | 2.5071 | 3.7950 | 2.6013 | 3.1535 | H10 | 3.2232 | 2.1429 | 1.1021 | 2.1596 | 2.7613 | 4.1407 | 3.5336 | 2.5686 | 1.7653 | | 2.5166 | 3.0641 | 3.7728 | 3.1154 | 2.5737 | H11 | 3.9649 | 2.7483 | 2.1635 | 1.1004 | 2.1640 | 4.6835 | 4.5178 | 2.6421 | 3.0732 | 2.5166 | | 1.7604 | 2.5089 | 3.0828 | 2.5255 | H12 | 3.5266 | 2.7620 | 2.1582 | 1.0994 | 2.1667 | 4.3452 | 3.7444 | 3.1795 | 2.5071 | 3.0641 | 1.7604 | | 2.5118 | 2.5291 | 3.0847 | H13 | 5.7223 | 4.7063 | 3.5067 | 2.1833 | 1.0972 | 6.6362 | 5.8672 | 4.9773 | 3.7950 | 3.7728 | 2.5089 | 2.5118 | | 1.7741 | 1.7745 | H14 | 5.0089 | 4.2178 | 2.8114 | 2.1784 | 1.0980 | 6.0493 | 4.8704 | 4.8391 | 2.6013 | 3.1154 | 3.0828 | 2.5291 | 1.7741 | | 1.7735 | H15 | 5.3264 | 4.2065 | 2.8143 | 2.1787 | 1.0981 | 6.2958 | 5.4895 | 4.4991 | 3.1535 | 2.5737 | 2.5255 | 3.0847 | 1.7745 | 1.7735 | |
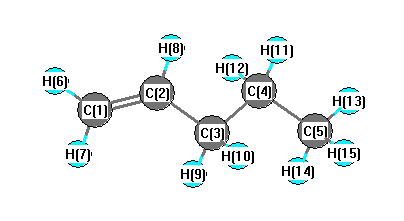 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
124.840 |
|
C1 |
C2 |
H8 |
119.145 |
| C2 |
C1 |
H6 |
121.880 |
|
C2 |
C1 |
H7 |
121.641 |
| C2 |
C3 |
C4 |
112.600 |
|
C2 |
C3 |
H9 |
109.512 |
| C2 |
C3 |
H10 |
109.551 |
|
C3 |
C2 |
H8 |
116.009 |
| C3 |
C4 |
C5 |
112.618 |
|
C3 |
C4 |
H11 |
109.238 |
| C3 |
C4 |
H12 |
108.886 |
|
C4 |
C3 |
H9 |
109.588 |
| C4 |
C3 |
H10 |
108.842 |
|
C4 |
C5 |
H13 |
111.378 |
| C4 |
C5 |
H14 |
110.932 |
|
C4 |
C5 |
H15 |
110.953 |
| C5 |
C4 |
H11 |
109.660 |
|
C5 |
C4 |
H12 |
109.928 |
| H6 |
C1 |
H7 |
116.478 |
|
H9 |
C3 |
H10 |
106.564 |
| H11 |
C4 |
H12 |
106.309 |
|
H13 |
C5 |
H14 |
107.837 |
| H13 |
C5 |
H15 |
107.870 |
|
H14 |
C5 |
H15 |
107.714 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -195.849945 |
| Energy at 298.15K | -195.860253 |
| HF Energy | -195.138714 |
| Nuclear repulsion energy | 173.009149 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3251 |
3096 |
20.56 |
|
|
|
| 2 |
A |
3172 |
3021 |
5.08 |
|
|
|
| 3 |
A |
3150 |
3000 |
29.79 |
|
|
|
| 4 |
A |
3148 |
2998 |
34.57 |
|
|
|
| 5 |
A |
3133 |
2983 |
38.58 |
|
|
|
| 6 |
A |
3093 |
2946 |
32.74 |
|
|
|
| 7 |
A |
3084 |
2937 |
25.62 |
|
|
|
| 8 |
A |
3063 |
2917 |
18.31 |
|
|
|
| 9 |
A |
3052 |
2906 |
31.82 |
|
|
|
| 10 |
A |
3041 |
2896 |
24.19 |
|
|
|
| 11 |
A |
1742 |
1659 |
4.58 |
|
|
|
| 12 |
A |
1558 |
1484 |
3.32 |
|
|
|
| 13 |
A |
1550 |
1476 |
8.87 |
|
|
|
| 14 |
A |
1544 |
1470 |
0.82 |
|
|
|
| 15 |
A |
1536 |
1463 |
3.69 |
|
|
|
| 16 |
A |
1493 |
1422 |
1.29 |
|
|
|
| 17 |
A |
1469 |
1399 |
5.38 |
|
|
|
| 18 |
A |
1418 |
1351 |
2.28 |
|
|
|
| 19 |
A |
1396 |
1329 |
1.21 |
|
|
|
| 20 |
A |
1349 |
1284 |
1.09 |
|
|
|
| 21 |
A |
1325 |
1262 |
0.19 |
|
|
|
| 22 |
A |
1291 |
1229 |
1.21 |
|
|
|
| 23 |
A |
1227 |
1169 |
0.40 |
|
|
|
| 24 |
A |
1131 |
1078 |
2.90 |
|
|
|
| 25 |
A |
1107 |
1054 |
2.79 |
|
|
|
| 26 |
A |
1043 |
993 |
3.00 |
|
|
|
| 27 |
A |
1026 |
977 |
12.80 |
|
|
|
| 28 |
A |
958 |
912 |
1.58 |
|
|
|
| 29 |
A |
930 |
886 |
37.20 |
|
|
|
| 30 |
A |
909 |
866 |
0.19 |
|
|
|
| 31 |
A |
888 |
845 |
1.21 |
|
|
|
| 32 |
A |
793 |
755 |
2.92 |
|
|
|
| 33 |
A |
647 |
616 |
7.12 |
|
|
|
| 34 |
A |
441 |
420 |
0.38 |
|
|
|
| 35 |
A |
402 |
383 |
1.43 |
|
|
|
| 36 |
A |
304 |
289 |
0.19 |
|
|
|
| 37 |
A |
233 |
222 |
0.06 |
|
|
|
| 38 |
A |
122 |
116 |
0.02 |
|
|
|
| 39 |
A |
90 |
86 |
0.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30553.7 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 29096.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.099 |
0.567 |
0.123 |
| C2 |
-1.179 |
-0.251 |
-0.406 |
| C3 |
-0.062 |
-0.907 |
0.363 |
| C4 |
1.332 |
-0.519 |
-0.157 |
| C5 |
1.648 |
0.967 |
0.024 |
| H6 |
-2.886 |
1.013 |
-0.482 |
| H7 |
-2.100 |
0.816 |
1.183 |
| H8 |
-1.216 |
-0.471 |
-1.476 |
| H9 |
-0.174 |
-2.001 |
0.300 |
| H10 |
-0.145 |
-0.642 |
1.428 |
| H11 |
2.087 |
-1.122 |
0.367 |
| H12 |
1.408 |
-0.787 |
-1.222 |
| H13 |
2.654 |
1.205 |
-0.345 |
| H14 |
0.929 |
1.594 |
-0.517 |
| H15 |
1.603 |
1.248 |
1.085 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
| C1 | | 1.3392 | 2.5256 | 3.6093 | 3.7695 | 1.0882 | 1.0897 | 2.1012 | 3.2135 | 2.6421 | 4.5204 | 3.9920 | 4.8180 | 3.2613 | 3.8847 |
C2 | 1.3392 | | 1.5063 | 2.5373 | 3.1078 | 2.1255 | 2.1239 | 1.0939 | 2.1375 | 2.1406 | 3.4675 | 2.7647 | 4.1002 | 2.8039 | 3.4933 | C3 | 2.5256 | 1.5063 | | 1.5375 | 2.5595 | 3.5180 | 2.7919 | 2.2142 | 1.1013 | 1.1004 | 2.1598 | 2.1643 | 3.5124 | 2.8308 | 2.8169 | C4 | 3.6093 | 2.5373 | 1.5375 | | 1.5297 | 4.4994 | 3.9184 | 2.8692 | 2.1619 | 2.1698 | 1.0996 | 1.1006 | 2.1804 | 2.1809 | 2.1760 | C5 | 3.7695 | 3.1078 | 2.5595 | 1.5297 | | 4.5625 | 3.9257 | 3.5382 | 3.4935 | 2.7885 | 2.1621 | 2.1648 | 1.0972 | 1.0964 | 1.0984 | H6 | 1.0882 | 2.1255 | 3.5180 | 4.4994 | 4.5625 | | 1.8520 | 2.4463 | 4.1289 | 3.7279 | 5.4784 | 4.7144 | 5.5449 | 3.8598 | 4.7601 | H7 | 1.0897 | 2.1239 | 2.7919 | 3.9184 | 3.9257 | 1.8520 | | 3.0844 | 3.5246 | 2.4506 | 4.6851 | 4.5448 | 5.0079 | 3.5596 | 3.7286 | H8 | 2.1012 | 1.0939 | 2.2142 | 2.8692 | 3.5382 | 2.4463 | 3.0844 | | 2.5645 | 3.0995 | 3.8379 | 2.6543 | 4.3661 | 3.1288 | 4.1781 | H9 | 3.2135 | 2.1375 | 1.1013 | 2.1619 | 3.4935 | 4.1289 | 3.5246 | 2.5645 | | 1.7660 | 2.4272 | 2.5079 | 4.3234 | 3.8484 | 3.7851 | H10 | 2.6421 | 2.1406 | 1.1004 | 2.1698 | 2.7885 | 3.7279 | 2.4506 | 3.0995 | 1.7660 | | 2.5175 | 3.0741 | 3.7931 | 3.1524 | 2.5970 | H11 | 4.5204 | 3.4675 | 2.1598 | 1.0996 | 2.1621 | 5.4784 | 4.6851 | 3.8379 | 2.4272 | 2.5175 | | 1.7602 | 2.4988 | 3.0822 | 2.5228 | H12 | 3.9920 | 2.7647 | 2.1643 | 1.1006 | 2.1648 | 4.7144 | 4.5448 | 2.6543 | 2.5079 | 3.0741 | 1.7602 | | 2.5084 | 2.5293 | 3.0819 | H13 | 4.8180 | 4.1002 | 3.5124 | 2.1804 | 1.0972 | 5.5449 | 5.0079 | 4.3661 | 4.3234 | 3.7931 | 2.4988 | 2.5084 | | 1.7759 | 1.7747 | H14 | 3.2613 | 2.8039 | 2.8308 | 2.1809 | 1.0964 | 3.8598 | 3.5596 | 3.1288 | 3.8484 | 3.1524 | 3.0822 | 2.5293 | 1.7759 | | 1.7715 | H15 | 3.8847 | 3.4933 | 2.8169 | 2.1760 | 1.0984 | 4.7601 | 3.7286 | 4.1781 | 3.7851 | 2.5970 | 2.5228 | 3.0819 | 1.7747 | 1.7715 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
125.040 |
|
C1 |
C2 |
H8 |
119.098 |
| C2 |
C1 |
H6 |
121.893 |
|
C2 |
C1 |
H7 |
121.608 |
| C2 |
C3 |
C4 |
112.942 |
|
C2 |
C3 |
H9 |
109.133 |
| C2 |
C3 |
H10 |
109.424 |
|
C3 |
C2 |
H8 |
115.861 |
| C3 |
C4 |
C5 |
113.123 |
|
C3 |
C4 |
H11 |
108.840 |
| C3 |
C4 |
H12 |
109.125 |
|
C4 |
C3 |
H9 |
108.907 |
| C4 |
C3 |
H10 |
109.568 |
|
C4 |
C5 |
H13 |
111.139 |
| C4 |
C5 |
H14 |
111.233 |
|
C4 |
C5 |
H15 |
110.716 |
| C5 |
C4 |
H11 |
109.553 |
|
C5 |
C4 |
H12 |
109.697 |
| H6 |
C1 |
H7 |
116.498 |
|
H9 |
C3 |
H10 |
106.669 |
| H11 |
C4 |
H12 |
106.268 |
|
H13 |
C5 |
H14 |
108.113 |
| H13 |
C5 |
H15 |
107.858 |
|
H14 |
C5 |
H15 |
107.634 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
