return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

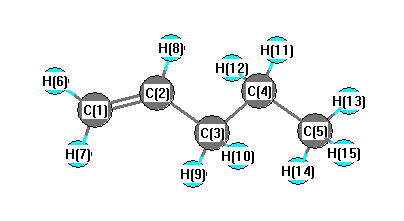
All results from a given calculation for CH2CHCH2CH2CH3 (1-pentene)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-196.268395
Energy at 298.15K-196.278611
HF Energy-196.054362
Nuclear repulsion energy169.743003
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3271 3105 20.80      
2 A 3189 3027 5.55      
3 A 3171 3010 29.48      
4 A 3152 2992 37.46      
5 A 3148 2988 54.19      
6 A 3114 2955 23.19      
7 A 3093 2935 7.40      
8 A 3075 2919 32.80      
9 A 3069 2914 29.38      
10 A 3047 2892 20.92      
11 A 1735 1647 7.40      
12 A 1559 1480 4.03      
13 A 1550 1471 6.00      
14 A 1544 1465 0.48      
15 A 1531 1453 2.22      
16 A 1491 1416 0.91      
17 A 1459 1385 1.39      
18 A 1420 1348 0.81      
19 A 1360 1291 0.49      
20 A 1348 1280 0.26      
21 A 1326 1259 4.70      
22 A 1298 1232 0.16      
23 A 1225 1163 0.39      
24 A 1140 1082 0.86      
25 A 1078 1024 2.37      
26 A 1054 1001 5.04      
27 A 1039 986 9.97      
28 A 972 923 0.64      
29 A 938 890 35.98      
30 A 908 862 5.35      
31 A 894 849 0.43      
32 A 763 724 2.30      
33 A 649 616 7.69      
34 A 440 418 1.24      
35 A 387 368 0.11      
36 A 255 242 0.02      
37 A 235 223 0.09      
38 A 106 101 0.02      
39 A 98 93 0.05      

Unscaled Zero Point Vibrational Energy (zpe) 30566.0 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 29013.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.60466 0.07059 0.06958

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.474 -0.224 -0.307
C2 1.415 0.347 0.272
C3 0.072 -0.303 0.446
C4 -1.051 0.444 -0.290
C5 -2.422 -0.198 -0.081
H6 3.419 0.298 -0.415
H7 2.432 -1.239 -0.691
H8 1.501 1.369 0.641
H9 0.117 -1.340 0.091
H10 -0.179 -0.348 1.515
H11 -1.076 1.486 0.052
H12 -0.813 0.477 -1.359
H13 -3.204 0.348 -0.616
H14 -2.431 -1.232 -0.441
H15 -2.692 -0.214 0.980

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.33532.51853.58784.90181.08471.08632.09332.63823.22083.95673.52245.71545.00965.3237
C21.33531.50212.53093.89192.11882.11651.08992.13602.13732.74762.76464.70374.21824.2048
C32.51851.50211.53642.55163.50752.78232.20831.09661.09902.16172.15673.50512.81332.8162
C43.58782.53091.53641.52884.47363.88962.86992.16592.15511.09691.09582.17992.17632.1767
C54.90183.89192.55161.52885.87165.00274.28642.78952.75712.16032.16321.09371.09461.0947
H61.08472.11883.50754.47365.87161.84732.43673.72034.13344.67224.33976.62606.04666.2884
H71.08632.11652.78233.88965.00271.84733.07312.44633.53304.50423.73225.85654.87025.4865
H82.09331.08992.20832.86994.28642.43673.07313.09122.55612.64633.18664.97644.83734.4947
H92.63822.13601.09662.16592.78953.72032.44633.09121.76033.06732.50423.79202.60533.1536
H103.22082.13731.09902.15512.75714.13343.53302.55611.76032.51113.05673.76523.11112.5724
H113.95672.74762.16171.09692.16034.67224.50422.64633.06732.51111.75442.50403.07662.5223
H123.52242.76462.15671.09582.16324.33973.73223.18662.50423.05671.75442.50692.52633.0787
H135.71544.70373.50512.17991.09376.62605.85654.97643.79203.76522.50402.50691.76781.7682
H145.00964.21822.81332.17631.09466.04664.87024.83732.60533.11113.07662.52631.76781.7670
H155.32374.20482.81622.17671.09476.28845.48654.49473.15362.57242.52233.07871.76821.7670

picture of 1-pentene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 125.049 C1 C2 H8 118.999
C2 C1 H6 121.874 C2 C1 H7 121.508
C2 C3 C4 112.803 C2 C3 H9 109.580
C2 C3 H10 109.533 C3 C2 H8 115.944
C3 C4 C5 112.702 C3 C4 H11 109.214
C3 C4 H12 108.884 C4 C3 H9 109.558
C4 C3 H10 108.579 C4 C5 H13 111.373
C4 C5 H14 111.031 C4 C5 H15 111.061
C5 C4 H11 109.627 C5 C4 H12 109.920
H6 C1 H7 116.617 H9 C3 H10 106.592
H11 C4 H12 106.286 H13 C5 H14 107.771
H13 C5 H15 107.805 H14 C5 H15 107.632
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.368      
2 C -0.075      
3 C -0.316      
4 C -0.274      
5 C -0.461      
6 H 0.153      
7 H 0.149      
8 H 0.146      
9 H 0.149      
10 H 0.150      
11 H 0.142      
12 H 0.152      
13 H 0.153      
14 H 0.150      
15 H 0.149      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 10.046 -0.333 -0.933
y -0.333 7.108 0.630
z -0.933 0.630 6.158


<r2> (average value of r2) Å2
<r2> 178.600
(<r2>)1/2 13.364