return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C(NH2)H2CH2CH2CH3 (1-Butanamine)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-213.070564
Energy at 298.15K-213.083307
Nuclear repulsion energy185.075255
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3465 3300 1.78      
2 A' 3136 2986 38.58      
3 A' 3059 2913 51.76      
4 A' 3059 2913 37.45      
5 A' 3047 2902 16.82      
6 A' 3030 2885 18.04      
7 A' 1728 1645 24.67      
8 A' 1563 1488 3.67      
9 A' 1552 1478 0.28      
10 A' 1543 1469 0.89      
11 A' 1539 1466 0.38      
12 A' 1467 1397 2.67      
13 A' 1456 1386 5.94      
14 A' 1416 1349 4.49      
15 A' 1320 1257 6.92      
16 A' 1181 1125 9.02      
17 A' 1125 1071 6.14      
18 A' 1104 1051 7.83      
19 A' 1059 1009 39.89      
20 A' 948 903 98.41      
21 A' 917 874 75.14      
22 A' 439 418 5.43      
23 A' 401 382 0.40      
24 A' 186 177 2.54      
25 A" 3553 3383 0.91      
26 A" 3132 2983 58.07      
27 A" 3106 2958 60.90      
28 A" 3085 2938 9.96      
29 A" 3061 2915 3.70      
30 A" 1550 1476 6.32      
31 A" 1425 1357 1.15      
32 A" 1361 1296 0.28      
33 A" 1344 1280 0.06      
34 A" 1271 1210 0.00      
35 A" 1073 1021 0.66      
36 A" 945 900 0.03      
37 A" 805 767 0.59      
38 A" 748 712 2.60      
39 A" 306 291 56.89      
40 A" 257 245 0.17      
41 A" 125 119 0.06      
42 A" 111 105 3.48      

Unscaled Zero Point Vibrational Energy (zpe) 33497.4 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 31899.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.59368 0.06463 0.06152

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 2.542 0.417 0.000
C2 1.337 -0.423 0.000
C3 0.000 0.326 0.000
C4 -1.214 -0.605 0.000
C5 -2.545 0.150 0.000
H6 2.518 1.035 0.813
H7 2.518 1.035 -0.813
H8 1.390 -1.080 0.880
H9 1.390 -1.080 -0.880
H10 -0.043 0.986 -0.882
H11 -0.043 0.986 0.882
H12 -1.165 -1.265 0.880
H13 -1.165 -1.265 -0.880
H14 -3.398 -0.541 0.000
H15 -2.632 0.791 0.887
H16 -2.632 0.791 -0.887

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
N11.46892.54383.89335.09431.02171.02172.08432.08432.79002.79004.16454.16456.01665.26285.2628
C21.46891.53252.55803.92402.04512.04511.09951.09952.16022.16022.78232.78234.73614.24404.2440
C32.54381.53251.53062.55122.73982.73982.16422.16421.10231.10232.15912.15913.50662.81592.8159
C43.89332.55801.53061.52994.15784.15782.78952.78952.16412.16411.10061.10062.18422.17862.1786
C55.09433.92402.55121.52995.20435.20434.21544.21542.78192.78192.16322.16321.09721.09801.0980
H61.02172.04512.73984.15785.20431.62682.39862.93513.07202.56274.34264.66066.17635.15665.4291
H71.02172.04512.73984.15785.20431.62682.93512.39862.56273.07204.66064.34266.17635.42915.1566
H82.08431.09952.16422.78954.21542.39862.93511.75913.07012.51452.56113.10724.89734.43594.7746
H92.08431.09952.16422.78954.21542.93512.39861.75912.51453.07013.10722.56114.89734.77464.4359
H102.79002.16021.10232.16412.78193.07202.56273.07012.51451.76393.07062.51483.79013.14152.5963
H112.79002.16021.10232.16412.78192.56273.07202.51453.07011.76392.51483.07063.79012.59633.1415
H124.16452.78232.15911.10062.16324.34264.66062.56113.10723.07062.51481.75992.50682.52583.0823
H134.16452.78232.15911.10062.16324.66064.34263.10722.56112.51483.07061.75992.50683.08232.5258
H146.01664.73613.50662.18421.09726.17636.17634.89734.89733.79013.79012.50682.50681.77431.7743
H155.26284.24402.81592.17861.09805.15665.42914.43594.77463.14152.59632.52583.08231.77431.7734
H165.26284.24402.81592.17861.09805.42915.15664.77464.43592.59633.14153.08232.52581.77431.7734

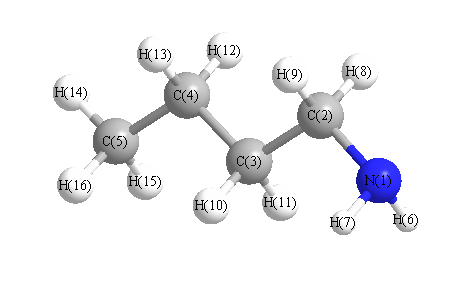
picture of 1-Butanamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C3 115.874 N1 C2 H8 107.613
N1 C2 H9 107.613 C2 N1 H6 109.068
C2 N1 H7 109.068 C2 C3 C4 113.250
C2 C3 H10 109.054 C2 C3 H11 109.054
C3 C2 H8 109.525 C3 C2 H9 109.525
C3 C4 C5 112.937 C3 C4 H12 109.189
C3 C4 H13 109.189 C4 C3 H10 109.483
C4 C3 H11 109.483 C4 C5 H14 111.417
C4 C5 H15 110.926 C4 C5 H16 110.926
C5 C4 H12 109.565 C5 C4 H13 109.565
H6 N1 H7 105.520 H8 C2 H9 106.248
H10 C3 H11 106.278 H12 C4 H13 106.170
H14 C5 H15 107.852 H14 C5 H16 107.852
H15 C5 H16 107.712
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability