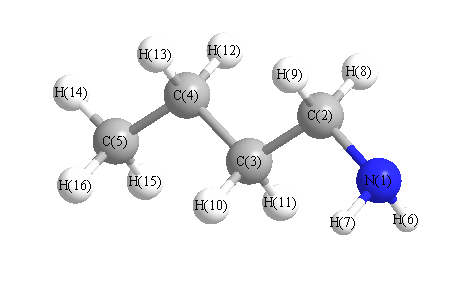
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3498 |
3327 |
1.34 |
|
|
|
| 2 |
A' |
3146 |
2991 |
37.35 |
|
|
|
| 3 |
A' |
3064 |
2914 |
37.78 |
|
|
|
| 4 |
A' |
3059 |
2909 |
74.76 |
|
|
|
| 5 |
A' |
3050 |
2901 |
8.20 |
|
|
|
| 6 |
A' |
3030 |
2882 |
16.70 |
|
|
|
| 7 |
A' |
1709 |
1625 |
23.82 |
|
|
|
| 8 |
A' |
1538 |
1462 |
4.81 |
|
|
|
| 9 |
A' |
1526 |
1452 |
0.49 |
|
|
|
| 10 |
A' |
1515 |
1441 |
0.44 |
|
|
|
| 11 |
A' |
1509 |
1435 |
0.52 |
|
|
|
| 12 |
A' |
1439 |
1368 |
2.44 |
|
|
|
| 13 |
A' |
1428 |
1358 |
8.32 |
|
|
|
| 14 |
A' |
1396 |
1328 |
4.33 |
|
|
|
| 15 |
A' |
1300 |
1236 |
6.13 |
|
|
|
| 16 |
A' |
1167 |
1109 |
6.11 |
|
|
|
| 17 |
A' |
1122 |
1067 |
9.92 |
|
|
|
| 18 |
A' |
1091 |
1037 |
5.54 |
|
|
|
| 19 |
A' |
1042 |
991 |
31.41 |
|
|
|
| 20 |
A' |
932 |
887 |
62.29 |
|
|
|
| 21 |
A' |
891 |
847 |
141.31 |
|
|
|
| 22 |
A' |
431 |
410 |
5.85 |
|
|
|
| 23 |
A' |
394 |
375 |
0.59 |
|
|
|
| 24 |
A' |
180 |
171 |
2.44 |
|
|
|
| 25 |
A" |
3588 |
3412 |
0.21 |
|
|
|
| 26 |
A" |
3139 |
2985 |
55.53 |
|
|
|
| 27 |
A" |
3108 |
2955 |
68.17 |
|
|
|
| 28 |
A" |
3089 |
2938 |
5.04 |
|
|
|
| 29 |
A" |
3063 |
2913 |
3.42 |
|
|
|
| 30 |
A" |
1527 |
1452 |
7.66 |
|
|
|
| 31 |
A" |
1413 |
1344 |
1.12 |
|
|
|
| 32 |
A" |
1347 |
1281 |
0.33 |
|
|
|
| 33 |
A" |
1326 |
1261 |
0.01 |
|
|
|
| 34 |
A" |
1244 |
1183 |
0.03 |
|
|
|
| 35 |
A" |
1061 |
1009 |
0.64 |
|
|
|
| 36 |
A" |
930 |
884 |
0.01 |
|
|
|
| 37 |
A" |
793 |
754 |
0.51 |
|
|
|
| 38 |
A" |
743 |
706 |
4.27 |
|
|
|
| 39 |
A" |
312 |
297 |
56.04 |
|
|
|
| 40 |
A" |
252 |
240 |
0.10 |
|
|
|
| 41 |
A" |
127 |
120 |
0.27 |
|
|
|
| 42 |
A" |
114 |
108 |
3.15 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 33313.9 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 31681.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.738 |
|
|
|
| 2 |
C |
-0.197 |
|
|
|
| 3 |
C |
-0.316 |
|
|
|
| 4 |
C |
-0.313 |
|
|
|
| 5 |
C |
-0.512 |
|
|
|
| 6 |
H |
0.310 |
|
|
|
| 7 |
H |
0.310 |
|
|
|
| 8 |
H |
0.166 |
|
|
|
| 9 |
H |
0.166 |
|
|
|
| 10 |
H |
0.149 |
|
|
|
| 11 |
H |
0.149 |
|
|
|
| 12 |
H |
0.161 |
|
|
|
| 13 |
H |
0.161 |
|
|
|
| 14 |
H |
0.170 |
|
|
|
| 15 |
H |
0.167 |
|
|
|
| 16 |
H |
0.167 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.176 |
0.908 |
0.000 |
1.486 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.699 |
3.141 |
0.000 |
| y |
3.141 |
-32.341 |
0.000 |
| z |
0.000 |
0.000 |
-31.825 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-8.616 |
3.141 |
0.000 |
| y |
3.141 |
3.921 |
0.000 |
| z |
0.000 |
0.000 |
4.695 |
|
| Polar |
|---|
| 3z2-r2 | 9.389 |
|---|
| x2-y2 | -8.358 |
|---|
| xy | 3.141 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.512 |
0.389 |
0.000 |
| y |
0.389 |
7.089 |
0.000 |
| z |
0.000 |
0.000 |
6.969 |
<r2> (average value of r
2) Å
2
| <r2> |
193.323 |
| (<r2>)1/2 |
13.904 |