return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CH2CH2CN (Butanenitrile)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-210.590453
Energy at 298.15K-210.597870
HF Energy-209.997080
Nuclear repulsion energy154.473622
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3245 2997 26.84      
2 A' 3182 2939 22.76      
3 A' 3175 2933 0.35      
4 A' 3160 2919 19.96      
5 A' 2495 2304 6.73      
6 A' 1603 1481 6.37      
7 A' 1589 1468 1.48      
8 A' 1572 1452 3.12      
9 A' 1518 1402 2.74      
10 A' 1486 1373 2.27      
11 A' 1385 1280 1.39      
12 A' 1184 1093 3.02      
13 A' 1115 1030 0.01      
14 A' 1007 930 3.45      
15 A' 930 859 2.20      
16 A' 554 512 1.04      
17 A' 364 336 0.19      
18 A' 174 161 7.06      
19 A" 3243 2995 50.08      
20 A" 3227 2981 0.48      
21 A" 3211 2966 0.01      
22 A" 1594 1473 8.48      
23 A" 1403 1296 0.04      
24 A" 1336 1234 0.05      
25 A" 1201 1109 0.14      
26 A" 926 855 0.09      
27 A" 781 721 3.49      
28 A" 403 372 0.96      
29 A" 253 234 0.01      
30 A" 104 96 4.53      

Unscaled Zero Point Vibrational Energy (zpe) 23709.3 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 21900.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
0.80260 0.07567 0.07190

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 -2.604 0.355 0.000
C2 -1.460 0.486 0.000
C3 0.000 0.634 0.000
C4 0.721 -0.717 0.000
C5 2.234 -0.543 0.000
H6 0.283 1.217 0.876
H7 0.283 1.217 -0.876
H8 0.409 -1.288 0.874
H9 0.409 -1.288 -0.874
H10 2.735 -1.510 0.000
H11 2.567 0.005 0.881
H12 2.567 0.005 -0.881

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12
N11.15112.61853.49354.92073.13703.13703.54103.54105.65505.25735.2573
C21.15111.46752.49103.83502.08292.08292.72072.72074.64544.15074.1507
C32.61851.46751.53142.52541.08981.08982.15012.15013.47482.78632.7863
C43.49352.49101.53141.52322.16812.16811.08961.08962.16412.16942.1694
C54.92073.83502.52541.52322.77032.77032.15662.15661.08841.08981.0898
H63.13702.08291.08982.16812.77031.75262.50743.05813.77032.58643.1270
H73.13702.08291.08982.16812.77031.75263.05812.50743.77033.12702.5864
H83.54102.72072.15011.08962.15662.50743.05811.74862.49492.51583.0677
H93.54102.72072.15011.08962.15663.05812.50741.74862.49493.06772.5158
H105.65504.64543.47482.16411.08843.77033.77032.49492.49491.76011.7601
H115.25734.15072.78632.16941.08982.58643.12702.51583.06771.76011.7624
H125.25734.15072.78632.16941.08983.12702.58643.06772.51581.76011.7624

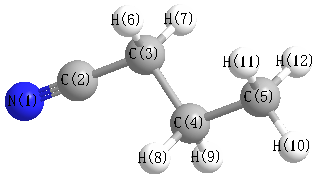
picture of Butanenitrile state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C3 179.266 C2 C3 C4 112.310
C2 C3 H6 108.171 C2 C3 H7 108.171
C3 C4 C5 111.532 C3 C4 H8 109.073
C3 C4 H9 109.073 C4 C3 H6 110.477
C4 C3 H7 110.477 C4 C5 H10 110.825
C4 C5 H11 111.165 C4 C5 H12 111.165
C5 C4 H8 110.155 C5 C4 H9 110.155
H6 C3 H7 107.050 H8 C4 H9 106.722
H10 C5 H11 107.810 H10 C5 H12 107.810
H11 C5 H12 107.918
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability