return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H2N2 (Malononitrile)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-224.762700
Energy at 298.15K-224.764337
HF Energy-224.526845
Nuclear repulsion energy124.154603
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3095 2937 0.26      
2 A1 2320 2202 0.03      
3 A1 1496 1420 11.88      
4 A1 910 864 5.55      
5 A1 578 548 0.31      
6 A1 145 138 10.26      
7 A2 1269 1204 0.00      
8 A2 366 347 0.00      
9 B1 3139 2979 1.69      
10 B1 964 915 0.58      
11 B1 338 321 0.43      
12 B2 2319 2201 0.21      
13 B2 1375 1306 3.61      
14 B2 997 947 7.04      
15 B2 364 345 0.63      

Unscaled Zero Point Vibrational Energy (zpe) 9836.8 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 9337.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.68649 0.09711 0.08645

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.845
C2 0.000 1.221 0.027
C3 0.000 -1.221 0.027
H4 0.882 0.000 1.494
H5 -0.882 0.000 1.494
N6 0.000 2.205 -0.599
N7 0.000 -2.205 -0.599

Atom - Atom Distances (Å)
  C1 C2 C3 H4 H5 N6 N7
C11.46961.46961.09511.09512.63592.6359
C21.46962.44252.10232.10231.16653.4834
C31.46962.44252.10232.10233.48341.1665
H41.09512.10232.10231.76473.16553.1655
H51.09512.10232.10231.76473.16553.1655
N62.63591.16653.48343.16553.16554.4109
N72.63593.48341.16653.16553.16554.4109

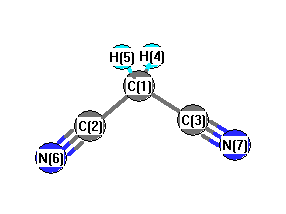
picture of Malononitrile state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 N6 178.660 C1 C3 N7 178.660
C2 C1 C3 112.400 C2 C1 H4 109.240
C2 C1 H5 109.240 C3 C1 H4 109.240
C3 C1 H5 109.240 H4 C1 H5 107.351
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.392      
2 C 0.359      
3 C 0.359      
4 H 0.269      
5 H 0.269      
6 N -0.432      
7 N -0.432      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 3.296 0.000 0.000
y 0.000 7.052 0.000
z 0.000 0.000 4.553


<r2> (average value of r2) Å2
<r2> 121.180
(<r2>)1/2 11.008