return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H9SH (1-Butanethiol)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-555.393374
Energy at 298.15K 
HF Energy-554.805104
Nuclear repulsion energy223.895021
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3236 2989 36.84      
2 A' 3190 2946 32.59      
3 A' 3159 2918 37.60      
4 A' 3156 2915 10.59      
5 A' 3148 2908 16.21      
6 A' 2842 2625 24.53      
7 A' 1608 1486 4.46      
8 A' 1596 1474 1.57      
9 A' 1590 1469 3.64      
10 A' 1587 1466 0.33      
11 A' 1516 1401 3.12      
12 A' 1502 1387 5.94      
13 A' 1433 1324 13.15      
14 A' 1344 1242 20.34      
15 A' 1205 1113 2.31      
16 A' 1138 1051 0.48      
17 A' 1101 1017 0.29      
18 A' 988 913 3.25      
19 A' 911 841 2.01      
20 A' 807 745 2.79      
21 A' 411 379 0.94      
22 A' 340 315 0.89      
23 A' 162 149 1.54      
24 A" 3247 2999 33.72      
25 A" 3233 2986 42.34      
26 A" 3203 2959 15.89      
27 A" 3181 2938 3.54      
28 A" 1595 1473 6.99      
29 A" 1409 1302 0.14      
30 A" 1388 1282 1.03      
31 A" 1311 1211 0.39      
32 A" 1156 1068 1.12      
33 A" 985 910 1.44      
34 A" 839 775 0.01      
35 A" 774 715 3.36      
36 A" 261 241 0.05      
37 A" 195 180 19.35      
38 A" 117 108 2.16      
39 A" 102 94 4.33      

Unscaled Zero Point Vibrational Energy (zpe) 30481.6 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 28155.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
0.53970 0.04466 0.04254

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 1.391 -1.837 0.000
C2 -0.218 -0.984 0.000
C3 0.000 0.523 0.000
C4 -1.315 1.296 0.000
C5 -1.105 2.805 0.000
H6 0.919 -3.087 0.000
H7 -0.783 -1.278 0.882
H8 -0.783 -1.278 -0.882
H9 0.588 0.804 -0.876
H10 0.588 0.804 0.876
H11 -1.903 1.009 0.874
H12 -1.903 1.009 -0.874
H13 -2.056 3.336 0.000
H14 -0.546 3.122 0.880
H15 -0.546 3.122 -0.880

Atom - Atom Distances (Å)
  S1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
S11.82182.73984.14025.27091.33542.41212.41212.89602.89604.44044.44046.21665.39615.3961
C21.82181.52272.52993.89142.39071.08781.08782.14792.14792.75202.75204.69464.21174.2117
C32.73981.52271.52562.53553.72482.15302.15301.09161.09162.15012.15013.48432.79752.7975
C44.14022.52991.52561.52394.91922.77242.77242.15222.15221.09211.09212.17062.16822.1682
C55.27093.89142.53551.52396.22974.18984.18982.76382.76382.15152.15151.08911.08991.0899
H61.33542.39073.72484.91926.22972.63552.63554.00144.00145.04995.04997.07836.43926.4392
H72.41211.08782.15302.77244.18982.63551.76363.05032.49312.54643.09334.86724.40634.7456
H82.41211.08782.15302.77244.18982.63551.76362.49313.05033.09332.54644.86724.74564.4063
H92.89602.14791.09162.15222.76384.00143.05032.49311.75123.05152.49993.76433.12112.5803
H102.89602.14791.09162.15222.76384.00142.49313.05031.75122.49993.05153.76432.58033.1211
H114.44042.75202.15011.09212.15155.04992.54643.09333.05152.49991.74862.49092.51163.0638
H124.44042.75202.15011.09212.15155.04993.09332.54642.49993.05151.74862.49093.06382.5116
H136.21664.69463.48432.17061.08917.07834.86724.86723.76433.76432.49092.49091.76111.7611
H145.39614.21172.79752.16821.08996.43924.40634.74563.12112.58032.51163.06381.76111.7607
H155.39614.21172.79752.16821.08996.43924.74564.40632.58033.12113.06382.51161.76111.7607

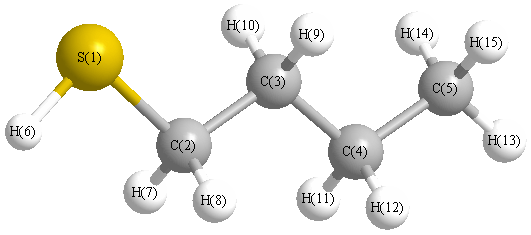
picture of 1-Butanethiol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C2 C3 109.684 S1 C2 H7 109.392
S1 C2 H8 109.392 C2 S1 H6 97.237
C2 C3 C4 112.188 C2 C3 H9 109.385
C2 C3 H10 109.385 C3 C2 H7 110.013
C3 C2 H8 110.013 C3 C4 C5 112.500
C3 C4 H11 109.331 C3 C4 H12 109.331
C4 C3 H9 109.527 C4 C3 H10 109.527
C4 C5 H13 111.243 C4 C5 H14 111.007
C4 C5 H15 111.007 C5 C4 H11 109.558
C5 C4 H12 109.558 H7 C2 H8 108.323
H9 C3 H10 106.675 H11 C4 H12 106.376
H13 C5 H14 107.841 H13 C5 H15 107.841
H14 C5 H15 107.748
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability