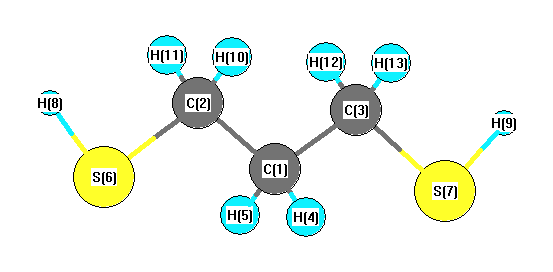
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3001 |
2951 |
53.07 |
|
|
|
| 2 |
A1 |
2986 |
2937 |
1.00 |
|
|
|
| 3 |
A1 |
2615 |
2572 |
14.92 |
|
|
|
| 4 |
A1 |
1484 |
1460 |
2.55 |
|
|
|
| 5 |
A1 |
1470 |
1446 |
0.07 |
|
|
|
| 6 |
A1 |
1274 |
1253 |
37.77 |
|
|
|
| 7 |
A1 |
1080 |
1062 |
2.52 |
|
|
|
| 8 |
A1 |
829 |
816 |
0.90 |
|
|
|
| 9 |
A1 |
771 |
759 |
1.27 |
|
|
|
| 10 |
A1 |
289 |
285 |
0.40 |
|
|
|
| 11 |
A1 |
123 |
121 |
0.18 |
|
|
|
| 12 |
A2 |
3045 |
2995 |
0.00 |
|
|
|
| 13 |
A2 |
1303 |
1281 |
0.00 |
|
|
|
| 14 |
A2 |
1093 |
1075 |
0.00 |
|
|
|
| 15 |
A2 |
823 |
809 |
0.00 |
|
|
|
| 16 |
A2 |
174 |
171 |
0.00 |
|
|
|
| 17 |
A2 |
85 |
84 |
0.00 |
|
|
|
| 18 |
B1 |
3060 |
3010 |
33.89 |
|
|
|
| 19 |
B1 |
3025 |
2975 |
0.02 |
|
|
|
| 20 |
B1 |
1252 |
1231 |
2.78 |
|
|
|
| 21 |
B1 |
1001 |
985 |
5.34 |
|
|
|
| 22 |
B1 |
746 |
734 |
5.09 |
|
|
|
| 23 |
B1 |
163 |
160 |
45.33 |
|
|
|
| 24 |
B1 |
98 |
96 |
0.04 |
|
|
|
| 25 |
B2 |
2992 |
2943 |
2.80 |
|
|
|
| 26 |
B2 |
2615 |
2572 |
33.60 |
|
|
|
| 27 |
B2 |
1464 |
1440 |
3.71 |
|
|
|
| 28 |
B2 |
1350 |
1328 |
22.13 |
|
|
|
| 29 |
B2 |
1213 |
1193 |
17.06 |
|
|
|
| 30 |
B2 |
1048 |
1031 |
0.08 |
|
|
|
| 31 |
B2 |
872 |
858 |
0.00 |
|
|
|
| 32 |
B2 |
690 |
679 |
3.74 |
|
|
|
| 33 |
B2 |
319 |
314 |
8.70 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22177.3 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 21811.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.306 |
|
|
|
| 2 |
C |
-0.442 |
|
|
|
| 3 |
C |
-0.442 |
|
|
|
| 4 |
H |
0.191 |
|
|
|
| 5 |
H |
0.191 |
|
|
|
| 6 |
S |
-0.082 |
|
|
|
| 7 |
S |
-0.082 |
|
|
|
| 8 |
H |
0.106 |
|
|
|
| 9 |
H |
0.106 |
|
|
|
| 10 |
H |
0.190 |
|
|
|
| 11 |
H |
0.190 |
|
|
|
| 12 |
H |
0.190 |
|
|
|
| 13 |
H |
0.190 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-3.079 |
3.079 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.378 |
0.000 |
0.000 |
| y |
0.000 |
-47.105 |
0.000 |
| z |
0.000 |
0.000 |
-46.040 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.805 |
0.000 |
0.000 |
| y |
0.000 |
0.604 |
0.000 |
| z |
0.000 |
0.000 |
2.201 |
|
| Polar |
|---|
| 3z2-r2 | 4.402 |
|---|
| x2-y2 | -2.273 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.053 |
0.000 |
0.000 |
| y |
0.000 |
13.508 |
0.000 |
| z |
0.000 |
0.000 |
8.665 |
<r2> (average value of r
2) Å
2
| <r2> |
349.367 |
| (<r2>)1/2 |
18.691 |