Jump to
S1C2
Vibrational Frequencies calculated at PBE1PBE/6-31G*
Geometric Data calculated at PBE1PBE/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at PBE1PBE/6-31G*
| | hartrees |
|---|
| Energy at 0K | -269.231364 |
| Energy at 298.15K | -269.240832 |
| HF Energy | -269.231364 |
| Nuclear repulsion energy | 193.972247 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3178 |
3020 |
22.19 |
|
|
|
| 2 |
A1 |
3013 |
2863 |
57.89 |
|
|
|
| 3 |
A1 |
2906 |
2762 |
57.02 |
|
|
|
| 4 |
A1 |
1582 |
1503 |
1.50 |
|
|
|
| 5 |
A1 |
1535 |
1459 |
3.37 |
|
|
|
| 6 |
A1 |
1501 |
1426 |
0.42 |
|
|
|
| 7 |
A1 |
1299 |
1235 |
12.96 |
|
|
|
| 8 |
A1 |
1206 |
1147 |
12.35 |
|
|
|
| 9 |
A1 |
1075 |
1022 |
58.67 |
|
|
|
| 10 |
A1 |
491 |
466 |
1.37 |
|
|
|
| 11 |
A1 |
220 |
209 |
2.03 |
|
|
|
| 12 |
A2 |
3062 |
2910 |
0.00 |
|
|
|
| 13 |
A2 |
1516 |
1441 |
0.00 |
|
|
|
| 14 |
A2 |
1269 |
1206 |
0.00 |
|
|
|
| 15 |
A2 |
1193 |
1134 |
0.00 |
|
|
|
| 16 |
A2 |
228 |
217 |
0.00 |
|
|
|
| 17 |
A2 |
85 |
81 |
0.00 |
|
|
|
| 18 |
B1 |
3063 |
2910 |
127.66 |
|
|
|
| 19 |
B1 |
2924 |
2778 |
141.05 |
|
|
|
| 20 |
B1 |
1517 |
1441 |
15.18 |
|
|
|
| 21 |
B1 |
1215 |
1155 |
20.02 |
|
|
|
| 22 |
B1 |
1165 |
1107 |
7.04 |
|
|
|
| 23 |
B1 |
239 |
227 |
5.82 |
|
|
|
| 24 |
B1 |
88 |
83 |
0.76 |
|
|
|
| 25 |
B2 |
3178 |
3020 |
24.75 |
|
|
|
| 26 |
B2 |
3011 |
2861 |
65.03 |
|
|
|
| 27 |
B2 |
1548 |
1471 |
0.00 |
|
|
|
| 28 |
B2 |
1518 |
1442 |
34.56 |
|
|
|
| 29 |
B2 |
1464 |
1391 |
91.10 |
|
|
|
| 30 |
B2 |
1257 |
1195 |
334.30 |
|
|
|
| 31 |
B2 |
1199 |
1139 |
104.02 |
|
|
|
| 32 |
B2 |
1053 |
1001 |
2.44 |
|
|
|
| 33 |
B2 |
432 |
410 |
3.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25113.7 cm
-1
Scaled (by 0.9503) Zero Point Vibrational Energy (zpe) 23865.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.306 |
| H2 |
-0.897 |
0.000 |
0.962 |
| H3 |
0.897 |
0.000 |
0.962 |
| O4 |
0.000 |
1.108 |
-0.527 |
| O5 |
0.000 |
-1.108 |
-0.527 |
| C6 |
0.000 |
2.307 |
0.198 |
| C7 |
0.000 |
-2.307 |
0.198 |
| H8 |
0.000 |
3.121 |
-0.530 |
| H9 |
0.000 |
-3.121 |
-0.530 |
| H10 |
-0.893 |
2.402 |
0.837 |
| H11 |
0.893 |
2.402 |
0.837 |
| H12 |
0.893 |
-2.402 |
0.837 |
| H13 |
-0.893 |
-2.402 |
0.837 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
O4 |
O5 |
C6 |
C7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.1120 | 1.1120 | 1.3860 | 1.3860 | 2.3099 | 2.3099 | 3.2309 | 3.2309 | 2.6170 | 2.6170 | 2.6170 | 2.6170 |
H2 | 1.1120 | | 1.7948 | 2.0617 | 2.0617 | 2.5909 | 2.5909 | 3.5739 | 3.5739 | 2.4050 | 2.9984 | 2.9984 | 2.4050 | H3 | 1.1120 | 1.7948 | | 2.0617 | 2.0617 | 2.5909 | 2.5909 | 3.5739 | 3.5739 | 2.9984 | 2.4050 | 2.4050 | 2.9984 | O4 | 1.3860 | 2.0617 | 2.0617 | | 2.2164 | 1.4015 | 3.4918 | 2.0129 | 4.2293 | 2.0812 | 2.0812 | 3.8701 | 3.8701 | O5 | 1.3860 | 2.0617 | 2.0617 | 2.2164 | | 3.4918 | 1.4015 | 4.2293 | 2.0129 | 3.8701 | 3.8701 | 2.0812 | 2.0812 | C6 | 2.3099 | 2.5909 | 2.5909 | 1.4015 | 3.4918 | | 4.6149 | 1.0919 | 5.4772 | 1.1020 | 1.1020 | 4.8355 | 4.8355 | C7 | 2.3099 | 2.5909 | 2.5909 | 3.4918 | 1.4015 | 4.6149 | | 5.4772 | 1.0919 | 4.8355 | 4.8355 | 1.1020 | 1.1020 | H8 | 3.2309 | 3.5739 | 3.5739 | 2.0129 | 4.2293 | 1.0919 | 5.4772 | | 6.2423 | 1.7842 | 1.7842 | 5.7592 | 5.7592 | H9 | 3.2309 | 3.5739 | 3.5739 | 4.2293 | 2.0129 | 5.4772 | 1.0919 | 6.2423 | | 5.7592 | 5.7592 | 1.7842 | 1.7842 | H10 | 2.6170 | 2.4050 | 2.9984 | 2.0812 | 3.8701 | 1.1020 | 4.8355 | 1.7842 | 5.7592 | | 1.7863 | 5.1249 | 4.8035 | H11 | 2.6170 | 2.9984 | 2.4050 | 2.0812 | 3.8701 | 1.1020 | 4.8355 | 1.7842 | 5.7592 | 1.7863 | | 4.8035 | 5.1249 | H12 | 2.6170 | 2.9984 | 2.4050 | 3.8701 | 2.0812 | 4.8355 | 1.1020 | 5.7592 | 1.7842 | 5.1249 | 4.8035 | | 1.7863 | H13 | 2.6170 | 2.4050 | 2.9984 | 3.8701 | 2.0812 | 4.8355 | 1.1020 | 5.7592 | 1.7842 | 4.8035 | 5.1249 | 1.7863 | |
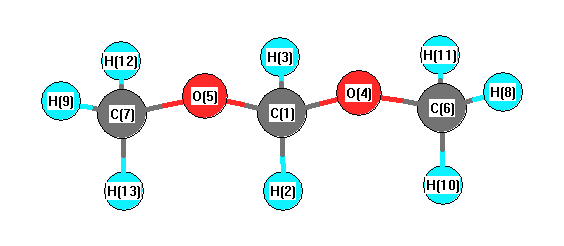 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O4 |
C6 |
111.924 |
|
C1 |
O5 |
C7 |
111.924 |
| H2 |
C1 |
H3 |
107.612 |
|
H2 |
C1 |
O4 |
110.772 |
| H2 |
C1 |
O5 |
110.772 |
|
H3 |
C1 |
O4 |
110.772 |
| H3 |
C1 |
O5 |
110.772 |
|
O4 |
C1 |
O5 |
106.179 |
| O4 |
C6 |
H8 |
107.011 |
|
O4 |
C6 |
H10 |
111.910 |
| O4 |
C6 |
H11 |
111.910 |
|
O5 |
C7 |
H9 |
107.011 |
| O5 |
C7 |
H12 |
111.910 |
|
O5 |
C7 |
H13 |
111.910 |
| H8 |
C6 |
H10 |
108.826 |
|
H8 |
C6 |
H11 |
108.826 |
| H9 |
C7 |
H12 |
108.826 |
|
H9 |
C7 |
H13 |
108.826 |
| H10 |
C6 |
H11 |
108.276 |
|
H12 |
C7 |
H13 |
108.276 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.196 |
|
|
|
| 2 |
H |
0.120 |
|
|
|
| 3 |
H |
0.120 |
|
|
|
| 4 |
O |
-0.449 |
|
|
|
| 5 |
O |
-0.449 |
|
|
|
| 6 |
C |
-0.277 |
|
|
|
| 7 |
C |
-0.277 |
|
|
|
| 8 |
H |
0.194 |
|
|
|
| 9 |
H |
0.194 |
|
|
|
| 10 |
H |
0.157 |
|
|
|
| 11 |
H |
0.157 |
|
|
|
| 12 |
H |
0.157 |
|
|
|
| 13 |
H |
0.157 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
2.395 |
2.395 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.723 |
0.000 |
0.000 |
| y |
0.000 |
-24.489 |
0.000 |
| z |
0.000 |
0.000 |
-33.058 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.950 |
0.000 |
0.000 |
| y |
0.000 |
7.902 |
0.000 |
| z |
0.000 |
0.000 |
-4.952 |
|
| Polar |
|---|
| 3z2-r2 | -9.904 |
|---|
| x2-y2 | -7.235 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.348 |
0.000 |
0.000 |
| y |
0.000 |
7.575 |
0.000 |
| z |
0.000 |
0.000 |
5.234 |
<r2> (average value of r
2) Å
2
| <r2> |
160.176 |
| (<r2>)1/2 |
12.656 |
