return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for NH(C2H5)2 (diethylamine)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-213.066504
Energy at 298.15K-213.079285
HF Energy-212.312313
Nuclear repulsion energy188.086019
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3474 3308 1.77      
2 A' 3156 3006 47.33      
3 A' 3133 2984 66.59      
4 A' 3081 2934 50.44      
5 A' 3063 2917 12.76      
6 A' 2971 2829 146.42      
7 A' 1581 1505 0.44      
8 A' 1550 1476 2.79      
9 A' 1541 1468 11.45      
10 A' 1479 1408 3.01      
11 A' 1439 1370 0.34      
12 A' 1353 1289 4.31      
13 A' 1275 1214 1.99      
14 A' 1200 1143 13.73      
15 A' 1095 1043 5.28      
16 A' 934 890 13.73      
17 A' 850 809 5.36      
18 A' 813 774 68.17      
19 A' 436 415 0.31      
20 A' 268 255 1.37      
21 A' 192 183 0.87      
22 A' 116 110 1.51      
23 A" 3156 3006 8.37      
24 A" 3133 2984 18.71      
25 A" 3077 2930 1.00      
26 A" 3063 2917 29.31      
27 A" 2966 2825 7.52      
28 A" 1570 1495 4.03      
29 A" 1554 1480 6.44      
30 A" 1546 1472 8.49      
31 A" 1524 1451 23.94      
32 A" 1462 1393 12.19      
33 A" 1397 1330 25.51      
34 A" 1317 1254 3.38      
35 A" 1195 1138 37.16      
36 A" 1144 1090 8.60      
37 A" 1097 1045 4.27      
38 A" 971 925 0.27      
39 A" 827 788 0.58      
40 A" 437 416 0.65      
41 A" 268 255 0.70      
42 A" 113 107 1.27      

Unscaled Zero Point Vibrational Energy (zpe) 33408.4 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 31814.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.58795 0.06999 0.06597

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.017 -0.290 0.000
C2 0.017 0.520 1.219
C3 0.017 0.520 -1.219
C4 0.017 -0.372 2.455
C5 0.017 -0.372 -2.455
H6 -0.825 -0.870 0.000
H7 -0.835 1.228 1.262
H8 0.930 1.134 1.206
H9 -0.835 1.228 -1.262
H10 0.930 1.134 -1.206
H11 0.061 0.228 3.373
H12 -0.894 -0.984 2.498
H13 0.880 -1.047 2.432
H14 0.061 0.228 -3.373
H15 -0.894 -0.984 -2.498
H16 0.880 -1.047 -2.432

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
N11.46351.46352.45632.45631.02292.14952.07692.14952.07693.41242.74792.68953.41242.74792.6895
C21.46352.43771.52443.78052.03201.10831.09932.71652.66282.17402.17442.16184.60104.11174.0659
C31.46352.43773.78051.52442.03202.71652.66281.10831.09934.60104.11174.06592.17402.17442.1618
C42.45631.52443.78054.90982.64282.16982.15804.13494.06251.09711.09861.09565.85855.07295.0085
C52.45633.78051.52444.90982.64284.13494.06252.16982.15805.85855.07295.00851.09711.09861.0956
H61.02292.03202.03202.64282.64282.44822.92392.44822.92393.65582.50132.97533.65582.50132.9753
H72.14951.10832.71652.16984.13492.44821.76792.52343.03552.50152.53393.07974.82484.36194.6648
H82.07691.09932.66282.15804.06252.92391.76793.03552.41302.50363.07832.50224.74814.64004.2425
H92.14952.71651.10834.13492.16982.44822.52343.03551.76794.82484.36194.66482.50152.53393.0797
H102.07692.66281.09934.06252.15802.92393.03552.41301.76794.74814.64004.24252.50363.07832.5022
H113.41242.17404.60101.09715.85853.65582.50152.50364.82484.74811.77331.78356.74536.06975.9995
H122.74792.17444.11171.09865.07292.50132.53393.07834.36194.64001.77331.77626.06974.99565.2398
H132.68952.16184.06591.09565.00852.97533.07972.50224.66484.24251.78351.77625.99955.23984.8646
H143.41244.60102.17405.85851.09713.65584.82484.74812.50152.50366.74536.06975.99951.77331.7835
H152.74794.11172.17445.07291.09862.50134.36194.64002.53393.07836.06974.99565.23981.77331.7762
H162.68954.06592.16185.00851.09562.97534.66484.24253.07972.50225.99955.23984.86461.78351.7762

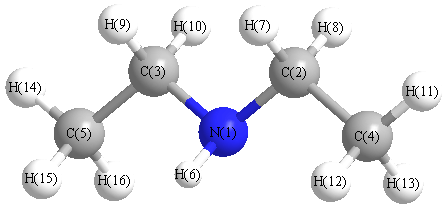
picture of diethylamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C4 110.573 N1 C2 H7 112.670
N1 C2 H8 107.423 N1 C3 C5 110.573
N1 C3 H9 112.670 N1 C3 H10 107.423
C2 N1 C3 112.784 C2 N1 H6 108.315
C2 C4 H11 111.005 C2 C4 H12 110.951
C2 C4 H13 110.122 C3 N1 H6 108.315
C3 C5 H14 111.005 C3 C5 H15 110.951
C3 C5 H16 110.122 C4 C2 H7 110.005
C4 C2 H8 109.609 C5 C3 H8 150.849
C5 C3 H10 109.609 H7 C2 H8 106.412
H9 C3 H10 106.412 H11 C4 H12 107.729
H11 C4 H13 108.851 H12 C4 H13 108.089
H14 C5 H15 107.729 H14 C5 H16 108.851
H15 C5 H16 108.089
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability