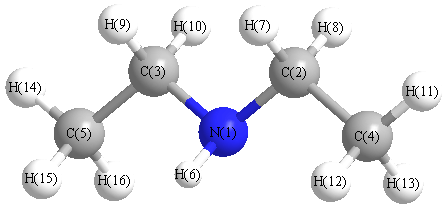
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3370 |
3315 |
3.73 |
|
|
|
| 2 |
A' |
3076 |
3025 |
40.43 |
|
|
|
| 3 |
A' |
3054 |
3003 |
61.25 |
|
|
|
| 4 |
A' |
2985 |
2935 |
60.28 |
|
|
|
| 5 |
A' |
2979 |
2930 |
15.41 |
|
|
|
| 6 |
A' |
2845 |
2798 |
186.35 |
|
|
|
| 7 |
A' |
1509 |
1484 |
0.82 |
|
|
|
| 8 |
A' |
1483 |
1459 |
2.12 |
|
|
|
| 9 |
A' |
1473 |
1448 |
13.94 |
|
|
|
| 10 |
A' |
1400 |
1377 |
3.32 |
|
|
|
| 11 |
A' |
1368 |
1345 |
0.24 |
|
|
|
| 12 |
A' |
1294 |
1273 |
2.98 |
|
|
|
| 13 |
A' |
1210 |
1190 |
1.12 |
|
|
|
| 14 |
A' |
1139 |
1120 |
12.09 |
|
|
|
| 15 |
A' |
1046 |
1029 |
5.77 |
|
|
|
| 16 |
A' |
894 |
880 |
12.44 |
|
|
|
| 17 |
A' |
808 |
795 |
1.93 |
|
|
|
| 18 |
A' |
762 |
749 |
73.46 |
|
|
|
| 19 |
A' |
418 |
411 |
0.17 |
|
|
|
| 20 |
A' |
256 |
252 |
0.91 |
|
|
|
| 21 |
A' |
181 |
178 |
0.91 |
|
|
|
| 22 |
A' |
109 |
107 |
1.04 |
|
|
|
| 23 |
A" |
3076 |
3025 |
11.25 |
|
|
|
| 24 |
A" |
3053 |
3003 |
14.47 |
|
|
|
| 25 |
A" |
2984 |
2935 |
23.48 |
|
|
|
| 26 |
A" |
2977 |
2928 |
6.14 |
|
|
|
| 27 |
A" |
2838 |
2791 |
18.77 |
|
|
|
| 28 |
A" |
1497 |
1472 |
3.84 |
|
|
|
| 29 |
A" |
1482 |
1458 |
3.24 |
|
|
|
| 30 |
A" |
1478 |
1453 |
12.27 |
|
|
|
| 31 |
A" |
1450 |
1426 |
17.17 |
|
|
|
| 32 |
A" |
1384 |
1361 |
13.16 |
|
|
|
| 33 |
A" |
1323 |
1301 |
38.77 |
|
|
|
| 34 |
A" |
1256 |
1235 |
5.02 |
|
|
|
| 35 |
A" |
1145 |
1126 |
48.20 |
|
|
|
| 36 |
A" |
1088 |
1070 |
11.87 |
|
|
|
| 37 |
A" |
1046 |
1029 |
4.46 |
|
|
|
| 38 |
A" |
931 |
915 |
0.19 |
|
|
|
| 39 |
A" |
793 |
780 |
0.40 |
|
|
|
| 40 |
A" |
416 |
409 |
0.54 |
|
|
|
| 41 |
A" |
257 |
253 |
0.71 |
|
|
|
| 42 |
A" |
116 |
114 |
1.26 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32123.7 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 31593.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.510 |
|
|
|
| 2 |
C |
-0.151 |
|
|
|
| 3 |
C |
-0.151 |
|
|
|
| 4 |
C |
-0.492 |
|
|
|
| 5 |
C |
-0.492 |
|
|
|
| 6 |
H |
0.288 |
|
|
|
| 7 |
H |
0.120 |
|
|
|
| 8 |
H |
0.153 |
|
|
|
| 9 |
H |
0.120 |
|
|
|
| 10 |
H |
0.153 |
|
|
|
| 11 |
H |
0.157 |
|
|
|
| 12 |
H |
0.151 |
|
|
|
| 13 |
H |
0.172 |
|
|
|
| 14 |
H |
0.157 |
|
|
|
| 15 |
H |
0.151 |
|
|
|
| 16 |
H |
0.172 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.788 |
0.331 |
0.000 |
0.854 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.451 |
1.653 |
0.000 |
| y |
1.653 |
-34.208 |
0.000 |
| z |
0.000 |
0.000 |
-32.932 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.119 |
1.653 |
0.000 |
| y |
1.653 |
-1.017 |
0.000 |
| z |
0.000 |
0.000 |
0.897 |
|
| Polar |
|---|
| 3z2-r2 | 1.795 |
|---|
| x2-y2 | 0.757 |
|---|
| xy | 1.653 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.967 |
0.106 |
0.000 |
| y |
0.106 |
7.374 |
0.000 |
| z |
0.000 |
0.000 |
9.586 |
<r2> (average value of r
2) Å
2
| <r2> |
188.748 |
| (<r2>)1/2 |
13.739 |