return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C2H3OC2H5 (Ethene, ethoxy-)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-232.154607
Energy at 298.15K-232.162976
HF Energy-231.925574
Nuclear repulsion energy173.823329
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3314 3146 7.65      
2 A 3223 3059 0.27      
3 A 3182 3020 25.52      
4 A 3175 3014 28.11      
5 A 3172 3011 20.40      
6 A 3095 2938 14.89      
7 A 3086 2929 47.69      
8 A 3044 2889 50.13      
9 A 1740 1652 132.34      
10 A 1574 1494 7.71      
11 A 1551 1472 3.47      
12 A 1535 1457 4.91      
13 A 1481 1406 8.01      
14 A 1460 1386 37.91      
15 A 1428 1356 57.99      
16 A 1368 1299 4.24      
17 A 1327 1260 0.35      
18 A 1245 1182 189.56      
19 A 1208 1147 29.19      
20 A 1174 1115 114.45      
21 A 1112 1056 28.65      
22 A 1005 954 8.32      
23 A 988 938 18.94      
24 A 872 828 6.13      
25 A 838 795 0.80      
26 A 830 788 52.10      
27 A 716 679 2.17      
28 A 501 475 2.49      
29 A 442 419 1.68      
30 A 264 251 0.70      
31 A 214 203 0.54      
32 A 84 80 3.53      
33 A 51 48 0.37      

Unscaled Zero Point Vibrational Energy (zpe) 25148.6 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 23871.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.79798 0.07869 0.07396

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -2.364 -0.317 0.202
H2 -2.289 -1.349 0.429
H3 -3.299 0.173 0.294
C4 -1.283 0.360 -0.204
H5 -1.343 1.400 -0.424
O6 -0.079 -0.241 -0.382
C7 0.993 0.446 0.280
H8 0.795 0.468 1.356
H9 1.059 1.477 -0.085
C10 2.298 -0.293 0.002
H11 2.222 -1.322 0.358
H12 2.504 -0.301 -1.069
H13 3.123 0.202 0.520

Atom - Atom Distances (Å)
  C1 H2 H3 C4 H5 O6 C7 H8 H9 C10 H11 H12 H13
C11.06001.05951.33862.09322.35943.44323.45353.87524.66664.69795.03075.5204
H21.06001.83172.08233.03022.60173.74383.69774.41164.72684.51235.12955.6306
H31.05951.83172.08512.41813.31604.30044.23964.56505.62425.72065.97936.4259
C41.33862.08232.08511.06451.35762.32832.60082.59743.64623.92873.93994.4677
H52.09323.03022.41811.06452.07232.61952.93392.42734.03834.55354.25534.7192
O62.35942.60173.31601.35762.07231.43512.07062.08212.40872.64802.67323.3558
C73.44323.74384.30042.32832.61951.43511.09431.09551.52592.15502.15872.1574
H83.45353.69774.23962.60082.93392.07061.09431.77932.16202.49763.06452.4881
H93.87524.41164.56502.59742.42732.08211.09551.77932.16193.06272.49232.4998
C104.66664.72685.62423.64624.03832.40871.52592.16202.16191.09131.08991.0922
H114.69794.51235.72063.92874.55352.64802.15502.49763.06271.09131.77661.7771
H125.03075.12955.97933.93994.25532.67322.15873.06452.49231.08991.77661.7772
H135.52045.63066.42594.46774.71923.35582.15742.48812.49981.09221.77711.7772

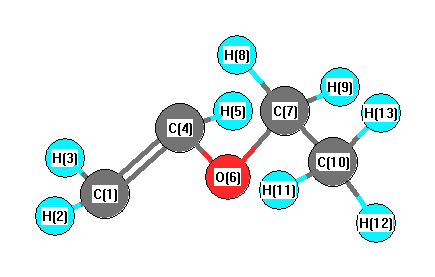
picture of Ethene, ethoxy- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C4 H5 120.734 C1 C4 O6 122.113
H2 C1 H3 119.592 H2 C1 C4 120.048
H3 C1 C4 120.360 C4 O6 C7 112.935
H5 C4 O6 117.138 O6 C7 H8 109.148
O6 C7 H9 110.002 O6 C7 C10 108.833
C7 C10 H11 109.734 C7 C10 H12 110.116
C7 C10 H13 109.874 H8 C7 H9 108.695
H8 C7 C10 110.112 H9 C7 C10 110.039
H11 C10 H12 109.077 H11 C10 H13 108.949
H12 C10 H13 109.066
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.427      
2 H 0.164      
3 H 0.155      
4 C 0.192      
5 H 0.144      
6 O -0.525      
7 C -0.016      
8 H 0.147      
9 H 0.142      
10 C -0.474      
11 H 0.172      
12 H 0.171      
13 H 0.153      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 9.865 0.749 -0.102
y 0.749 6.347 -0.256
z -0.102 -0.256 4.559


<r2> (average value of r2) Å2
<r2> 161.989
(<r2>)1/2 12.727