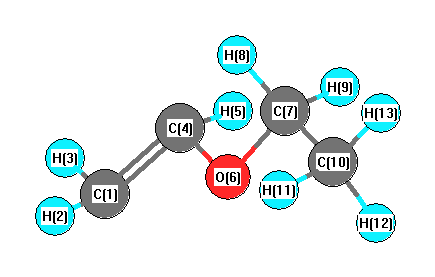
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3217 |
3163 |
7.21 |
|
|
|
| 2 |
A |
3126 |
3074 |
0.06 |
|
|
|
| 3 |
A |
3083 |
3032 |
22.92 |
|
|
|
| 4 |
A |
3080 |
3029 |
22.07 |
|
|
|
| 5 |
A |
3063 |
3013 |
25.00 |
|
|
|
| 6 |
A |
3000 |
2951 |
14.15 |
|
|
|
| 7 |
A |
2969 |
2920 |
48.56 |
|
|
|
| 8 |
A |
2931 |
2882 |
56.94 |
|
|
|
| 9 |
A |
1686 |
1658 |
138.62 |
|
|
|
| 10 |
A |
1502 |
1477 |
7.79 |
|
|
|
| 11 |
A |
1482 |
1458 |
5.53 |
|
|
|
| 12 |
A |
1466 |
1442 |
6.07 |
|
|
|
| 13 |
A |
1414 |
1391 |
4.09 |
|
|
|
| 14 |
A |
1391 |
1368 |
44.37 |
|
|
|
| 15 |
A |
1361 |
1338 |
49.98 |
|
|
|
| 16 |
A |
1315 |
1293 |
4.84 |
|
|
|
| 17 |
A |
1270 |
1249 |
0.17 |
|
|
|
| 18 |
A |
1207 |
1187 |
212.25 |
|
|
|
| 19 |
A |
1152 |
1133 |
14.04 |
|
|
|
| 20 |
A |
1128 |
1109 |
101.45 |
|
|
|
| 21 |
A |
1071 |
1054 |
29.15 |
|
|
|
| 22 |
A |
969 |
953 |
4.63 |
|
|
|
| 23 |
A |
942 |
926 |
18.26 |
|
|
|
| 24 |
A |
837 |
823 |
3.82 |
|
|
|
| 25 |
A |
803 |
790 |
0.81 |
|
|
|
| 26 |
A |
772 |
760 |
51.77 |
|
|
|
| 27 |
A |
680 |
669 |
3.97 |
|
|
|
| 28 |
A |
486 |
478 |
3.10 |
|
|
|
| 29 |
A |
427 |
420 |
1.71 |
|
|
|
| 30 |
A |
253 |
249 |
0.48 |
|
|
|
| 31 |
A |
204 |
201 |
0.37 |
|
|
|
| 32 |
A |
84 |
83 |
2.55 |
|
|
|
| 33 |
A |
37 |
36 |
0.23 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24204.0 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 23804.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.417 |
|
|
|
| 2 |
H |
0.152 |
|
|
|
| 3 |
H |
0.148 |
|
|
|
| 4 |
C |
0.148 |
|
|
|
| 5 |
H |
0.128 |
|
|
|
| 6 |
O |
-0.403 |
|
|
|
| 7 |
C |
-0.076 |
|
|
|
| 8 |
H |
0.152 |
|
|
|
| 9 |
H |
0.148 |
|
|
|
| 10 |
C |
-0.501 |
|
|
|
| 11 |
H |
0.179 |
|
|
|
| 12 |
H |
0.178 |
|
|
|
| 13 |
H |
0.163 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.207 |
1.218 |
0.036 |
1.715 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.328 |
-0.111 |
0.154 |
| y |
-0.111 |
-30.226 |
-0.352 |
| z |
0.154 |
-0.352 |
-32.817 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.194 |
-0.111 |
0.154 |
| y |
-0.111 |
0.846 |
-0.352 |
| z |
0.154 |
-0.352 |
-3.039 |
|
| Polar |
|---|
| 3z2-r2 | -6.079 |
|---|
| x2-y2 | 0.899 |
|---|
| xy | -0.111 |
|---|
| xz | 0.154 |
|---|
| yz | -0.352 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.514 |
0.747 |
-0.033 |
| y |
0.747 |
6.620 |
-0.181 |
| z |
-0.033 |
-0.181 |
4.686 |
<r2> (average value of r
2) Å
2
| <r2> |
163.612 |
| (<r2>)1/2 |
12.791 |