Jump to
S1C2
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -230.362374 |
| Energy at 298.15K | -230.368185 |
| HF Energy | -229.760271 |
| Nuclear repulsion energy | 159.495219 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3401 |
3141 |
7.11 |
|
|
|
| 2 |
A1 |
3314 |
3061 |
7.56 |
|
|
|
| 3 |
A1 |
3293 |
3042 |
17.94 |
|
|
|
| 4 |
A1 |
1857 |
1715 |
4.38 |
|
|
|
| 5 |
A1 |
1522 |
1406 |
0.20 |
|
|
|
| 6 |
A1 |
1439 |
1330 |
3.77 |
|
|
|
| 7 |
A1 |
1282 |
1184 |
20.90 |
|
|
|
| 8 |
A1 |
915 |
845 |
3.38 |
|
|
|
| 9 |
A1 |
544 |
503 |
0.59 |
|
|
|
| 10 |
A1 |
243 |
225 |
0.01 |
|
|
|
| 11 |
A2 |
1024 |
946 |
0.00 |
|
|
|
| 12 |
A2 |
918 |
848 |
0.00 |
|
|
|
| 13 |
A2 |
753 |
695 |
0.00 |
|
|
|
| 14 |
A2 |
77 |
71 |
0.00 |
|
|
|
| 15 |
B1 |
1034 |
955 |
43.29 |
|
|
|
| 16 |
B1 |
920 |
850 |
124.10 |
|
|
|
| 17 |
B1 |
727 |
672 |
1.67 |
|
|
|
| 18 |
B1 |
59 |
55 |
2.47 |
|
|
|
| 19 |
B2 |
3401 |
3141 |
2.91 |
|
|
|
| 20 |
B2 |
3311 |
3058 |
0.07 |
|
|
|
| 21 |
B2 |
3285 |
3034 |
5.46 |
|
|
|
| 22 |
B2 |
1814 |
1676 |
422.07 |
|
|
|
| 23 |
B2 |
1512 |
1397 |
77.59 |
|
|
|
| 24 |
B2 |
1407 |
1299 |
4.52 |
|
|
|
| 25 |
B2 |
1295 |
1196 |
467.75 |
|
|
|
| 26 |
B2 |
1090 |
1007 |
7.30 |
|
|
|
| 27 |
B2 |
508 |
470 |
6.48 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20472.4 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 18910.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
0.328 |
| C2 |
0.000 |
1.169 |
-0.373 |
| C3 |
0.000 |
-1.169 |
-0.373 |
| C4 |
0.000 |
2.341 |
0.235 |
| C5 |
0.000 |
-2.341 |
0.235 |
| H6 |
0.000 |
1.065 |
-1.450 |
| H7 |
0.000 |
-1.065 |
-1.450 |
| H8 |
0.000 |
3.247 |
-0.347 |
| H9 |
0.000 |
2.419 |
1.310 |
| H10 |
0.000 |
-3.247 |
-0.347 |
| H11 |
0.000 |
-2.419 |
1.310 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3627 | 1.3627 | 2.3430 | 2.3430 | 2.0724 | 2.0724 | 3.3167 | 2.6108 | 3.3167 | 2.6108 |
C2 | 1.3627 | | 2.3377 | 1.3205 | 3.5623 | 1.0824 | 2.4799 | 2.0785 | 2.0965 | 4.4162 | 3.9629 | C3 | 1.3627 | 2.3377 | | 3.5623 | 1.3205 | 2.4799 | 1.0824 | 4.4162 | 3.9629 | 2.0785 | 2.0965 | C4 | 2.3430 | 1.3205 | 3.5623 | | 4.6824 | 2.1140 | 3.8000 | 1.0770 | 1.0782 | 5.6187 | 4.8800 | C5 | 2.3430 | 3.5623 | 1.3205 | 4.6824 | | 3.8000 | 2.1140 | 5.6187 | 4.8800 | 1.0770 | 1.0782 | H6 | 2.0724 | 1.0824 | 2.4799 | 2.1140 | 3.8000 | | 2.1295 | 2.4453 | 3.0747 | 4.4508 | 4.4448 | H7 | 2.0724 | 2.4799 | 1.0824 | 3.8000 | 2.1140 | 2.1295 | | 4.4508 | 4.4448 | 2.4453 | 3.0747 | H8 | 3.3167 | 2.0785 | 4.4162 | 1.0770 | 5.6187 | 2.4453 | 4.4508 | | 1.8531 | 6.4945 | 5.9036 | H9 | 2.6108 | 2.0965 | 3.9629 | 1.0782 | 4.8800 | 3.0747 | 4.4448 | 1.8531 | | 5.9036 | 4.8378 | H10 | 3.3167 | 4.4162 | 2.0785 | 5.6187 | 1.0770 | 4.4508 | 2.4453 | 6.4945 | 5.9036 | | 1.8531 | H11 | 2.6108 | 3.9629 | 2.0965 | 4.8800 | 1.0782 | 4.4448 | 3.0747 | 5.9036 | 4.8378 | 1.8531 | |
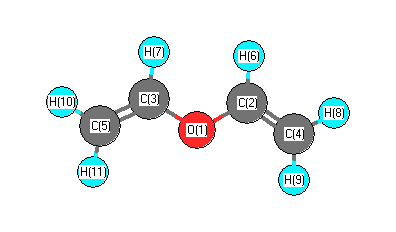 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
121.662 |
|
O1 |
C2 |
H6 |
115.418 |
| O1 |
C3 |
C5 |
121.662 |
|
O1 |
C3 |
H7 |
115.418 |
| C2 |
O1 |
C3 |
118.126 |
|
C2 |
C4 |
H8 |
119.871 |
| C2 |
C4 |
H9 |
121.534 |
|
C3 |
C5 |
H10 |
119.871 |
| C3 |
C5 |
H11 |
121.534 |
|
C4 |
C2 |
H6 |
122.920 |
| C5 |
C3 |
H7 |
122.920 |
|
H8 |
C4 |
H9 |
118.596 |
| H10 |
C5 |
H11 |
118.596 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -230.363491 |
| Energy at 298.15K | -230.369690 |
| HF Energy | -229.760958 |
| Nuclear repulsion energy | 162.294851 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3397 |
3138 |
8.81 |
|
|
|
| 2 |
A |
3396 |
3137 |
6.15 |
|
|
|
| 3 |
A |
3343 |
3088 |
11.37 |
|
|
|
| 4 |
A |
3313 |
3061 |
5.94 |
|
|
|
| 5 |
A |
3306 |
3054 |
1.81 |
|
|
|
| 6 |
A |
3293 |
3042 |
5.06 |
|
|
|
| 7 |
A |
1832 |
1692 |
94.84 |
|
|
|
| 8 |
A |
1811 |
1673 |
160.42 |
|
|
|
| 9 |
A |
1526 |
1410 |
1.84 |
|
|
|
| 10 |
A |
1508 |
1393 |
28.05 |
|
|
|
| 11 |
A |
1439 |
1330 |
49.01 |
|
|
|
| 12 |
A |
1405 |
1298 |
3.17 |
|
|
|
| 13 |
A |
1327 |
1226 |
293.24 |
|
|
|
| 14 |
A |
1203 |
1112 |
16.32 |
|
|
|
| 15 |
A |
1072 |
990 |
40.71 |
|
|
|
| 16 |
A |
1062 |
981 |
3.95 |
|
|
|
| 17 |
A |
1049 |
969 |
33.71 |
|
|
|
| 18 |
A |
948 |
876 |
59.90 |
|
|
|
| 19 |
A |
934 |
863 |
15.23 |
|
|
|
| 20 |
A |
926 |
855 |
60.80 |
|
|
|
| 21 |
A |
780 |
721 |
4.36 |
|
|
|
| 22 |
A |
760 |
702 |
0.82 |
|
|
|
| 23 |
A |
612 |
565 |
2.84 |
|
|
|
| 24 |
A |
478 |
441 |
3.61 |
|
|
|
| 25 |
A |
300 |
277 |
1.96 |
|
|
|
| 26 |
A |
197 |
182 |
9.85 |
|
|
|
| 27 |
A |
93 |
86 |
0.80 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20655.8 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 19079.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-0.039 |
-0.809 |
0.052 |
| C2 |
1.282 |
-0.483 |
-0.009 |
| C3 |
-0.929 |
0.191 |
0.338 |
| C4 |
1.803 |
0.727 |
-0.149 |
| C5 |
-2.124 |
0.232 |
-0.220 |
| H6 |
1.886 |
-1.374 |
0.059 |
| H7 |
-0.591 |
0.894 |
1.088 |
| H8 |
2.874 |
0.835 |
-0.175 |
| H9 |
1.208 |
1.619 |
-0.259 |
| H10 |
-2.835 |
0.985 |
0.079 |
| H11 |
-2.420 |
-0.487 |
-0.968 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3621 | 1.3690 | 2.4066 | 2.3461 | 2.0063 | 2.0686 | 3.3532 | 2.7470 | 3.3217 | 2.6101 |
C2 | 1.3621 | | 2.3372 | 1.3241 | 3.4865 | 1.0790 | 2.5708 | 2.0737 | 2.1175 | 4.3712 | 3.8239 | C3 | 1.3690 | 2.3372 | | 2.8258 | 1.3197 | 3.2329 | 1.0819 | 3.8913 | 2.6380 | 2.0805 | 2.0949 | C4 | 2.4066 | 1.3241 | 2.8258 | | 3.9586 | 2.1127 | 2.6998 | 1.0774 | 1.0778 | 4.6501 | 4.4693 | C5 | 2.3461 | 3.4865 | 1.3197 | 3.9586 | | 4.3286 | 2.1213 | 5.0350 | 3.6094 | 1.0777 | 1.0784 | H6 | 2.0063 | 1.0790 | 3.2329 | 2.1127 | 4.3286 | | 3.5129 | 2.4316 | 3.0851 | 5.2772 | 4.5148 | H7 | 2.0686 | 2.5708 | 1.0819 | 2.6998 | 2.1213 | 3.5129 | | 3.6891 | 2.3613 | 2.4617 | 3.0787 | H8 | 3.3532 | 2.0737 | 3.8913 | 1.0774 | 5.0350 | 2.4316 | 3.6891 | | 1.8435 | 5.7166 | 5.5142 | H9 | 2.7470 | 2.1175 | 2.6380 | 1.0778 | 3.6094 | 3.0851 | 2.3613 | 1.8435 | | 4.1058 | 4.2540 | H10 | 3.3217 | 4.3712 | 2.0805 | 4.6501 | 1.0777 | 5.2772 | 2.4617 | 5.7166 | 4.1058 | | 1.8529 | H11 | 2.6101 | 3.8239 | 2.0949 | 4.4693 | 1.0784 | 4.5148 | 3.0787 | 5.5142 | 4.2540 | 1.8529 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
127.235 |
|
O1 |
C2 |
H6 |
110.006 |
| O1 |
C3 |
C5 |
121.508 |
|
O1 |
C3 |
H7 |
114.622 |
| C2 |
O1 |
C3 |
117.685 |
|
C2 |
C4 |
H8 |
119.059 |
| C2 |
C4 |
H9 |
123.333 |
|
C3 |
C5 |
H10 |
120.080 |
| C3 |
C5 |
H11 |
121.424 |
|
C4 |
C2 |
H6 |
122.754 |
| C5 |
C3 |
H7 |
123.778 |
|
H8 |
C4 |
H9 |
117.595 |
| H10 |
C5 |
H11 |
118.491 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
