Jump to
S1C2
Energy calculated at PBE1PBE/6-31G*
| | hartrees |
|---|
| Energy at 0K | -230.918233 |
| Energy at 298.15K | -230.924007 |
| HF Energy | -230.918233 |
| Nuclear repulsion energy | 159.123893 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3316 |
3151 |
5.10 |
|
|
|
| 2 |
A1 |
3223 |
3063 |
0.86 |
|
|
|
| 3 |
A1 |
3187 |
3029 |
30.52 |
|
|
|
| 4 |
A1 |
1784 |
1695 |
4.73 |
|
|
|
| 5 |
A1 |
1454 |
1382 |
0.19 |
|
|
|
| 6 |
A1 |
1377 |
1308 |
5.60 |
|
|
|
| 7 |
A1 |
1235 |
1173 |
22.31 |
|
|
|
| 8 |
A1 |
879 |
835 |
1.86 |
|
|
|
| 9 |
A1 |
530 |
503 |
0.42 |
|
|
|
| 10 |
A1 |
233 |
222 |
0.00 |
|
|
|
| 11 |
A2 |
979 |
930 |
0.00 |
|
|
|
| 12 |
A2 |
853 |
811 |
0.00 |
|
|
|
| 13 |
A2 |
724 |
688 |
0.00 |
|
|
|
| 14 |
A2 |
115 |
109 |
0.00 |
|
|
|
| 15 |
B1 |
989 |
940 |
40.43 |
|
|
|
| 16 |
B1 |
857 |
814 |
122.89 |
|
|
|
| 17 |
B1 |
693 |
659 |
5.18 |
|
|
|
| 18 |
B1 |
84 |
80 |
1.54 |
|
|
|
| 19 |
B2 |
3316 |
3151 |
1.49 |
|
|
|
| 20 |
B2 |
3222 |
3062 |
0.53 |
|
|
|
| 21 |
B2 |
3182 |
3024 |
5.30 |
|
|
|
| 22 |
B2 |
1737 |
1651 |
456.93 |
|
|
|
| 23 |
B2 |
1447 |
1375 |
75.73 |
|
|
|
| 24 |
B2 |
1350 |
1282 |
13.91 |
|
|
|
| 25 |
B2 |
1264 |
1202 |
404.15 |
|
|
|
| 26 |
B2 |
1041 |
990 |
2.27 |
|
|
|
| 27 |
B2 |
491 |
466 |
6.76 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19780.5 cm
-1
Scaled (by 0.9503) Zero Point Vibrational Energy (zpe) 18797.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
0.328 |
| C2 |
0.000 |
1.167 |
-0.371 |
| C3 |
0.000 |
-1.167 |
-0.371 |
| C4 |
0.000 |
2.349 |
0.235 |
| C5 |
0.000 |
-2.349 |
0.235 |
| H6 |
0.000 |
1.057 |
-1.456 |
| H7 |
0.000 |
-1.057 |
-1.456 |
| H8 |
0.000 |
3.255 |
-0.357 |
| H9 |
0.000 |
2.431 |
1.317 |
| H10 |
0.000 |
-3.255 |
-0.357 |
| H11 |
0.000 |
-2.431 |
1.317 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3597 | 1.3597 | 2.3504 | 2.3504 | 2.0727 | 2.0727 | 3.3267 | 2.6244 | 3.3267 | 2.6244 |
C2 | 1.3597 | | 2.3331 | 1.3283 | 3.5670 | 1.0901 | 2.4736 | 2.0890 | 2.1087 | 4.4221 | 3.9737 | C3 | 1.3597 | 2.3331 | | 3.5670 | 1.3283 | 2.4736 | 1.0901 | 4.4221 | 3.9737 | 2.0890 | 2.1087 | C4 | 2.3504 | 1.3283 | 3.5670 | | 4.6971 | 2.1278 | 3.8017 | 1.0830 | 1.0846 | 5.6352 | 4.9004 | C5 | 2.3504 | 3.5670 | 1.3283 | 4.6971 | | 3.8017 | 2.1278 | 5.6352 | 4.9004 | 1.0830 | 1.0846 | H6 | 2.0727 | 1.0901 | 2.4736 | 2.1278 | 3.8017 | | 2.1131 | 2.4582 | 3.0941 | 4.4498 | 4.4550 | H7 | 2.0727 | 2.4736 | 1.0901 | 3.8017 | 2.1278 | 2.1131 | | 4.4498 | 4.4550 | 2.4582 | 3.0941 | H8 | 3.3267 | 2.0890 | 4.4221 | 1.0830 | 5.6352 | 2.4582 | 4.4498 | | 1.8655 | 6.5110 | 5.9276 | H9 | 2.6244 | 2.1087 | 3.9737 | 1.0846 | 4.9004 | 3.0941 | 4.4550 | 1.8655 | | 5.9276 | 4.8620 | H10 | 3.3267 | 4.4221 | 2.0890 | 5.6352 | 1.0830 | 4.4498 | 2.4582 | 6.5110 | 5.9276 | | 1.8655 | H11 | 2.6244 | 3.9737 | 2.1087 | 4.9004 | 1.0846 | 4.4550 | 3.0941 | 5.9276 | 4.8620 | 1.8655 | |
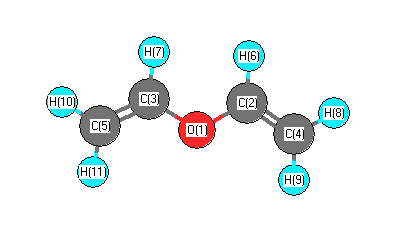 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
121.937 |
|
O1 |
C2 |
H6 |
115.125 |
| O1 |
C3 |
C5 |
121.937 |
|
O1 |
C3 |
H7 |
115.125 |
| C2 |
O1 |
C3 |
118.168 |
|
C2 |
C4 |
H8 |
119.721 |
| C2 |
C4 |
H9 |
121.504 |
|
C3 |
C5 |
H10 |
119.721 |
| C3 |
C5 |
H11 |
121.504 |
|
C4 |
C2 |
H6 |
122.938 |
| C5 |
C3 |
H7 |
122.938 |
|
H8 |
C4 |
H9 |
118.774 |
| H10 |
C5 |
H11 |
118.774 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.437 |
|
|
|
| 2 |
C |
0.152 |
|
|
|
| 3 |
C |
0.152 |
|
|
|
| 4 |
C |
-0.464 |
|
|
|
| 5 |
C |
-0.464 |
|
|
|
| 6 |
H |
0.167 |
|
|
|
| 7 |
H |
0.167 |
|
|
|
| 8 |
H |
0.177 |
|
|
|
| 9 |
H |
0.185 |
|
|
|
| 10 |
H |
0.177 |
|
|
|
| 11 |
H |
0.185 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.334 |
1.334 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.331 |
0.000 |
0.000 |
| y |
0.000 |
-27.023 |
0.000 |
| z |
0.000 |
0.000 |
-27.045 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.297 |
0.000 |
0.000 |
| y |
0.000 |
3.165 |
0.000 |
| z |
0.000 |
0.000 |
3.132 |
|
| Polar |
|---|
| 3z2-r2 | 6.264 |
|---|
| x2-y2 | -6.308 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.712 |
0.000 |
0.000 |
| y |
0.000 |
12.216 |
0.000 |
| z |
0.000 |
0.000 |
6.147 |
<r2> (average value of r
2) Å
2
| <r2> |
147.096 |
| (<r2>)1/2 |
12.128 |
Jump to
S1C1
Energy calculated at PBE1PBE/6-31G*
| | hartrees |
|---|
| Energy at 0K | -230.918605 |
| Energy at 298.15K | -230.924676 |
| HF Energy | -230.918605 |
| Nuclear repulsion energy | 161.785393 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3312 |
3147 |
4.22 |
|
|
|
| 2 |
A |
3309 |
3144 |
6.63 |
|
|
|
| 3 |
A |
3246 |
3085 |
10.67 |
|
|
|
| 4 |
A |
3220 |
3060 |
1.62 |
|
|
|
| 5 |
A |
3218 |
3058 |
3.08 |
|
|
|
| 6 |
A |
3195 |
3036 |
10.17 |
|
|
|
| 7 |
A |
1756 |
1669 |
95.76 |
|
|
|
| 8 |
A |
1735 |
1648 |
184.39 |
|
|
|
| 9 |
A |
1459 |
1386 |
1.63 |
|
|
|
| 10 |
A |
1441 |
1369 |
30.75 |
|
|
|
| 11 |
A |
1384 |
1315 |
67.98 |
|
|
|
| 12 |
A |
1343 |
1276 |
5.08 |
|
|
|
| 13 |
A |
1280 |
1216 |
261.90 |
|
|
|
| 14 |
A |
1161 |
1103 |
16.41 |
|
|
|
| 15 |
A |
1030 |
979 |
28.25 |
|
|
|
| 16 |
A |
1013 |
962 |
5.88 |
|
|
|
| 17 |
A |
998 |
948 |
32.31 |
|
|
|
| 18 |
A |
899 |
854 |
19.09 |
|
|
|
| 19 |
A |
878 |
835 |
48.82 |
|
|
|
| 20 |
A |
859 |
816 |
61.56 |
|
|
|
| 21 |
A |
743 |
706 |
4.45 |
|
|
|
| 22 |
A |
727 |
691 |
1.58 |
|
|
|
| 23 |
A |
595 |
565 |
2.41 |
|
|
|
| 24 |
A |
458 |
435 |
2.46 |
|
|
|
| 25 |
A |
289 |
274 |
1.67 |
|
|
|
| 26 |
A |
212 |
201 |
9.21 |
|
|
|
| 27 |
A |
96 |
91 |
0.32 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19926.4 cm
-1
Scaled (by 0.9503) Zero Point Vibrational Energy (zpe) 18936.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-0.045 |
-0.793 |
0.046 |
| C2 |
1.281 |
-0.493 |
0.005 |
| C3 |
-0.923 |
0.221 |
0.301 |
| C4 |
1.834 |
0.708 |
-0.149 |
| C5 |
-2.155 |
0.217 |
-0.194 |
| H6 |
1.867 |
-1.402 |
0.106 |
| H7 |
-0.545 |
0.986 |
0.978 |
| H8 |
2.914 |
0.795 |
-0.150 |
| H9 |
1.254 |
1.613 |
-0.299 |
| H10 |
-2.855 |
0.994 |
0.091 |
| H11 |
-2.490 |
-0.560 |
-0.873 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3601 | 1.3655 | 2.4131 | 2.3518 | 2.0076 | 2.0696 | 3.3640 | 2.7555 | 3.3300 | 2.6222 |
C2 | 1.3601 | | 2.3359 | 1.3317 | 3.5145 | 1.0858 | 2.5434 | 2.0858 | 2.1278 | 4.3956 | 3.8723 | C3 | 1.3655 | 2.3359 | | 2.8358 | 1.3275 | 3.2339 | 1.0894 | 3.9061 | 2.6523 | 2.0906 | 2.1075 | C4 | 2.4131 | 1.3317 | 2.8358 | | 4.0195 | 2.1257 | 2.6469 | 1.0836 | 1.0846 | 4.7031 | 4.5639 | C5 | 2.3518 | 3.5145 | 1.3275 | 4.0195 | | 4.3462 | 2.1346 | 5.1022 | 3.6849 | 1.0836 | 1.0849 | H6 | 2.0076 | 1.0858 | 3.2339 | 2.1257 | 4.3462 | | 3.5045 | 2.4468 | 3.1024 | 5.2947 | 4.5441 | H7 | 2.0696 | 2.5434 | 1.0894 | 2.6469 | 2.1346 | 3.5045 | | 3.6433 | 2.2930 | 2.4735 | 3.0980 | H8 | 3.3640 | 2.0858 | 3.9061 | 1.0836 | 5.1022 | 2.4468 | 3.6433 | | 1.8566 | 5.7769 | 5.6180 | H9 | 2.7555 | 2.1278 | 2.6523 | 1.0846 | 3.6849 | 3.1024 | 2.2930 | 1.8566 | | 4.1727 | 4.3663 | H10 | 3.3300 | 4.3956 | 2.0906 | 4.7031 | 1.0836 | 5.2947 | 2.4735 | 5.7769 | 4.1727 | | 1.8651 | H11 | 2.6222 | 3.8723 | 2.1075 | 4.5639 | 1.0849 | 4.5441 | 3.0980 | 5.6180 | 4.3663 | 1.8651 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
127.384 |
|
O1 |
C2 |
H6 |
109.822 |
| O1 |
C3 |
C5 |
121.681 |
|
O1 |
C3 |
H7 |
114.466 |
| C2 |
O1 |
C3 |
117.965 |
|
C2 |
C4 |
H8 |
119.082 |
| C2 |
C4 |
H9 |
123.096 |
|
C3 |
C5 |
H10 |
119.902 |
| C3 |
C5 |
H11 |
121.436 |
|
C4 |
C2 |
H6 |
122.789 |
| C5 |
C3 |
H7 |
123.767 |
|
H8 |
C4 |
H9 |
117.806 |
| H10 |
C5 |
H11 |
118.656 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.443 |
|
|
|
| 2 |
C |
0.126 |
|
|
|
| 3 |
C |
0.129 |
|
|
|
| 4 |
C |
-0.457 |
|
|
|
| 5 |
C |
-0.441 |
|
|
|
| 6 |
H |
0.182 |
|
|
|
| 7 |
H |
0.178 |
|
|
|
| 8 |
H |
0.180 |
|
|
|
| 9 |
H |
0.182 |
|
|
|
| 10 |
H |
0.178 |
|
|
|
| 11 |
H |
0.186 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.358 |
0.670 |
0.283 |
0.810 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.582 |
-1.677 |
0.567 |
| y |
-1.677 |
-28.961 |
0.970 |
| z |
0.567 |
0.970 |
-31.900 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.849 |
-1.677 |
0.567 |
| y |
-1.677 |
0.280 |
0.970 |
| z |
0.567 |
0.970 |
-4.129 |
|
| Polar |
|---|
| 3z2-r2 | -8.258 |
|---|
| x2-y2 | 2.379 |
|---|
| xy | -1.677 |
|---|
| xz | 0.567 |
|---|
| yz | 0.970 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.212 |
0.315 |
0.624 |
| y |
0.315 |
6.187 |
0.275 |
| z |
0.624 |
0.275 |
3.558 |
<r2> (average value of r
2) Å
2
| <r2> |
129.742 |
| (<r2>)1/2 |
11.390 |
