return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH2CHOCHCH2 (Vinyl ether)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-230.934952
Energy at 298.15K-230.940670
HF Energy-230.707673
Nuclear repulsion energy158.541268
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3320 3152 6.89      
2 A1 3229 3065 1.76      
3 A1 3201 3039 28.99      
4 A1 1763 1674 4.16      
5 A1 1470 1396 0.09      
6 A1 1386 1315 4.48      
7 A1 1226 1164 19.41      
8 A1 875 831 3.15      
9 A1 526 499 0.49      
10 A1 234 222 0.00      
11 A2 974 924 0.00      
12 A2 841 798 0.00      
13 A2 722 685 0.00      
14 A2 102 97 0.00      
15 B1 983 933 44.62      
16 B1 844 801 104.31      
17 B1 689 654 3.05      
18 B1 74 71 1.70      
19 B2 3320 3152 2.16      
20 B2 3228 3064 0.03      
21 B2 3194 3032 5.45      
22 B2 1718 1630 398.34      
23 B2 1459 1385 70.31      
24 B2 1357 1288 8.96      
25 B2 1239 1176 423.81      
26 B2 1048 995 7.48      
27 B2 491 466 7.19      

Unscaled Zero Point Vibrational Energy (zpe) 19755.8 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 18752.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
1.16358 0.08343 0.07784

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 0.000 0.000 0.332
C2 0.000 1.174 -0.375
C3 0.000 -1.174 -0.375
C4 0.000 2.357 0.236
C5 0.000 -2.357 0.236
H6 0.000 1.063 -1.457
H7 0.000 -1.063 -1.457
H8 0.000 3.264 -0.351
H9 0.000 2.434 1.316
H10 0.000 -3.264 -0.351
H11 0.000 -2.434 1.316

Atom - Atom Distances (Å)
  O1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11
O11.37051.37052.35862.35862.08092.08093.33502.62553.33502.6255
C21.37052.34811.33143.58331.08762.48502.09042.10914.43843.9848
C31.37052.34813.58331.33142.48501.08764.43843.98482.09042.1091
C42.35861.33143.58334.71332.13113.81611.08131.08265.65154.9108
C52.35863.58331.33144.71333.81612.13115.65154.91081.08131.0826
H62.08091.08762.48502.13113.81612.12612.46343.09374.46644.4633
H72.08092.48501.08763.81612.13112.12614.46644.46332.46343.0937
H83.33502.09044.43841.08135.65152.46344.46641.86286.52865.9372
H92.62552.10913.98481.08264.91083.09374.46331.86285.93724.8679
H103.33504.43842.09045.65151.08134.46642.46346.52865.93721.8628
H112.62553.98482.10914.91081.08264.46333.09375.93724.86791.8628

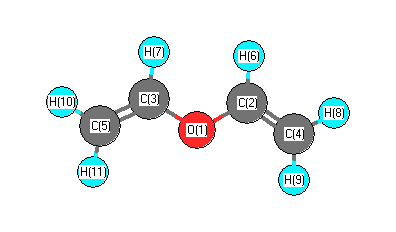
picture of Vinyl ether state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C2 C4 121.596 O1 C2 H6 115.195
O1 C3 C5 121.596 O1 C3 H7 115.195
C2 O1 C3 117.891 C2 C4 H8 119.731
C2 C4 H9 121.445 C3 C5 H10 119.731
C3 C5 H11 121.445 C4 C2 H6 123.208
C5 C3 H7 123.208 H8 C4 H9 118.824
H10 C5 H11 118.824
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 O -0.507      
2 C 0.188      
3 C 0.188      
4 C -0.430      
5 C -0.430      
6 H 0.159      
7 H 0.159      
8 H 0.164      
9 H 0.174      
10 H 0.164      
11 H 0.174      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 2.656 0.000 0.000
y 0.000 12.007 0.000
z 0.000 0.000 6.124


<r2> (average value of r2) Å2
<r2> 148.102
(<r2>)1/2 12.170