Jump to
S1C2
Energy calculated at HSEh1PBE/6-31G*
| | hartrees |
|---|
| Energy at 0K | -230.938077 |
| Energy at 298.15K | -230.943853 |
| Nuclear repulsion energy | 159.113719 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3314 |
3152 |
5.74 |
|
|
|
| 2 |
A1 |
3221 |
3063 |
0.81 |
|
|
|
| 3 |
A1 |
3186 |
3030 |
32.15 |
|
|
|
| 4 |
A1 |
1782 |
1695 |
4.75 |
|
|
|
| 5 |
A1 |
1454 |
1383 |
0.17 |
|
|
|
| 6 |
A1 |
1376 |
1309 |
5.59 |
|
|
|
| 7 |
A1 |
1233 |
1173 |
22.33 |
|
|
|
| 8 |
A1 |
878 |
835 |
1.97 |
|
|
|
| 9 |
A1 |
530 |
504 |
0.42 |
|
|
|
| 10 |
A1 |
233 |
222 |
0.00 |
|
|
|
| 11 |
A2 |
979 |
931 |
0.00 |
|
|
|
| 12 |
A2 |
853 |
811 |
0.00 |
|
|
|
| 13 |
A2 |
724 |
689 |
0.00 |
|
|
|
| 14 |
A2 |
117 |
111 |
0.00 |
|
|
|
| 15 |
B1 |
989 |
941 |
40.11 |
|
|
|
| 16 |
B1 |
856 |
814 |
120.92 |
|
|
|
| 17 |
B1 |
693 |
659 |
5.14 |
|
|
|
| 18 |
B1 |
85 |
81 |
1.53 |
|
|
|
| 19 |
B2 |
3314 |
3152 |
1.66 |
|
|
|
| 20 |
B2 |
3221 |
3063 |
0.40 |
|
|
|
| 21 |
B2 |
3181 |
3025 |
5.12 |
|
|
|
| 22 |
B2 |
1735 |
1650 |
453.33 |
|
|
|
| 23 |
B2 |
1447 |
1376 |
74.34 |
|
|
|
| 24 |
B2 |
1349 |
1283 |
14.10 |
|
|
|
| 25 |
B2 |
1262 |
1200 |
405.17 |
|
|
|
| 26 |
B2 |
1041 |
990 |
2.47 |
|
|
|
| 27 |
B2 |
490 |
466 |
6.85 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19770.3 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 18801.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
0.328 |
| C2 |
0.000 |
1.167 |
-0.371 |
| C3 |
0.000 |
-1.167 |
-0.371 |
| C4 |
0.000 |
2.349 |
0.235 |
| C5 |
0.000 |
-2.349 |
0.235 |
| H6 |
0.000 |
1.057 |
-1.455 |
| H7 |
0.000 |
-1.057 |
-1.455 |
| H8 |
0.000 |
3.256 |
-0.356 |
| H9 |
0.000 |
2.431 |
1.316 |
| H10 |
0.000 |
-3.256 |
-0.356 |
| H11 |
0.000 |
-2.431 |
1.316 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3601 | 1.3601 | 2.3506 | 2.3506 | 2.0725 | 2.0725 | 3.3268 | 2.6246 | 3.3268 | 2.6246 |
C2 | 1.3601 | | 2.3340 | 1.3282 | 3.5677 | 1.0900 | 2.4740 | 2.0888 | 2.1084 | 4.4228 | 3.9741 | C3 | 1.3601 | 2.3340 | | 3.5677 | 1.3282 | 2.4740 | 1.0900 | 4.4228 | 3.9741 | 2.0888 | 2.1084 | C4 | 2.3506 | 1.3282 | 3.5677 | | 4.6976 | 2.1278 | 3.8020 | 1.0828 | 1.0844 | 5.6357 | 4.9007 | C5 | 2.3506 | 3.5677 | 1.3282 | 4.6976 | | 3.8020 | 2.1278 | 5.6357 | 4.9007 | 1.0828 | 1.0844 | H6 | 2.0725 | 1.0900 | 2.4740 | 2.1278 | 3.8020 | | 2.1133 | 2.4585 | 3.0939 | 4.4503 | 4.4550 | H7 | 2.0725 | 2.4740 | 1.0900 | 3.8020 | 2.1278 | 2.1133 | | 4.4503 | 4.4550 | 2.4585 | 3.0939 | H8 | 3.3268 | 2.0888 | 4.4228 | 1.0828 | 5.6357 | 2.4585 | 4.4503 | | 1.8650 | 6.5116 | 5.9278 | H9 | 2.6246 | 2.1084 | 3.9741 | 1.0844 | 4.9007 | 3.0939 | 4.4550 | 1.8650 | | 5.9278 | 4.8623 | H10 | 3.3268 | 4.4228 | 2.0888 | 5.6357 | 1.0828 | 4.4503 | 2.4585 | 6.5116 | 5.9278 | | 1.8650 | H11 | 2.6246 | 3.9741 | 2.1084 | 4.9007 | 1.0844 | 4.4550 | 3.0939 | 5.9278 | 4.8623 | 1.8650 | |
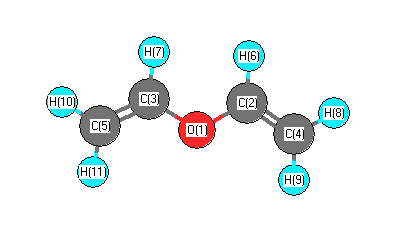 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
121.948 |
|
O1 |
C2 |
H6 |
115.091 |
| O1 |
C3 |
C5 |
121.948 |
|
O1 |
C3 |
H7 |
115.091 |
| C2 |
O1 |
C3 |
118.197 |
|
C2 |
C4 |
H8 |
119.738 |
| C2 |
C4 |
H9 |
121.504 |
|
C3 |
C5 |
H10 |
119.738 |
| C3 |
C5 |
H11 |
121.504 |
|
C4 |
C2 |
H6 |
122.961 |
| C5 |
C3 |
H7 |
122.961 |
|
H8 |
C4 |
H9 |
118.758 |
| H10 |
C5 |
H11 |
118.758 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.435 |
|
|
|
| 2 |
C |
0.154 |
|
|
|
| 3 |
C |
0.154 |
|
|
|
| 4 |
C |
-0.458 |
|
|
|
| 5 |
C |
-0.458 |
|
|
|
| 6 |
H |
0.164 |
|
|
|
| 7 |
H |
0.164 |
|
|
|
| 8 |
H |
0.175 |
|
|
|
| 9 |
H |
0.183 |
|
|
|
| 10 |
H |
0.175 |
|
|
|
| 11 |
H |
0.183 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.330 |
1.330 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.304 |
0.000 |
0.000 |
| y |
0.000 |
-27.038 |
0.000 |
| z |
0.000 |
0.000 |
-27.071 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.249 |
0.000 |
0.000 |
| y |
0.000 |
3.150 |
0.000 |
| z |
0.000 |
0.000 |
3.099 |
|
| Polar |
|---|
| 3z2-r2 | 6.199 |
|---|
| x2-y2 | -6.266 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.709 |
0.000 |
0.000 |
| y |
0.000 |
12.247 |
0.000 |
| z |
0.000 |
0.000 |
6.159 |
<r2> (average value of r
2) Å
2
| <r2> |
147.128 |
| (<r2>)1/2 |
12.130 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G*
| | hartrees |
|---|
| Energy at 0K | -230.938446 |
| Energy at 298.15K | -230.944517 |
| HF Energy | -230.938446 |
| Nuclear repulsion energy | 161.780199 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3311 |
3149 |
4.65 |
|
|
|
| 2 |
A |
3308 |
3146 |
7.25 |
|
|
|
| 3 |
A |
3246 |
3087 |
11.18 |
|
|
|
| 4 |
A |
3219 |
3062 |
1.53 |
|
|
|
| 5 |
A |
3217 |
3060 |
2.94 |
|
|
|
| 6 |
A |
3194 |
3037 |
10.72 |
|
|
|
| 7 |
A |
1754 |
1668 |
95.99 |
|
|
|
| 8 |
A |
1733 |
1648 |
181.58 |
|
|
|
| 9 |
A |
1458 |
1387 |
1.39 |
|
|
|
| 10 |
A |
1441 |
1370 |
29.44 |
|
|
|
| 11 |
A |
1384 |
1316 |
66.47 |
|
|
|
| 12 |
A |
1343 |
1277 |
5.18 |
|
|
|
| 13 |
A |
1277 |
1215 |
264.53 |
|
|
|
| 14 |
A |
1159 |
1103 |
16.32 |
|
|
|
| 15 |
A |
1029 |
979 |
29.37 |
|
|
|
| 16 |
A |
1012 |
963 |
5.36 |
|
|
|
| 17 |
A |
997 |
949 |
32.13 |
|
|
|
| 18 |
A |
898 |
854 |
19.37 |
|
|
|
| 19 |
A |
878 |
835 |
47.97 |
|
|
|
| 20 |
A |
858 |
816 |
60.59 |
|
|
|
| 21 |
A |
742 |
706 |
4.39 |
|
|
|
| 22 |
A |
727 |
691 |
1.55 |
|
|
|
| 23 |
A |
595 |
566 |
2.41 |
|
|
|
| 24 |
A |
457 |
435 |
2.46 |
|
|
|
| 25 |
A |
289 |
275 |
1.64 |
|
|
|
| 26 |
A |
213 |
202 |
9.20 |
|
|
|
| 27 |
A |
96 |
91 |
0.30 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19917.8 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 18941.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-0.045 |
-0.793 |
0.046 |
| C2 |
1.281 |
-0.494 |
0.006 |
| C3 |
-0.923 |
0.222 |
0.300 |
| C4 |
1.834 |
0.708 |
-0.149 |
| C5 |
-2.156 |
0.217 |
-0.193 |
| H6 |
1.867 |
-1.402 |
0.107 |
| H7 |
-0.544 |
0.988 |
0.974 |
| H8 |
2.914 |
0.796 |
-0.149 |
| H9 |
1.253 |
1.612 |
-0.299 |
| H10 |
-2.855 |
0.994 |
0.091 |
| H11 |
-2.491 |
-0.562 |
-0.870 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3605 | 1.3658 | 2.4129 | 2.3520 | 2.0079 | 2.0696 | 3.3641 | 2.7543 | 3.3302 | 2.6222 |
C2 | 1.3605 | | 2.3366 | 1.3316 | 3.5155 | 1.0855 | 2.5428 | 2.0858 | 2.1272 | 4.3965 | 3.8737 | C3 | 1.3658 | 2.3366 | | 2.8355 | 1.3273 | 3.2344 | 1.0892 | 3.9057 | 2.6509 | 2.0905 | 2.1072 | C4 | 2.4129 | 1.3316 | 2.8355 | | 4.0201 | 2.1256 | 2.6445 | 1.0834 | 1.0845 | 4.7035 | 4.5653 | C5 | 2.3520 | 3.5155 | 1.3273 | 4.0201 | | 4.3469 | 2.1346 | 5.1027 | 3.6848 | 1.0834 | 1.0847 | H6 | 2.0079 | 1.0855 | 3.2344 | 2.1256 | 4.3469 | | 3.5042 | 2.4474 | 3.1020 | 5.2952 | 4.5450 | H7 | 2.0696 | 2.5428 | 1.0892 | 2.6445 | 2.1346 | 3.5042 | | 3.6406 | 2.2891 | 2.4738 | 3.0977 | H8 | 3.3641 | 2.0858 | 3.9057 | 1.0834 | 5.1027 | 2.4474 | 3.6406 | | 1.8562 | 5.7769 | 5.6196 | H9 | 2.7543 | 2.1272 | 2.6509 | 1.0845 | 3.6848 | 3.1020 | 2.2891 | 1.8562 | | 4.1723 | 4.3671 | H10 | 3.3302 | 4.3965 | 2.0905 | 4.7035 | 1.0834 | 5.2952 | 2.4738 | 5.7769 | 4.1723 | | 1.8646 | H11 | 2.6222 | 3.8737 | 2.1072 | 4.5653 | 1.0847 | 4.5450 | 3.0977 | 5.6196 | 4.3671 | 1.8646 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
127.344 |
|
O1 |
C2 |
H6 |
109.834 |
| O1 |
C3 |
C5 |
121.686 |
|
O1 |
C3 |
H7 |
114.441 |
| C2 |
O1 |
C3 |
117.969 |
|
C2 |
C4 |
H8 |
119.118 |
| C2 |
C4 |
H9 |
123.070 |
|
C3 |
C5 |
H10 |
119.916 |
| C3 |
C5 |
H11 |
121.437 |
|
C4 |
C2 |
H6 |
122.818 |
| C5 |
C3 |
H7 |
123.789 |
|
H8 |
C4 |
H9 |
117.796 |
| H10 |
C5 |
H11 |
118.641 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.441 |
|
|
|
| 2 |
C |
0.128 |
|
|
|
| 3 |
C |
0.131 |
|
|
|
| 4 |
C |
-0.451 |
|
|
|
| 5 |
C |
-0.436 |
|
|
|
| 6 |
H |
0.179 |
|
|
|
| 7 |
H |
0.175 |
|
|
|
| 8 |
H |
0.177 |
|
|
|
| 9 |
H |
0.179 |
|
|
|
| 10 |
H |
0.176 |
|
|
|
| 11 |
H |
0.183 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.358 |
0.668 |
0.281 |
0.809 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.591 |
-1.668 |
0.564 |
| y |
-1.668 |
-28.970 |
0.959 |
| z |
0.564 |
0.959 |
-31.893 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.841 |
-1.668 |
0.564 |
| y |
-1.668 |
0.272 |
0.959 |
| z |
0.564 |
0.959 |
-4.113 |
|
| Polar |
|---|
| 3z2-r2 | -8.226 |
|---|
| x2-y2 | 2.379 |
|---|
| xy | -1.668 |
|---|
| xz | 0.564 |
|---|
| yz | 0.959 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.239 |
0.314 |
0.622 |
| y |
0.314 |
6.195 |
0.277 |
| z |
0.622 |
0.277 |
3.553 |
<r2> (average value of r
2) Å
2
| <r2> |
129.761 |
| (<r2>)1/2 |
11.391 |
