return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H4S (Thiophene)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-551.845582
Energy at 298.15K-551.850109
HF Energy-551.290035
Nuclear repulsion energy203.178769
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3380 3122 2.31      
2 A1 3343 3088 4.05      
3 A1 1551 1433 20.85      
4 A1 1489 1376 0.05      
5 A1 1175 1085 5.32      
6 A1 1102 1018 0.91      
7 A1 889 821 32.39      
8 A1 645 596 0.53      
9 A2 947 875 0.00      
10 A2 744 687 0.00      
11 A2 585 540 0.00      
12 B1 941 870 1.39      
13 B1 770 711 127.93      
14 B1 470 434 0.37      
15 B2 3377 3120 0.74      
16 B2 3329 3075 5.05      
17 B2 1678 1550 0.22      
18 B2 1360 1257 9.91      
19 B2 1177 1087 3.50      
20 B2 934 863 0.84      
21 B2 797 736 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 15341.9 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 14171.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
0.26959 0.18134 0.10841

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 0.000 0.000 1.190
C2 0.000 1.231 -0.010
C3 0.000 -1.231 -0.010
C4 0.000 0.715 -1.263
C5 0.000 -0.715 -1.263
H6 0.000 2.269 0.277
H7 0.000 -2.269 0.277
H8 0.000 1.318 -2.158
H9 0.000 -1.318 -2.158

Atom - Atom Distances (Å)
  S1 C2 C3 C4 C5 H6 H7 H8 H9
S11.71991.71992.55532.55532.44582.44583.59803.5980
C21.71992.46301.35512.31451.07643.51202.14913.3331
C31.71992.46302.31451.35513.51201.07643.33312.1491
C42.55531.35512.31451.42942.18783.35751.07892.2207
C52.55532.31451.35511.42943.35752.18782.22071.0789
H62.44581.07643.51202.18783.35754.53762.61384.3348
H72.44583.51201.07643.35752.18784.53764.33482.6138
H83.59802.14913.33311.07892.22072.61384.33482.6355
H93.59803.33312.14912.22071.07894.33482.61382.6355

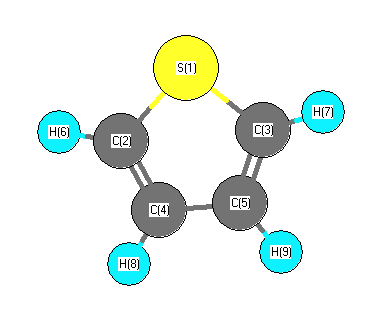
picture of Thiophene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C2 C4 111.856 S1 C2 H6 120.248
S1 C3 C5 111.856 S1 C3 H7 120.248
C2 S1 C3 91.452 C2 C4 C5 112.418
C2 C4 H8 123.599 C3 C5 C4 112.418
C3 C5 H9 123.599 C4 C2 H6 127.896
C4 C5 H9 123.983 C5 C3 H7 127.896
C5 C4 H8 123.983
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability