return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H14 (Hexane)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1Ag
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-236.751678
Energy at 298.15K-236.766771
HF Energy-236.491952
Nuclear repulsion energy243.791708
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3149 2989 0.00      
2 Ag 3075 2918 0.00      
3 Ag 3062 2906 0.00      
4 Ag 3046 2891 0.00      
5 Ag 1559 1480 0.00      
6 Ag 1542 1464 0.00      
7 Ag 1536 1458 0.00      
8 Ag 1460 1386 0.00      
9 Ag 1446 1373 0.00      
10 Ag 1372 1302 0.00      
11 Ag 1186 1126 0.00      
12 Ag 1097 1041 0.00      
13 Ag 1039 986 0.00      
14 Ag 927 879 0.00      
15 Ag 371 353 0.00      
16 Ag 299 284 0.00      
17 Au 3147 2987 120.38      
18 Au 3111 2953 83.54      
19 Au 3083 2927 0.30      
20 Au 1551 1472 11.91      
21 Au 1367 1297 0.84      
22 Au 1286 1221 0.00      
23 Au 1037 984 0.42      
24 Au 825 783 1.21      
25 Au 745 707 4.16      
26 Au 261 247 0.00      
27 Au 102 97 0.00      
28 Au 74 70 0.01      
29 Bg 3147 2987 0.00      
30 Bg 3100 2942 0.00      
31 Bg 3072 2916 0.00      
32 Bg 1551 1472 0.00      
33 Bg 1362 1292 0.00      
34 Bg 1340 1272 0.00      
35 Bg 1241 1178 0.00      
36 Bg 928 881 0.00      
37 Bg 761 722 0.00      
38 Bg 249 237 0.00      
39 Bg 151 144 0.00      
40 Bu 3149 2989 81.31      
41 Bu 3074 2918 73.85      
42 Bu 3065 2909 99.29      
43 Bu 3049 2894 11.14      
44 Bu 1564 1485 5.33      
45 Bu 1550 1472 1.46      
46 Bu 1536 1458 0.50      
47 Bu 1459 1385 3.57      
48 Bu 1426 1354 2.32      
49 Bu 1300 1234 4.43      
50 Bu 1102 1046 4.45      
51 Bu 1080 1025 0.56      
52 Bu 911 865 2.83      
53 Bu 471 447 0.08      
54 Bu 133 126 0.01      

Unscaled Zero Point Vibrational Energy (zpe) 42261.2 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 40114.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.48723 0.03779 0.03649

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.424 2.919 0.000
C2 -1.424 -2.919 0.000
C3 0.009 0.770 0.000
C4 -0.009 -0.770 0.000
C5 -1.424 -1.378 0.000
C6 1.424 1.378 0.000
H7 -2.439 -3.327 0.000
H8 2.439 3.327 0.000
H9 0.904 3.303 0.882
H10 0.904 3.303 -0.882
H11 -0.904 -3.303 -0.882
H12 -0.904 -3.303 0.882
H13 1.960 1.019 -0.885
H14 1.960 1.019 0.885
H15 -1.960 -1.019 -0.885
H16 -1.960 -1.019 0.885
H17 -0.535 1.133 0.880
H18 -0.535 1.133 -0.880
H19 0.535 -1.133 -0.880
H20 0.535 -1.133 0.880

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18 H19 H20
C16.49592.57373.95725.15591.54047.34441.09351.09381.09386.70146.70142.16342.16345.26675.26672.79292.79294.24114.2411
C26.49593.95722.57371.54045.15591.09357.34446.70146.70141.09381.09385.26675.26672.16342.16344.24114.24112.79292.7929
C32.57373.95721.53942.58231.54124.77233.52812.82782.82784.26564.26562.15662.15662.80262.80261.09611.09612.16162.1616
C43.95722.57371.53941.54122.58233.52814.77234.26564.26562.82782.82782.80262.80262.15662.15662.16162.16161.09611.0961
C55.15591.54042.58231.54123.96442.19706.08845.30225.30222.17982.17984.24034.24031.09491.09492.80622.80622.16162.1616
C61.54045.15591.54122.58233.96446.08842.19702.17982.17985.30225.30221.09491.09494.24034.24032.16162.16162.80622.8062
H77.34441.09354.77233.52812.19706.08848.25087.47737.47731.77031.77036.24626.24622.51852.51854.92924.92923.79863.7986
H81.09357.34443.52814.77236.08842.19708.25081.77031.77037.47737.47732.51852.51856.24626.24623.79863.79864.92924.9292
H91.09386.70142.82784.26565.30222.17987.47731.77031.76467.07206.84833.07462.51615.47695.18402.60313.14344.78754.4514
H101.09386.70142.82784.26565.30222.17987.47731.77031.76466.84837.07202.51613.07465.18405.47693.14342.60314.45144.7875
H116.70141.09384.26562.82782.17985.30221.77037.47737.07206.84831.76465.18405.47692.51613.07464.78754.45142.60313.1434
H126.70141.09384.26562.82782.17985.30221.77037.47736.84837.07201.76465.47695.18403.07462.51614.45144.78753.14342.6031
H132.16345.26672.15662.80264.24031.09496.24622.51853.07462.51615.18405.47691.76954.41704.75833.05752.49702.58093.1264
H142.16345.26672.15662.80264.24031.09496.24622.51852.51613.07465.47695.18401.76954.75834.41702.49703.05753.12642.5809
H155.26672.16342.80262.15661.09494.24032.51856.24625.47695.18402.51613.07464.41704.75831.76953.12642.58092.49703.0575
H165.26672.16342.80262.15661.09494.24032.51856.24625.18405.47693.07462.51614.75834.41701.76952.58093.12643.05752.4970
H172.79294.24111.09612.16162.80622.16164.92923.79862.60313.14344.78754.45143.05752.49703.12642.58091.75943.06242.5065
H182.79294.24111.09612.16162.80622.16164.92923.79863.14342.60314.45144.78752.49703.05752.58093.12641.75942.50653.0624
H194.24112.79292.16161.09612.16162.80623.79864.92924.78754.45142.60313.14342.58093.12642.49703.05753.06242.50651.7594
H204.24112.79292.16161.09612.16162.80623.79864.92924.45144.78753.14342.60313.12642.58093.05752.49702.50653.06241.7594

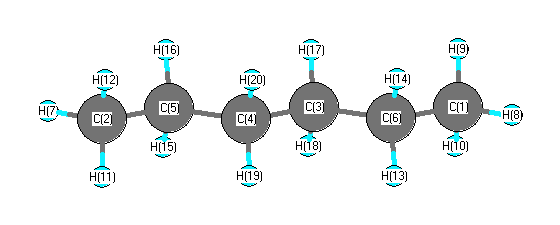
picture of Hexane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C6 C3 113.269 C1 C6 H13 109.190
C1 C6 H14 109.190 C2 C5 C4 113.269
C2 C5 H15 109.190 C2 C5 H16 109.190
C3 C4 C5 113.909 C3 C4 H19 109.044
C3 C4 H20 109.044 C3 C6 H13 108.613
C3 C6 H14 108.613 C4 C3 C6 113.909
C4 C3 H17 109.044 C4 C3 H18 109.044
C4 C5 H15 108.613 C4 C5 H16 108.613
C5 C2 H7 111.935 C5 C2 H11 110.538
C5 C2 H12 110.538 C5 C4 H19 108.925
C5 C4 H20 108.925 C6 C1 H8 111.935
C6 C1 H9 110.538 C6 C1 H10 110.538
C6 C3 H17 108.925 C6 C3 H18 108.925
H7 C2 H11 108.070 H7 C2 H12 108.070
H8 C1 H9 108.070 H8 C1 H10 108.070
H9 C1 H10 107.538 H11 C2 H12 107.538
H13 C6 H14 107.824 H15 C5 H16 107.824
H17 C3 H18 106.751 H19 C4 H20 106.751
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.459      
2 C -0.459      
3 C -0.270      
4 C -0.270      
5 C -0.272      
6 C -0.272      
7 H 0.150      
8 H 0.150      
9 H 0.148      
10 H 0.148      
11 H 0.148      
12 H 0.148      
13 H 0.140      
14 H 0.140      
15 H 0.140      
16 H 0.140      
17 H 0.137      
18 H 0.137      
19 H 0.137      
20 H 0.137      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 9.434 1.120 0.000
y 1.120 10.704 0.000
z 0.000 0.000 8.596


<r2> (average value of r2) Å2
<r2> 318.857
(<r2>)1/2 17.857