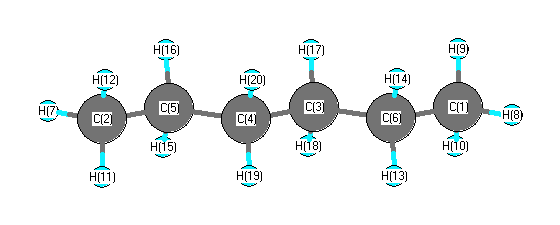
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3145 |
2991 |
0.00 |
|
|
|
| 2 |
Ag |
3064 |
2914 |
0.00 |
|
|
|
| 3 |
Ag |
3051 |
2901 |
0.00 |
|
|
|
| 4 |
Ag |
3034 |
2885 |
0.00 |
|
|
|
| 5 |
Ag |
1536 |
1460 |
0.00 |
|
|
|
| 6 |
Ag |
1518 |
1444 |
0.00 |
|
|
|
| 7 |
Ag |
1512 |
1438 |
0.00 |
|
|
|
| 8 |
Ag |
1438 |
1368 |
0.00 |
|
|
|
| 9 |
Ag |
1434 |
1364 |
0.00 |
|
|
|
| 10 |
Ag |
1353 |
1286 |
0.00 |
|
|
|
| 11 |
Ag |
1180 |
1122 |
0.00 |
|
|
|
| 12 |
Ag |
1103 |
1049 |
0.00 |
|
|
|
| 13 |
Ag |
1048 |
996 |
0.00 |
|
|
|
| 14 |
Ag |
921 |
875 |
0.00 |
|
|
|
| 15 |
Ag |
371 |
353 |
0.00 |
|
|
|
| 16 |
Ag |
296 |
282 |
0.00 |
|
|
|
| 17 |
Au |
3140 |
2986 |
111.33 |
|
|
|
| 18 |
Au |
3102 |
2950 |
81.20 |
|
|
|
| 19 |
Au |
3074 |
2924 |
0.38 |
|
|
|
| 20 |
Au |
1528 |
1453 |
15.04 |
|
|
|
| 21 |
Au |
1352 |
1285 |
0.88 |
|
|
|
| 22 |
Au |
1268 |
1206 |
0.01 |
|
|
|
| 23 |
Au |
1025 |
974 |
0.63 |
|
|
|
| 24 |
Au |
813 |
773 |
1.74 |
|
|
|
| 25 |
Au |
738 |
702 |
5.76 |
|
|
|
| 26 |
Au |
260 |
247 |
0.00 |
|
|
|
| 27 |
Au |
104 |
99 |
0.00 |
|
|
|
| 28 |
Au |
74 |
70 |
0.01 |
|
|
|
| 29 |
Bg |
3139 |
2985 |
0.00 |
|
|
|
| 30 |
Bg |
3091 |
2939 |
0.00 |
|
|
|
| 31 |
Bg |
3061 |
2911 |
0.00 |
|
|
|
| 32 |
Bg |
1528 |
1453 |
0.00 |
|
|
|
| 33 |
Bg |
1347 |
1281 |
0.00 |
|
|
|
| 34 |
Bg |
1324 |
1259 |
0.00 |
|
|
|
| 35 |
Bg |
1220 |
1161 |
0.00 |
|
|
|
| 36 |
Bg |
915 |
871 |
0.00 |
|
|
|
| 37 |
Bg |
752 |
715 |
0.00 |
|
|
|
| 38 |
Bg |
249 |
236 |
0.00 |
|
|
|
| 39 |
Bg |
151 |
143 |
0.00 |
|
|
|
| 40 |
Bu |
3145 |
2991 |
75.55 |
|
|
|
| 41 |
Bu |
3064 |
2913 |
79.50 |
|
|
|
| 42 |
Bu |
3054 |
2904 |
98.12 |
|
|
|
| 43 |
Bu |
3038 |
2889 |
11.55 |
|
|
|
| 44 |
Bu |
1540 |
1464 |
8.74 |
|
|
|
| 45 |
Bu |
1527 |
1452 |
1.52 |
|
|
|
| 46 |
Bu |
1511 |
1437 |
0.47 |
|
|
|
| 47 |
Bu |
1438 |
1368 |
6.60 |
|
|
|
| 48 |
Bu |
1409 |
1340 |
1.19 |
|
|
|
| 49 |
Bu |
1281 |
1219 |
2.71 |
|
|
|
| 50 |
Bu |
1100 |
1046 |
4.06 |
|
|
|
| 51 |
Bu |
1080 |
1027 |
2.46 |
|
|
|
| 52 |
Bu |
913 |
868 |
4.17 |
|
|
|
| 53 |
Bu |
466 |
443 |
0.09 |
|
|
|
| 54 |
Bu |
132 |
126 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 41974.6 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 39917.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.513 |
|
|
|
| 2 |
C |
-0.513 |
|
|
|
| 3 |
C |
-0.304 |
|
|
|
| 4 |
C |
-0.304 |
|
|
|
| 5 |
C |
-0.310 |
|
|
|
| 6 |
C |
-0.310 |
|
|
|
| 7 |
H |
0.168 |
|
|
|
| 8 |
H |
0.168 |
|
|
|
| 9 |
H |
0.166 |
|
|
|
| 10 |
H |
0.166 |
|
|
|
| 11 |
H |
0.166 |
|
|
|
| 12 |
H |
0.166 |
|
|
|
| 13 |
H |
0.159 |
|
|
|
| 14 |
H |
0.159 |
|
|
|
| 15 |
H |
0.159 |
|
|
|
| 16 |
H |
0.159 |
|
|
|
| 17 |
H |
0.155 |
|
|
|
| 18 |
H |
0.155 |
|
|
|
| 19 |
H |
0.155 |
|
|
|
| 20 |
H |
0.155 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.321 |
-0.084 |
0.000 |
| y |
-0.084 |
-42.254 |
0.000 |
| z |
0.000 |
0.000 |
-40.523 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.068 |
-0.084 |
0.000 |
| y |
-0.084 |
-1.332 |
0.000 |
| z |
0.000 |
0.000 |
1.265 |
|
| Polar |
|---|
| 3z2-r2 | 2.529 |
|---|
| x2-y2 | 0.933 |
|---|
| xy | -0.084 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.566 |
1.149 |
0.000 |
| y |
1.149 |
10.893 |
0.000 |
| z |
0.000 |
0.000 |
8.675 |
<r2> (average value of r
2) Å
2
| <r2> |
317.010 |
| (<r2>)1/2 |
17.805 |