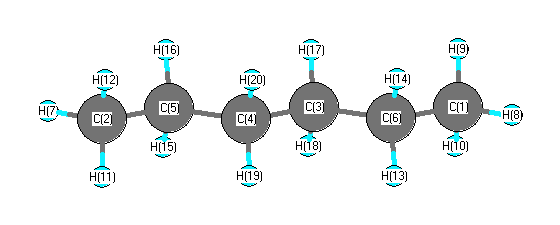
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3056 |
3005 |
0.00 |
|
|
|
| 2 |
Ag |
2977 |
2928 |
0.00 |
|
|
|
| 3 |
Ag |
2960 |
2911 |
0.00 |
|
|
|
| 4 |
Ag |
2943 |
2895 |
0.00 |
|
|
|
| 5 |
Ag |
1490 |
1466 |
0.00 |
|
|
|
| 6 |
Ag |
1472 |
1448 |
0.00 |
|
|
|
| 7 |
Ag |
1464 |
1440 |
0.00 |
|
|
|
| 8 |
Ag |
1391 |
1368 |
0.00 |
|
|
|
| 9 |
Ag |
1373 |
1351 |
0.00 |
|
|
|
| 10 |
Ag |
1308 |
1286 |
0.00 |
|
|
|
| 11 |
Ag |
1138 |
1120 |
0.00 |
|
|
|
| 12 |
Ag |
1062 |
1044 |
0.00 |
|
|
|
| 13 |
Ag |
1011 |
994 |
0.00 |
|
|
|
| 14 |
Ag |
891 |
876 |
0.00 |
|
|
|
| 15 |
Ag |
359 |
353 |
0.00 |
|
|
|
| 16 |
Ag |
288 |
283 |
0.00 |
|
|
|
| 17 |
Au |
3049 |
2999 |
107.54 |
|
|
|
| 18 |
Au |
3010 |
2960 |
85.47 |
|
|
|
| 19 |
Au |
2981 |
2932 |
0.23 |
|
|
|
| 20 |
Au |
1482 |
1457 |
13.81 |
|
|
|
| 21 |
Au |
1308 |
1287 |
0.81 |
|
|
|
| 22 |
Au |
1225 |
1205 |
0.02 |
|
|
|
| 23 |
Au |
991 |
975 |
0.69 |
|
|
|
| 24 |
Au |
789 |
776 |
1.90 |
|
|
|
| 25 |
Au |
720 |
708 |
6.22 |
|
|
|
| 26 |
Au |
252 |
247 |
0.00 |
|
|
|
| 27 |
Au |
100 |
98 |
0.00 |
|
|
|
| 28 |
Au |
73 |
72 |
0.01 |
|
|
|
| 29 |
Bg |
3049 |
2998 |
0.00 |
|
|
|
| 30 |
Bg |
2998 |
2948 |
0.00 |
|
|
|
| 31 |
Bg |
2969 |
2920 |
0.00 |
|
|
|
| 32 |
Bg |
1482 |
1458 |
0.00 |
|
|
|
| 33 |
Bg |
1301 |
1280 |
0.00 |
|
|
|
| 34 |
Bg |
1281 |
1260 |
0.00 |
|
|
|
| 35 |
Bg |
1180 |
1160 |
0.00 |
|
|
|
| 36 |
Bg |
887 |
873 |
0.00 |
|
|
|
| 37 |
Bg |
732 |
720 |
0.00 |
|
|
|
| 38 |
Bg |
240 |
236 |
0.00 |
|
|
|
| 39 |
Bg |
149 |
147 |
0.00 |
|
|
|
| 40 |
Bu |
3056 |
3005 |
75.36 |
|
|
|
| 41 |
Bu |
2977 |
2928 |
80.50 |
|
|
|
| 42 |
Bu |
2963 |
2914 |
103.32 |
|
|
|
| 43 |
Bu |
2948 |
2899 |
9.94 |
|
|
|
| 44 |
Bu |
1495 |
1470 |
6.96 |
|
|
|
| 45 |
Bu |
1481 |
1457 |
1.44 |
|
|
|
| 46 |
Bu |
1464 |
1440 |
0.48 |
|
|
|
| 47 |
Bu |
1389 |
1366 |
4.15 |
|
|
|
| 48 |
Bu |
1359 |
1337 |
2.68 |
|
|
|
| 49 |
Bu |
1239 |
1218 |
4.76 |
|
|
|
| 50 |
Bu |
1063 |
1045 |
4.39 |
|
|
|
| 51 |
Bu |
1043 |
1025 |
2.12 |
|
|
|
| 52 |
Bu |
882 |
868 |
4.06 |
|
|
|
| 53 |
Bu |
453 |
445 |
0.11 |
|
|
|
| 54 |
Bu |
126 |
124 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 40682.3 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 40011.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.482 |
|
|
|
| 2 |
C |
-0.482 |
|
|
|
| 3 |
C |
-0.274 |
|
|
|
| 4 |
C |
-0.274 |
|
|
|
| 5 |
C |
-0.279 |
|
|
|
| 6 |
C |
-0.279 |
|
|
|
| 7 |
H |
0.155 |
|
|
|
| 8 |
H |
0.155 |
|
|
|
| 9 |
H |
0.155 |
|
|
|
| 10 |
H |
0.155 |
|
|
|
| 11 |
H |
0.155 |
|
|
|
| 12 |
H |
0.155 |
|
|
|
| 13 |
H |
0.145 |
|
|
|
| 14 |
H |
0.145 |
|
|
|
| 15 |
H |
0.145 |
|
|
|
| 16 |
H |
0.145 |
|
|
|
| 17 |
H |
0.140 |
|
|
|
| 18 |
H |
0.140 |
|
|
|
| 19 |
H |
0.140 |
|
|
|
| 20 |
H |
0.140 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.416 |
-0.066 |
0.000 |
| y |
-0.066 |
-42.276 |
0.000 |
| z |
0.000 |
0.000 |
-40.681 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.063 |
-0.066 |
0.000 |
| y |
-0.066 |
-1.228 |
0.000 |
| z |
0.000 |
0.000 |
1.165 |
|
| Polar |
|---|
| 3z2-r2 | 2.330 |
|---|
| x2-y2 | 0.860 |
|---|
| xy | -0.066 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.874 |
1.290 |
0.000 |
| y |
1.290 |
11.388 |
0.000 |
| z |
0.000 |
0.000 |
8.859 |
<r2> (average value of r
2) Å
2
| <r2> |
321.362 |
| (<r2>)1/2 |
17.927 |