return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H8Cl2 (1,4-Dichlorobutane)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1Ag
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-1077.112469
Energy at 298.15K-1077.121552
HF Energy-1076.854833
Nuclear repulsion energy326.155995
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3132 2973 0.00      
2 Ag 3084 2928 0.00      
3 Ag 1542 1463 0.00      
4 Ag 1540 1461 0.00      
5 Ag 1436 1363 0.00      
6 Ag 1341 1272 0.00      
7 Ag 1107 1051 0.00      
8 Ag 1058 1004 0.00      
9 Ag 776 736 0.00      
10 Ag 336 319 0.00      
11 Ag 225 214 0.00      
12 Au 3195 3033 28.52      
13 Au 3141 2981 20.00      
14 Au 1354 1285 2.05      
15 Au 1165 1106 1.65      
16 Au 926 879 0.81      
17 Au 762 724 4.08      
18 Au 111 105 1.16      
19 Au 61 58 5.45      
20 Bg 3193 3031 0.00      
21 Bg 3123 2964 0.00      
22 Bg 1359 1290 0.00      
23 Bg 1297 1231 0.00      
24 Bg 1112 1056 0.00      
25 Bg 800 759 0.00      
26 Bg 156 149 0.00      
27 Bu 3133 2974 42.86      
28 Bu 3091 2934 19.38      
29 Bu 1558 1479 5.66      
30 Bu 1536 1458 1.12      
31 Bu 1397 1326 71.31      
32 Bu 1272 1208 11.00      
33 Bu 1056 1002 17.45      
34 Bu 752 714 77.99      
35 Bu 425 403 14.99      
36 Bu 104 99 5.00      

Unscaled Zero Point Vibrational Energy (zpe) 25827.9 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 24515.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.52584 0.01804 0.01767

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
Cl1 -0.406 -3.419 0.000
Cl2 0.406 3.419 0.000
C3 -0.512 1.880 0.000
C4 0.512 -1.880 0.000
C5 0.406 0.657 0.000
C6 -0.406 -0.657 0.000
H7 -1.149 1.882 0.885
H8 -1.149 1.882 -0.885
H9 1.149 -1.882 -0.885
H10 1.149 -1.882 0.885
H11 1.049 0.702 -0.884
H12 -1.049 -0.702 -0.884
H13 -1.049 -0.702 0.884
H14 1.049 0.702 0.884

Atom - Atom Distances (Å)
  Cl1 Cl2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14
Cl16.88655.29991.79254.15602.76275.42545.42542.35952.35954.45862.92922.92924.4586
Cl26.88651.79255.29992.76274.15602.35952.35955.42545.42542.92924.45864.45862.9292
C35.29991.79253.89621.52932.53841.09081.09084.20604.20602.14592.78082.78082.1459
C41.79255.29993.89622.53841.52934.20604.20601.09081.09082.78082.14592.14592.7808
C54.15602.76271.52932.53841.54412.16892.16892.78892.78891.09392.17782.17781.0939
C62.76274.15602.53841.52931.54412.78892.78892.16892.16892.17781.09391.09392.1778
H75.42542.35951.09084.20602.16892.78891.77014.75194.40993.05853.13262.58522.4949
H85.42542.35951.09084.20602.16892.78891.77014.40994.75192.49492.58523.13263.0585
H92.35955.42544.20601.09082.78892.16894.75194.40991.77012.58522.49493.05853.1326
H102.35955.42544.20601.09082.78892.16894.40994.75191.77013.13263.05852.49492.5852
H114.45862.92922.14592.78081.09392.17783.05852.49492.58523.13262.52343.08141.7684
H122.92924.45862.78082.14592.17781.09393.13262.58522.49493.05852.52341.76843.0814
H132.92924.45862.78082.14592.17781.09392.58523.13263.05852.49493.08141.76842.5234
H144.45862.92922.14592.78081.09392.17782.49493.05853.13262.58521.76843.08142.5234

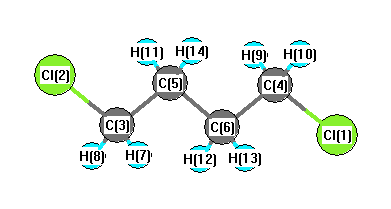
picture of 1,4-Dichlorobutane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
Cl1 C4 C6 112.305 Cl1 C4 H9 107.326
Cl1 C4 H10 107.326 Cl2 C3 C5 112.305
Cl2 C3 H7 107.326 Cl2 C3 H8 107.326
C3 C5 C6 111.366 C3 C5 H11 108.641
C3 C5 H14 108.641 C4 C6 C5 111.366
C4 C6 H12 108.641 C4 C6 H13 108.641
C5 C3 H7 110.631 C5 C3 H8 110.631
C5 C6 H12 110.121 C5 C6 H13 110.121
C6 C4 H9 110.631 C6 C4 H10 110.631
C6 C5 H11 110.121 C6 C5 H14 110.121
H7 C3 H8 108.455 H9 C4 H10 108.455
H11 C5 H14 107.863 H12 C6 H13 107.863
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 Cl -0.092      
2 Cl -0.092      
3 C -0.376      
4 C -0.376      
5 C -0.288      
6 C -0.288      
7 H 0.204      
8 H 0.204      
9 H 0.204      
10 H 0.204      
11 H 0.174      
12 H 0.174      
13 H 0.174      
14 H 0.174      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 7.790 1.395 0.000
y 1.395 12.731 0.000
z 0.000 0.000 6.938


<r2> (average value of r2) Å2
<r2> 520.172
(<r2>)1/2 22.807