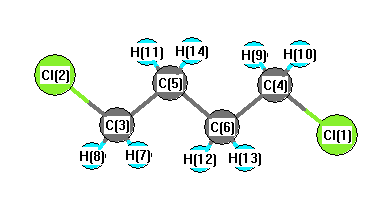
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3117 |
2965 |
0.00 |
|
|
|
| 2 |
Ag |
3073 |
2922 |
0.00 |
|
|
|
| 3 |
Ag |
1518 |
1444 |
0.00 |
|
|
|
| 4 |
Ag |
1517 |
1442 |
0.00 |
|
|
|
| 5 |
Ag |
1421 |
1352 |
0.00 |
|
|
|
| 6 |
Ag |
1324 |
1259 |
0.00 |
|
|
|
| 7 |
Ag |
1110 |
1055 |
0.00 |
|
|
|
| 8 |
Ag |
1063 |
1011 |
0.00 |
|
|
|
| 9 |
Ag |
786 |
747 |
0.00 |
|
|
|
| 10 |
Ag |
334 |
318 |
0.00 |
|
|
|
| 11 |
Ag |
226 |
215 |
0.00 |
|
|
|
| 12 |
Au |
3182 |
3026 |
26.89 |
|
|
|
| 13 |
Au |
3132 |
2978 |
16.79 |
|
|
|
| 14 |
Au |
1339 |
1273 |
2.16 |
|
|
|
| 15 |
Au |
1155 |
1098 |
1.83 |
|
|
|
| 16 |
Au |
914 |
870 |
1.06 |
|
|
|
| 17 |
Au |
755 |
718 |
5.51 |
|
|
|
| 18 |
Au |
112 |
107 |
1.10 |
|
|
|
| 19 |
Au |
62 |
59 |
5.49 |
|
|
|
| 20 |
Bg |
3180 |
3024 |
0.00 |
|
|
|
| 21 |
Bg |
3113 |
2960 |
0.00 |
|
|
|
| 22 |
Bg |
1344 |
1278 |
0.00 |
|
|
|
| 23 |
Bg |
1282 |
1219 |
0.00 |
|
|
|
| 24 |
Bg |
1098 |
1044 |
0.00 |
|
|
|
| 25 |
Bg |
790 |
751 |
0.00 |
|
|
|
| 26 |
Bg |
157 |
149 |
0.00 |
|
|
|
| 27 |
Bu |
3118 |
2966 |
43.45 |
|
|
|
| 28 |
Bu |
3081 |
2930 |
18.16 |
|
|
|
| 29 |
Bu |
1533 |
1458 |
8.18 |
|
|
|
| 30 |
Bu |
1514 |
1439 |
0.99 |
|
|
|
| 31 |
Bu |
1380 |
1312 |
71.78 |
|
|
|
| 32 |
Bu |
1255 |
1194 |
7.14 |
|
|
|
| 33 |
Bu |
1063 |
1011 |
18.88 |
|
|
|
| 34 |
Bu |
761 |
724 |
79.42 |
|
|
|
| 35 |
Bu |
425 |
405 |
14.81 |
|
|
|
| 36 |
Bu |
104 |
99 |
5.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25668.8 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 24411.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Cl |
-0.067 |
|
|
|
| 2 |
Cl |
-0.067 |
|
|
|
| 3 |
C |
-0.434 |
|
|
|
| 4 |
C |
-0.434 |
|
|
|
| 5 |
C |
-0.321 |
|
|
|
| 6 |
C |
-0.321 |
|
|
|
| 7 |
H |
0.221 |
|
|
|
| 8 |
H |
0.221 |
|
|
|
| 9 |
H |
0.221 |
|
|
|
| 10 |
H |
0.221 |
|
|
|
| 11 |
H |
0.191 |
|
|
|
| 12 |
H |
0.191 |
|
|
|
| 13 |
H |
0.191 |
|
|
|
| 14 |
H |
0.191 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.989 |
-4.447 |
0.000 |
| y |
-4.447 |
-68.584 |
0.000 |
| z |
0.000 |
0.000 |
-50.162 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
9.384 |
-4.447 |
0.000 |
| y |
-4.447 |
-18.509 |
0.000 |
| z |
0.000 |
0.000 |
9.125 |
|
| Polar |
|---|
| 3z2-r2 | 18.249 |
|---|
| x2-y2 | 18.595 |
|---|
| xy | -4.447 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.863 |
1.347 |
0.000 |
| y |
1.347 |
12.844 |
0.000 |
| z |
0.000 |
0.000 |
6.991 |
<r2> (average value of r
2) Å
2
| <r2> |
516.167 |
| (<r2>)1/2 |
22.719 |