return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H9N (Pentanenitrile)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-250.395960
Energy at 298.15K-250.405449
HF Energy-250.127325
Nuclear repulsion energy210.496321
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3158 2998 32.23      
2 A' 3091 2934 34.74      
3 A' 3082 2926 10.15      
4 A' 3079 2923 12.96      
5 A' 3063 2907 19.76      
6 A' 2305 2188 3.08      
7 A' 1561 1481 5.37      
8 A' 1547 1469 0.79      
9 A' 1539 1460 1.10      
10 A' 1521 1444 2.51      
11 A' 1462 1388 2.09      
12 A' 1440 1367 0.52      
13 A' 1392 1321 2.67      
14 A' 1307 1241 1.91      
15 A' 1150 1092 3.90      
16 A' 1087 1031 0.23      
17 A' 1041 988 0.55      
18 A' 952 904 0.16      
19 A' 921 874 2.87      
20 A' 539 512 0.87      
21 A' 388 368 0.74      
22 A' 282 268 1.19      
23 A' 126 120 5.61      
24 A" 3154 2993 58.20      
25 A" 3140 2981 15.07      
26 A" 3120 2962 2.11      
27 A" 3096 2938 8.62      
28 A" 1550 1472 7.04      
29 A" 1363 1294 0.16      
30 A" 1346 1277 0.63      
31 A" 1273 1209 0.04      
32 A" 1167 1108 0.00      
33 A" 956 908 0.75      
34 A" 811 770 0.03      
35 A" 749 711 3.98      
36 A" 384 364 0.54      
37 A" 251 238 0.10      
38 A" 116 110 0.26      
39 A" 89 85 3.80      

Unscaled Zero Point Vibrational Energy (zpe) 29298.8 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 27810.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.50494 0.04409 0.04181

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.402 0.000
C2 1.418 0.991 0.000
C3 1.407 2.528 0.000
C4 -0.002 -1.151 0.000
N5 -2.431 -2.121 0.000
C6 -1.337 -1.713 0.000
H7 2.421 2.935 0.000
H8 -0.546 0.755 0.880
H9 -0.546 0.755 -0.880
H10 1.954 0.631 0.884
H11 1.954 0.631 -0.884
H12 0.888 2.910 -0.883
H13 0.888 2.910 0.883
H14 0.543 -1.507 -0.882
H15 0.543 -1.507 0.882

Atom - Atom Distances (Å)
  C1 C2 C3 C4 N5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.53582.55001.55303.50342.50203.50451.09401.09402.15712.15712.80322.80322.17132.1713
C21.53581.53762.57024.94983.86062.18772.16542.16541.09491.09492.17782.17782.78952.7895
C32.55001.53763.94026.02885.05171.09312.78102.78102.16392.16391.09291.09294.21974.2197
C41.55302.57023.94022.61551.44884.75092.16892.16892.79002.79004.25004.25001.09601.0960
N53.50344.94986.02882.61551.16727.00793.54923.54925.25205.25206.09126.09123.16243.1624
C62.50203.86065.05171.44881.16725.97762.73672.73674.13634.13635.20575.20572.08732.0873
H73.50452.18771.09314.75097.00795.97763.78613.78612.51192.51191.77001.77004.90254.9025
H81.09402.16542.78102.16893.54922.73673.78611.75912.50353.06253.13152.58843.06672.5103
H91.09402.16542.78102.16893.54922.73673.78611.75913.06252.50352.58843.13152.51033.0667
H102.15711.09492.16392.79005.25204.13632.51192.50353.06251.76883.07472.51593.11132.5612
H112.15711.09492.16392.79005.25204.13632.51193.06252.50351.76882.51593.07472.56123.1113
H122.80322.17781.09294.25006.09125.20571.77003.13152.58843.07472.51591.76614.42974.7684
H132.80322.17781.09294.25006.09125.20571.77002.58843.13152.51593.07471.76614.76844.4297
H142.17132.78954.21971.09603.16242.08734.90253.06672.51033.11132.56124.42974.76841.7641
H152.17132.78954.21971.09603.16242.08734.90252.51033.06672.56123.11134.76844.42971.7641

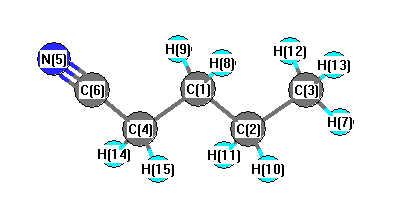
picture of Pentanenitrile state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 112.133 C1 C2 H10 109.013
C1 C2 H11 109.013 C1 C4 C6 112.872
C1 C4 H14 108.881 C1 C4 H15 108.881
C2 C1 C4 112.627 C2 C1 H8 109.712
C2 C1 H9 109.712 C2 C3 H7 111.403
C2 C3 H12 110.627 C2 C3 H13 110.627
C3 C2 H10 109.418 C3 C2 H11 109.418
C4 C1 H8 108.804 C4 C1 H9 108.804
C4 C6 N5 177.667 C6 C4 H14 109.434
C6 C4 H15 109.434 H7 C3 H12 108.127
H7 C3 H13 108.127 H8 C1 H9 107.018
H10 C2 H11 107.745 H12 C3 H13 107.804
H14 C4 H15 107.176
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.282      
2 C -0.279      
3 C -0.462      
4 C -0.351      
5 N -0.478      
6 C 0.336      
7 H 0.159      
8 H 0.170      
9 H 0.170      
10 H 0.149      
11 H 0.149      
12 H 0.156      
13 H 0.156      
14 H 0.205      
15 H 0.205      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 9.285 1.715 0.000
y 1.715 8.400 0.000
z 0.000 0.000 6.597


<r2> (average value of r2) Å2
<r2> 259.042
(<r2>)1/2 16.095