return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H9N (Pentanenitrile)

using model chemistry: MP3/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP3/6-31G*
 hartrees
Energy at 0K-249.872260
Energy at 298.15K-249.881786
HF Energy-249.030513
Nuclear repulsion energy210.750157
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3183 2988 28.21      
2 A' 3115 2924 19.81      
3 A' 3105 2915 14.03      
4 A' 3100 2910 15.72      
5 A' 3087 2897 17.54      
6 A' 2447 2297 4.66      
7 A' 1573 1476 7.38      
8 A' 1559 1463 0.68      
9 A' 1554 1458 1.51      
10 A' 1538 1444 3.21      
11 A' 1479 1388 3.08      
12 A' 1466 1376 1.26      
13 A' 1407 1321 1.69      
14 A' 1316 1235 0.62      
15 A' 1165 1094 3.90      
16 A' 1111 1043 0.17      
17 A' 1067 1002 0.58      
18 A' 967 908 0.69      
19 A' 936 878 3.49      
20 A' 541 508 0.76      
21 A' 395 371 0.75      
22 A' 284 266 1.23      
23 A' 126 118 5.99      
24 A" 3178 2983 48.16      
25 A" 3166 2972 13.00      
26 A" 3147 2953 5.41      
27 A" 3122 2930 6.81      
28 A" 1560 1464 7.93      
29 A" 1373 1288 0.16      
30 A" 1357 1273 0.65      
31 A" 1286 1207 0.02      
32 A" 1178 1106 0.01      
33 A" 966 907 0.69      
34 A" 820 769 0.03      
35 A" 752 705 4.05      
36 A" 377 354 0.62      
37 A" 253 238 0.11      
38 A" 117 110 0.13      
39 A" 89 84 4.07      

Unscaled Zero Point Vibrational Energy (zpe) 29629.2 cm-1
Scaled (by 0.9386) Zero Point Vibrational Energy (zpe) 27810.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP3/6-31G*
ABC
0.50309 0.04427 0.04196

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP3/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.405 0.000
C2 1.421 0.969 0.000
C3 1.441 2.497 0.000
C4 -0.005 -1.131 0.000
N5 -2.450 -2.101 0.000
C6 -1.368 -1.684 0.000
H7 2.466 2.879 0.000
H8 -0.545 0.768 0.879
H9 -0.545 0.768 -0.879
H10 1.960 0.593 0.879
H11 1.960 0.593 -0.879
H12 0.933 2.895 -0.885
H13 0.933 2.895 0.885
H14 0.522 -1.514 -0.881
H15 0.522 -1.514 0.881

Atom - Atom Distances (Å)
  C1 C2 C3 C4 N5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.52902.54021.53593.50442.49723.49301.09621.09622.15662.15662.80202.80202.17562.1756
C21.52901.52842.53794.94043.84922.17712.16352.16351.09771.09772.17482.17482.78402.7840
C32.54021.52843.90526.02315.03701.09402.77672.77672.16092.16091.09501.09504.20864.2086
C41.53592.53793.90522.63061.47124.70962.16072.16072.75752.75754.22674.22671.09561.0956
N53.50444.94046.02312.63061.15966.99733.55353.55355.24185.24186.09756.09753.15483.1548
C62.49723.84925.03701.47121.15965.95982.73142.73144.12714.12715.20025.20022.09192.0919
H73.49302.17711.09404.70966.99735.95983.78153.78152.50092.50091.77031.77034.88414.8841
H81.09622.16352.77672.16073.55352.73143.78151.75882.51163.06603.13412.59033.07312.5193
H91.09622.16352.77672.16073.55352.73143.78151.75883.06602.51162.59033.13412.51933.0731
H102.15661.09772.16092.75755.24184.12712.50092.51163.06601.75813.07662.52073.09912.5511
H112.15661.09772.16092.75755.24184.12712.50093.06602.51161.75812.52073.07662.55113.0991
H122.80202.17481.09504.22676.09755.20021.77033.13412.59033.07662.52071.76994.42814.7671
H132.80202.17481.09504.22676.09755.20021.77032.59033.13412.52073.07661.76994.76714.4281
H142.17562.78404.20861.09563.15482.09194.88413.07312.51933.09912.55114.42814.76711.7611
H152.17562.78404.20861.09563.15482.09194.88412.51933.07312.55113.09914.76714.42811.7611

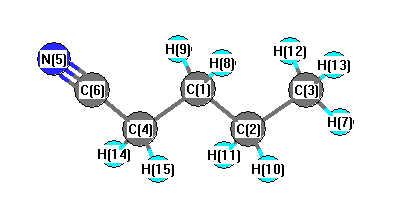
picture of Pentanenitrile state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 112.374 C1 C2 H10 109.282
C1 C2 H11 109.282 C1 C4 C6 112.271
C1 C4 H14 110.419 C1 C4 H15 110.419
C2 C1 C4 111.804 C2 C1 H8 109.901
C2 C1 H9 109.901 C2 C3 H7 111.154
C2 C3 H12 110.912 C2 C3 H13 110.912
C3 C2 H10 109.654 C3 C2 H11 109.654
C4 C1 H8 109.208 C4 C1 H9 109.208
C4 C6 N5 178.946 C6 C4 H14 108.287
C6 C4 H15 108.287 H7 C3 H12 107.942
H7 C3 H13 107.942 H8 C1 H9 106.680
H10 C2 H11 106.417 H12 C3 H13 107.838
H14 C4 H15 106.978
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability