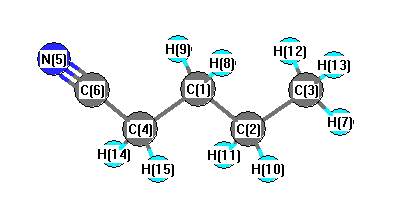
Vibrational Frequencies calculated at B3LYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3119 |
2995 |
35.31 |
|
|
|
| 2 |
A' |
3057 |
2936 |
41.79 |
|
|
|
| 3 |
A' |
3048 |
2927 |
9.32 |
|
|
|
| 4 |
A' |
3045 |
2924 |
11.97 |
|
|
|
| 5 |
A' |
3027 |
2907 |
20.83 |
|
|
|
| 6 |
A' |
2368 |
2274 |
10.54 |
|
|
|
| 7 |
A' |
1541 |
1480 |
5.14 |
|
|
|
| 8 |
A' |
1527 |
1467 |
0.59 |
|
|
|
| 9 |
A' |
1518 |
1457 |
1.12 |
|
|
|
| 10 |
A' |
1500 |
1440 |
2.43 |
|
|
|
| 11 |
A' |
1442 |
1385 |
1.76 |
|
|
|
| 12 |
A' |
1421 |
1364 |
0.41 |
|
|
|
| 13 |
A' |
1376 |
1322 |
3.10 |
|
|
|
| 14 |
A' |
1295 |
1243 |
2.74 |
|
|
|
| 15 |
A' |
1134 |
1089 |
3.97 |
|
|
|
| 16 |
A' |
1066 |
1024 |
0.24 |
|
|
|
| 17 |
A' |
1023 |
982 |
0.51 |
|
|
|
| 18 |
A' |
944 |
907 |
0.20 |
|
|
|
| 19 |
A' |
908 |
872 |
2.67 |
|
|
|
| 20 |
A' |
543 |
522 |
0.94 |
|
|
|
| 21 |
A' |
383 |
368 |
0.80 |
|
|
|
| 22 |
A' |
282 |
270 |
1.28 |
|
|
|
| 23 |
A' |
126 |
121 |
5.61 |
|
|
|
| 24 |
A" |
3114 |
2990 |
70.29 |
|
|
|
| 25 |
A" |
3102 |
2979 |
10.82 |
|
|
|
| 26 |
A" |
3080 |
2958 |
0.97 |
|
|
|
| 27 |
A" |
3055 |
2934 |
9.25 |
|
|
|
| 28 |
A" |
1529 |
1469 |
6.84 |
|
|
|
| 29 |
A" |
1347 |
1294 |
0.14 |
|
|
|
| 30 |
A" |
1330 |
1277 |
0.61 |
|
|
|
| 31 |
A" |
1257 |
1207 |
0.06 |
|
|
|
| 32 |
A" |
1153 |
1108 |
0.00 |
|
|
|
| 33 |
A" |
945 |
907 |
0.85 |
|
|
|
| 34 |
A" |
802 |
770 |
0.03 |
|
|
|
| 35 |
A" |
743 |
713 |
4.27 |
|
|
|
| 36 |
A" |
393 |
378 |
0.57 |
|
|
|
| 37 |
A" |
247 |
237 |
0.09 |
|
|
|
| 38 |
A" |
115 |
111 |
0.42 |
|
|
|
| 39 |
A" |
89 |
85 |
3.72 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28997.3 cm
-1
Scaled (by 0.9603) Zero Point Vibrational Energy (zpe) 27846.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.256 |
0.092 |
|
0.096 |
| 2 |
C |
-0.252 |
0.129 |
|
0.121 |
| 3 |
C |
-0.444 |
-0.230 |
|
-0.330 |
| 4 |
C |
-0.341 |
-0.173 |
|
-0.347 |
| 5 |
N |
-0.474 |
-0.462 |
|
-0.472 |
| 6 |
C |
0.348 |
0.352 |
|
0.403 |
| 7 |
H |
0.150 |
0.062 |
|
0.091 |
| 8 |
H |
0.157 |
0.001 |
|
0.017 |
| 9 |
H |
0.157 |
0.001 |
|
0.017 |
| 10 |
H |
0.138 |
-0.019 |
|
-0.006 |
| 11 |
H |
0.138 |
-0.019 |
|
-0.006 |
| 12 |
H |
0.149 |
0.053 |
|
0.080 |
| 13 |
H |
0.149 |
0.053 |
|
0.080 |
| 14 |
H |
0.191 |
0.080 |
|
0.129 |
| 15 |
H |
0.191 |
0.080 |
|
0.129 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.706 |
1.867 |
0.000 |
4.149 |
| CHELPG |
3.680 |
1.849 |
0.000 |
4.118 |
| AIM |
|
|
|
|
| ESP |
3.709 |
1.869 |
0.000 |
4.153 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.496 |
-8.026 |
0.000 |
| y |
-8.026 |
-41.879 |
0.000 |
| z |
0.000 |
0.000 |
-36.514 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-8.300 |
-8.026 |
0.000 |
| y |
-8.026 |
0.126 |
0.000 |
| z |
0.000 |
0.000 |
8.174 |
|
| Polar |
|---|
| 3z2-r2 | 16.347 |
|---|
| x2-y2 | -5.617 |
|---|
| xy | -8.026 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.464 |
1.803 |
0.000 |
| y |
1.803 |
8.567 |
0.000 |
| z |
0.000 |
0.000 |
6.650 |
<r2> (average value of r
2) Å
2
| <r2> |
259.725 |
| (<r2>)1/2 |
16.116 |