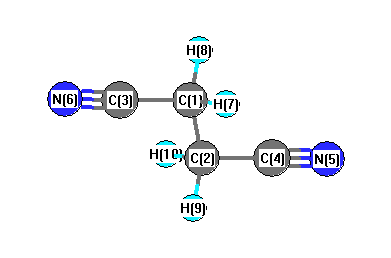
Vibrational Frequencies calculated at B3LYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3072 |
2950 |
0.00 |
|
|
|
| 2 |
Ag |
2375 |
2280 |
0.00 |
|
|
|
| 3 |
Ag |
1498 |
1439 |
0.00 |
|
|
|
| 4 |
Ag |
1401 |
1346 |
0.00 |
|
|
|
| 5 |
Ag |
1026 |
985 |
0.00 |
|
|
|
| 6 |
Ag |
975 |
936 |
0.00 |
|
|
|
| 7 |
Ag |
515 |
494 |
0.00 |
|
|
|
| 8 |
Ag |
224 |
216 |
0.00 |
|
|
|
| 9 |
Au |
3128 |
3003 |
1.60 |
|
|
|
| 10 |
Au |
1226 |
1177 |
0.13 |
|
|
|
| 11 |
Au |
780 |
749 |
3.64 |
|
|
|
| 12 |
Au |
409 |
393 |
0.22 |
|
|
|
| 13 |
Au |
80 |
77 |
20.58 |
|
|
|
| 14 |
Bg |
3109 |
2986 |
0.00 |
|
|
|
| 15 |
Bg |
1329 |
1276 |
0.00 |
|
|
|
| 16 |
Bg |
1052 |
1010 |
0.00 |
|
|
|
| 17 |
Bg |
369 |
354 |
0.00 |
|
|
|
| 18 |
Bu |
3081 |
2959 |
3.85 |
|
|
|
| 19 |
Bu |
2376 |
2282 |
7.43 |
|
|
|
| 20 |
Bu |
1506 |
1446 |
12.70 |
|
|
|
| 21 |
Bu |
1310 |
1258 |
2.36 |
|
|
|
| 22 |
Bu |
937 |
900 |
5.51 |
|
|
|
| 23 |
Bu |
530 |
509 |
1.19 |
|
|
|
| 24 |
Bu |
132 |
127 |
21.90 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16219.5 cm
-1
Scaled (by 0.9603) Zero Point Vibrational Energy (zpe) 15575.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.343 |
-0.011 |
|
-0.114 |
| 2 |
C |
-0.343 |
-0.011 |
|
-0.083 |
| 3 |
C |
0.356 |
0.288 |
|
0.306 |
| 4 |
C |
0.356 |
0.288 |
|
0.297 |
| 5 |
N |
-0.454 |
-0.418 |
|
-0.414 |
| 6 |
N |
-0.454 |
-0.418 |
|
-0.417 |
| 7 |
H |
0.221 |
0.071 |
|
0.110 |
| 8 |
H |
0.221 |
0.071 |
|
0.110 |
| 9 |
H |
0.221 |
0.071 |
|
0.103 |
| 10 |
H |
0.221 |
0.071 |
|
0.103 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
0.000 |
0.000 |
0.000 |
0.000 |
| AIM |
|
|
|
|
| ESP |
-0.001 |
-0.004 |
0.000 |
0.004 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-52.707 |
12.915 |
0.000 |
| y |
12.915 |
-39.277 |
0.000 |
| z |
0.000 |
0.000 |
-32.483 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-16.827 |
12.915 |
0.000 |
| y |
12.915 |
3.318 |
0.000 |
| z |
0.000 |
0.000 |
13.509 |
|
| Polar |
|---|
| 3z2-r2 | 27.018 |
|---|
| x2-y2 | -13.430 |
|---|
| xy | 12.915 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.336 |
-2.264 |
0.000 |
| y |
-2.264 |
6.157 |
0.000 |
| z |
0.000 |
0.000 |
4.624 |
<r2> (average value of r
2) Å
2
| <r2> |
210.842 |
| (<r2>)1/2 |
14.520 |