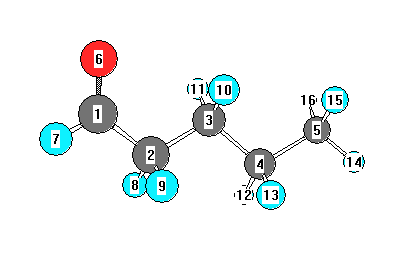
Vibrational Frequencies calculated at PBE1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3151 |
2994 |
32.74 |
|
|
|
| 2 |
A' |
3079 |
2926 |
30.87 |
|
|
|
| 3 |
A' |
3068 |
2915 |
20.95 |
|
|
|
| 4 |
A' |
3051 |
2899 |
26.71 |
|
|
|
| 5 |
A' |
3038 |
2887 |
18.59 |
|
|
|
| 6 |
A' |
2926 |
2780 |
156.88 |
|
|
|
| 7 |
A' |
1868 |
1775 |
144.27 |
|
|
|
| 8 |
A' |
1536 |
1459 |
7.41 |
|
|
|
| 9 |
A' |
1520 |
1445 |
1.00 |
|
|
|
| 10 |
A' |
1511 |
1436 |
1.43 |
|
|
|
| 11 |
A' |
1476 |
1403 |
15.08 |
|
|
|
| 12 |
A' |
1440 |
1368 |
11.61 |
|
|
|
| 13 |
A' |
1438 |
1366 |
1.50 |
|
|
|
| 14 |
A' |
1432 |
1361 |
7.92 |
|
|
|
| 15 |
A' |
1387 |
1318 |
14.91 |
|
|
|
| 16 |
A' |
1292 |
1227 |
4.09 |
|
|
|
| 17 |
A' |
1155 |
1098 |
11.78 |
|
|
|
| 18 |
A' |
1100 |
1045 |
0.58 |
|
|
|
| 19 |
A' |
1065 |
1012 |
1.00 |
|
|
|
| 20 |
A' |
938 |
892 |
1.23 |
|
|
|
| 21 |
A' |
913 |
867 |
12.95 |
|
|
|
| 22 |
A' |
701 |
666 |
12.92 |
|
|
|
| 23 |
A' |
398 |
378 |
1.77 |
|
|
|
| 24 |
A' |
293 |
278 |
4.55 |
|
|
|
| 25 |
A' |
137 |
130 |
5.05 |
|
|
|
| 26 |
A" |
3145 |
2988 |
52.05 |
|
|
|
| 27 |
A" |
3124 |
2969 |
21.68 |
|
|
|
| 28 |
A" |
3084 |
2931 |
7.66 |
|
|
|
| 29 |
A" |
3069 |
2916 |
7.86 |
|
|
|
| 30 |
A" |
1525 |
1449 |
8.04 |
|
|
|
| 31 |
A" |
1344 |
1277 |
0.21 |
|
|
|
| 32 |
A" |
1324 |
1258 |
0.08 |
|
|
|
| 33 |
A" |
1249 |
1187 |
0.00 |
|
|
|
| 34 |
A" |
1169 |
1111 |
0.03 |
|
|
|
| 35 |
A" |
987 |
938 |
1.39 |
|
|
|
| 36 |
A" |
862 |
819 |
0.12 |
|
|
|
| 37 |
A" |
750 |
713 |
2.69 |
|
|
|
| 38 |
A" |
674 |
640 |
3.95 |
|
|
|
| 39 |
A" |
240 |
228 |
0.00 |
|
|
|
| 40 |
A" |
184 |
174 |
0.89 |
|
|
|
| 41 |
A" |
111 |
106 |
1.04 |
|
|
|
| 42 |
A" |
68 |
65 |
1.87 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31408.5 cm
-1
Scaled (by 0.9503) Zero Point Vibrational Energy (zpe) 29847.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.242 |
|
|
|
| 2 |
C |
-0.427 |
|
|
|
| 3 |
C |
-0.309 |
|
|
|
| 4 |
C |
-0.321 |
|
|
|
| 5 |
C |
-0.520 |
|
|
|
| 6 |
O |
-0.385 |
|
|
|
| 7 |
H |
0.130 |
|
|
|
| 8 |
H |
0.193 |
|
|
|
| 9 |
H |
0.193 |
|
|
|
| 10 |
H |
0.181 |
|
|
|
| 11 |
H |
0.181 |
|
|
|
| 12 |
H |
0.162 |
|
|
|
| 13 |
H |
0.162 |
|
|
|
| 14 |
H |
0.173 |
|
|
|
| 15 |
H |
0.173 |
|
|
|
| 16 |
H |
0.173 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.568 |
-0.107 |
0.000 |
2.571 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.185 |
3.997 |
0.000 |
| y |
3.997 |
-36.390 |
0.000 |
| z |
0.000 |
0.000 |
-36.824 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.578 |
3.997 |
0.000 |
| y |
3.997 |
4.114 |
0.000 |
| z |
0.000 |
0.000 |
3.463 |
|
| Polar |
|---|
| 3z2-r2 | 6.926 |
|---|
| x2-y2 | -7.795 |
|---|
| xy | 3.997 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.484 |
-0.434 |
0.000 |
| y |
-0.434 |
9.385 |
0.000 |
| z |
0.000 |
0.000 |
6.628 |
<r2> (average value of r
2) Å
2
| <r2> |
250.094 |
| (<r2>)1/2 |
15.814 |