return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10O (Pentanal)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-271.447980
Energy at 298.15K-271.458794
HF Energy-271.174548
Nuclear repulsion energy230.664350
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3153 2993 37.07      
2 A' 3086 2929 32.50      
3 A' 3076 2920 19.69      
4 A' 3059 2904 28.50      
5 A' 3046 2891 17.45      
6 A' 2942 2792 156.66      
7 A' 1810 1718 117.79      
8 A' 1560 1481 5.20      
9 A' 1545 1466 0.52      
10 A' 1536 1458 1.43      
11 A' 1503 1426 13.59      
12 A' 1461 1387 2.40      
13 A' 1451 1378 5.62      
14 A' 1442 1369 5.63      
15 A' 1402 1330 15.16      
16 A' 1309 1243 2.68      
17 A' 1159 1101 10.62      
18 A' 1094 1039 1.07      
19 A' 1061 1007 2.04      
20 A' 939 891 0.68      
21 A' 912 866 12.43      
22 A' 698 662 12.49      
23 A' 401 380 1.91      
24 A' 297 282 4.50      
25 A' 141 134 4.90      
26 A" 3150 2990 59.20      
27 A" 3129 2970 24.13      
28 A" 3091 2934 9.27      
29 A" 3075 2919 9.30      
30 A" 1549 1470 6.23      
31 A" 1360 1291 0.21      
32 A" 1341 1273 0.09      
33 A" 1268 1204 0.01      
34 A" 1184 1124 0.03      
35 A" 997 946 1.15      
36 A" 874 829 0.03      
37 A" 761 723 1.80      
38 A" 688 653 2.74      
39 A" 247 235 0.00      
40 A" 177 168 0.97      
41 A" 112 107 0.98      
42 A" 72 68 1.99      

Unscaled Zero Point Vibrational Energy (zpe) 31576.8 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 29972.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.34292 0.04888 0.04417

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.826 -2.064 0.000
C2 -0.368 -1.134 0.000
C3 0.000 0.347 0.000
C4 -1.227 1.263 0.000
C5 -0.855 2.748 0.000
O6 1.983 -1.703 0.000
H7 0.566 -3.146 0.000
H8 -0.986 -1.392 0.873
H9 -0.986 -1.392 -0.873
H10 0.624 0.562 -0.875
H11 0.624 0.562 0.875
H12 -1.847 1.042 0.878
H13 -1.847 1.042 -0.878
H14 -1.747 3.381 0.000
H15 -0.262 3.003 -0.884
H16 -0.262 3.003 0.884

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 O6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
C11.51312.54843.90895.09621.21231.11332.12012.12012.77462.77464.18974.18976.02165.25715.2571
C21.51311.52692.54663.91232.41952.21791.09961.09962.15092.15092.77312.77314.72094.23244.2324
C32.54841.52691.53082.54792.85273.53882.18142.18141.09541.09542.15912.15913.50052.81142.8114
C43.90892.54661.53081.53024.37074.75952.80512.80512.16342.16341.09711.09712.18062.17712.1771
C55.09623.91232.54791.53025.27866.06234.23244.23242.78002.78002.15962.15961.09391.09451.0945
O61.21232.41952.85274.37075.27862.02283.11043.11042.78272.78274.79294.79296.30565.28895.2889
H71.11332.21793.53884.75956.06232.02282.49912.49913.81003.81004.91174.91176.92446.26746.2674
H82.12011.09962.18142.80514.23243.11042.49911.74543.07582.53132.58123.11874.91114.78824.4545
H92.12011.09962.18142.80514.23243.11042.49911.74542.53133.07583.11872.58124.91114.45454.7882
H102.77462.15091.09542.16342.78002.78273.81003.07582.53131.74913.06632.51643.78562.59723.1363
H112.77462.15091.09542.16342.78002.78273.81002.53133.07581.74912.51643.06633.78563.13632.5972
H124.18972.77312.15911.09712.15964.79294.91172.58123.11873.06632.51641.75532.50043.07602.5219
H134.18972.77312.15911.09712.15964.79294.91173.11872.58122.51643.06631.75532.50042.52193.0760
H146.02164.72093.50052.18061.09396.30566.92444.91114.91113.78563.78562.50042.50041.76881.7688
H155.25714.23242.81142.17711.09455.28896.26744.78824.45452.59723.13633.07602.52191.76881.7672
H165.25714.23242.81142.17711.09455.28896.26744.45454.78823.13632.59722.52193.07601.76881.7672

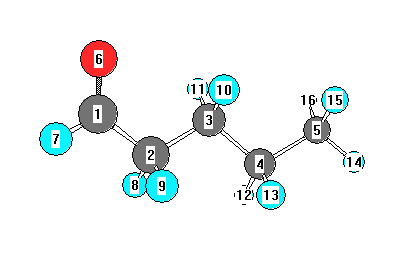
picture of Pentanal state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 113.922 C1 C2 H8 107.422
C1 C2 H9 107.422 C2 C1 O6 124.823
C2 C1 H7 114.369 C2 C3 C4 112.783
C2 C3 H10 109.112 C2 C3 H11 109.112
C3 C2 H8 111.267 C3 C2 H9 111.267
C3 C4 C5 112.685 C3 C4 H12 109.386
C3 C4 H13 109.386 C4 C3 H10 109.823
C4 C3 H11 109.823 C4 C5 H14 111.314
C4 C5 H15 111.000 C4 C5 H16 111.000
C5 C4 H12 109.462 C5 C4 H13 109.462
O6 C1 H7 120.808 H8 C2 H9 105.053
H10 C3 H11 105.955 H12 C4 H13 106.257
H14 C5 H15 107.848 H14 C5 H16 107.848
H15 C5 H16 107.672
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C 0.286      
2 C -0.385      
3 C -0.266      
4 C -0.277      
5 C -0.460      
6 O -0.431      
7 H 0.127      
8 H 0.172      
9 H 0.172      
10 H 0.162      
11 H 0.162      
12 H 0.141      
13 H 0.141      
14 H 0.152      
15 H 0.153      
16 H 0.153      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 8.408 -0.406 0.000
y -0.406 9.246 0.000
z 0.000 0.000 6.581


<r2> (average value of r2) Å2
<r2> 251.489
(<r2>)1/2 15.858