Jump to
S1C2
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -308.517286 |
| Energy at 298.15K | -308.528952 |
| HF Energy | -308.228955 |
| Nuclear repulsion energy | 244.654545 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3779 |
3587 |
0.00 |
|
|
|
| 2 |
Ag |
3074 |
2918 |
0.00 |
|
|
|
| 3 |
Ag |
3020 |
2867 |
0.00 |
|
|
|
| 4 |
Ag |
1578 |
1498 |
0.00 |
|
|
|
| 5 |
Ag |
1544 |
1465 |
0.00 |
|
|
|
| 6 |
Ag |
1499 |
1423 |
0.00 |
|
|
|
| 7 |
Ag |
1416 |
1344 |
0.00 |
|
|
|
| 8 |
Ag |
1293 |
1227 |
0.00 |
|
|
|
| 9 |
Ag |
1101 |
1045 |
0.00 |
|
|
|
| 10 |
Ag |
1080 |
1025 |
0.00 |
|
|
|
| 11 |
Ag |
1053 |
1000 |
0.00 |
|
|
|
| 12 |
Ag |
366 |
347 |
0.00 |
|
|
|
| 13 |
Ag |
342 |
324 |
0.00 |
|
|
|
| 14 |
Au |
3135 |
2975 |
84.76 |
|
|
|
| 15 |
Au |
3051 |
2896 |
107.66 |
|
|
|
| 16 |
Au |
1353 |
1284 |
2.35 |
|
|
|
| 17 |
Au |
1241 |
1178 |
3.52 |
|
|
|
| 18 |
Au |
968 |
919 |
2.79 |
|
|
|
| 19 |
Au |
772 |
733 |
0.27 |
|
|
|
| 20 |
Au |
287 |
272 |
254.72 |
|
|
|
| 21 |
Au |
98 |
93 |
19.30 |
|
|
|
| 22 |
Au |
83 |
78 |
2.99 |
|
|
|
| 23 |
Bg |
3112 |
2954 |
0.00 |
|
|
|
| 24 |
Bg |
3052 |
2897 |
0.00 |
|
|
|
| 25 |
Bg |
1345 |
1277 |
0.00 |
|
|
|
| 26 |
Bg |
1314 |
1248 |
0.00 |
|
|
|
| 27 |
Bg |
1207 |
1145 |
0.00 |
|
|
|
| 28 |
Bg |
835 |
792 |
0.00 |
|
|
|
| 29 |
Bg |
283 |
268 |
0.00 |
|
|
|
| 30 |
Bg |
159 |
151 |
0.00 |
|
|
|
| 31 |
Bu |
3779 |
3587 |
30.40 |
|
|
|
| 32 |
Bu |
3082 |
2925 |
61.39 |
|
|
|
| 33 |
Bu |
3020 |
2867 |
105.54 |
|
|
|
| 34 |
Bu |
1582 |
1501 |
6.30 |
|
|
|
| 35 |
Bu |
1553 |
1474 |
3.69 |
|
|
|
| 36 |
Bu |
1498 |
1422 |
20.61 |
|
|
|
| 37 |
Bu |
1337 |
1269 |
82.89 |
|
|
|
| 38 |
Bu |
1231 |
1169 |
54.82 |
|
|
|
| 39 |
Bu |
1095 |
1039 |
192.39 |
|
|
|
| 40 |
Bu |
1012 |
961 |
0.89 |
|
|
|
| 41 |
Bu |
527 |
500 |
40.13 |
|
|
|
| 42 |
Bu |
141 |
134 |
6.52 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31646.1 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 30038.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
1.490 |
2.682 |
0.000 |
| O2 |
-1.490 |
-2.682 |
0.000 |
| C3 |
1.490 |
1.248 |
0.000 |
| C4 |
-1.490 |
-1.248 |
0.000 |
| C5 |
0.047 |
0.770 |
0.000 |
| C6 |
-0.047 |
-0.770 |
0.000 |
| H7 |
2.412 |
2.982 |
0.000 |
| H8 |
-2.412 |
-2.982 |
0.000 |
| H9 |
-0.449 |
1.180 |
0.885 |
| H10 |
-0.449 |
1.180 |
-0.885 |
| H11 |
0.449 |
-1.180 |
0.885 |
| H12 |
0.449 |
-1.180 |
-0.885 |
| H13 |
-1.998 |
-0.851 |
-0.889 |
| H14 |
-1.998 |
-0.851 |
0.889 |
| H15 |
1.998 |
0.851 |
-0.889 |
| H16 |
1.998 |
0.851 |
0.889 |
Atom - Atom Distances (Å)
| |
O1 |
O2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
| O1 | | 6.1362 | 1.4343 | 4.9317 | 2.3959 | 3.7782 | 0.9694 | 6.8778 | 2.6076 | 2.6076 | 4.0963 | 4.0963 | 5.0439 | 5.0439 | 2.0980 | 2.0980 |
O2 | 6.1362 | | 4.9317 | 1.4343 | 3.7782 | 2.3959 | 6.8778 | 0.9694 | 4.0963 | 4.0963 | 2.6076 | 2.6076 | 2.0980 | 2.0980 | 5.0439 | 5.0439 | C3 | 1.4343 | 4.9317 | | 3.8866 | 1.5203 | 2.5358 | 1.9641 | 5.7544 | 2.1324 | 2.1324 | 2.7854 | 2.7854 | 4.1668 | 4.1668 | 1.0984 | 1.0984 | C4 | 4.9317 | 1.4343 | 3.8866 | | 2.5358 | 1.5203 | 5.7544 | 1.9641 | 2.7854 | 2.7854 | 2.1324 | 2.1324 | 1.0984 | 1.0984 | 4.1668 | 4.1668 | C5 | 2.3959 | 3.7782 | 1.5203 | 2.5358 | | 1.5421 | 3.2385 | 4.4853 | 1.0940 | 1.0940 | 2.1784 | 2.1784 | 2.7566 | 2.7566 | 2.1462 | 2.1462 | C6 | 3.7782 | 2.3959 | 2.5358 | 1.5203 | 1.5421 | | 4.4853 | 3.2385 | 2.1784 | 2.1784 | 1.0940 | 1.0940 | 2.1462 | 2.1462 | 2.7566 | 2.7566 | H7 | 0.9694 | 6.8778 | 1.9641 | 5.7544 | 3.2385 | 4.4853 | | 7.6704 | 3.4951 | 3.4951 | 4.6856 | 4.6856 | 5.9102 | 5.9102 | 2.3458 | 2.3458 | H8 | 6.8778 | 0.9694 | 5.7544 | 1.9641 | 4.4853 | 3.2385 | 7.6704 | | 4.6856 | 4.6856 | 3.4951 | 3.4951 | 2.3458 | 2.3458 | 5.9102 | 5.9102 | H9 | 2.6076 | 4.0963 | 2.1324 | 2.7854 | 1.0940 | 2.1784 | 3.4951 | 4.6856 | | 1.7695 | 2.5249 | 3.0832 | 3.1097 | 2.5542 | 3.0406 | 2.4696 | H10 | 2.6076 | 4.0963 | 2.1324 | 2.7854 | 1.0940 | 2.1784 | 3.4951 | 4.6856 | 1.7695 | | 3.0832 | 2.5249 | 2.5542 | 3.1097 | 2.4696 | 3.0406 | H11 | 4.0963 | 2.6076 | 2.7854 | 2.1324 | 2.1784 | 1.0940 | 4.6856 | 3.4951 | 2.5249 | 3.0832 | | 1.7695 | 3.0406 | 2.4696 | 3.1097 | 2.5542 | H12 | 4.0963 | 2.6076 | 2.7854 | 2.1324 | 2.1784 | 1.0940 | 4.6856 | 3.4951 | 3.0832 | 2.5249 | 1.7695 | | 2.4696 | 3.0406 | 2.5542 | 3.1097 | H13 | 5.0439 | 2.0980 | 4.1668 | 1.0984 | 2.7566 | 2.1462 | 5.9102 | 2.3458 | 3.1097 | 2.5542 | 3.0406 | 2.4696 | | 1.7781 | 4.3440 | 4.6938 | H14 | 5.0439 | 2.0980 | 4.1668 | 1.0984 | 2.7566 | 2.1462 | 5.9102 | 2.3458 | 2.5542 | 3.1097 | 2.4696 | 3.0406 | 1.7781 | | 4.6938 | 4.3440 | H15 | 2.0980 | 5.0439 | 1.0984 | 4.1668 | 2.1462 | 2.7566 | 2.3458 | 5.9102 | 3.0406 | 2.4696 | 3.1097 | 2.5542 | 4.3440 | 4.6938 | | 1.7781 | H16 | 2.0980 | 5.0439 | 1.0984 | 4.1668 | 2.1462 | 2.7566 | 2.3458 | 5.9102 | 2.4696 | 3.0406 | 2.5542 | 3.1097 | 4.6938 | 4.3440 | 1.7781 | |
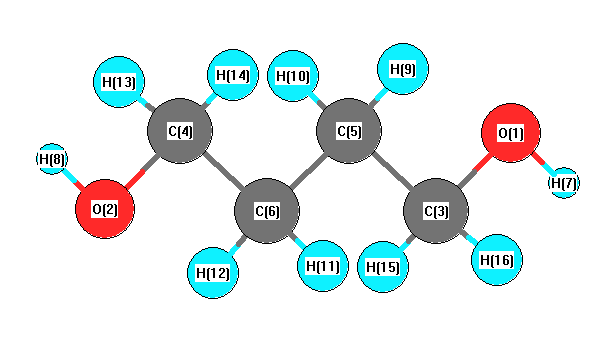 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C3 |
C5 |
108.331 |
|
O1 |
C3 |
H15 |
111.173 |
| O1 |
C3 |
H16 |
111.173 |
|
O2 |
C4 |
C6 |
108.331 |
| O2 |
C4 |
H13 |
111.173 |
|
O2 |
C4 |
H14 |
111.173 |
| C3 |
O1 |
H7 |
108.031 |
|
C3 |
C5 |
C6 |
111.797 |
| C3 |
C5 |
H9 |
108.200 |
|
C3 |
C5 |
H10 |
108.200 |
| C4 |
O2 |
H8 |
108.031 |
|
C4 |
C6 |
C5 |
111.797 |
| C4 |
C6 |
H11 |
108.200 |
|
C4 |
C6 |
H12 |
108.200 |
| C5 |
C3 |
H15 |
109.020 |
|
C5 |
C3 |
H16 |
109.020 |
| C5 |
C6 |
H11 |
110.293 |
|
C5 |
C6 |
H12 |
110.293 |
| C6 |
C4 |
H13 |
109.020 |
|
C6 |
C4 |
H14 |
109.020 |
| C6 |
C5 |
H9 |
110.293 |
|
C6 |
C5 |
H10 |
110.293 |
| H9 |
C5 |
H10 |
107.940 |
|
H11 |
C6 |
H12 |
107.940 |
| H13 |
C4 |
H14 |
108.082 |
|
H15 |
C3 |
H16 |
108.082 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.675 |
|
|
|
| 2 |
O |
-0.675 |
|
|
|
| 3 |
C |
-0.010 |
|
|
|
| 4 |
C |
-0.010 |
|
|
|
| 5 |
C |
-0.298 |
|
|
|
| 6 |
C |
-0.298 |
|
|
|
| 7 |
H |
0.410 |
|
|
|
| 8 |
H |
0.410 |
|
|
|
| 9 |
H |
0.157 |
|
|
|
| 10 |
H |
0.157 |
|
|
|
| 11 |
H |
0.157 |
|
|
|
| 12 |
H |
0.157 |
|
|
|
| 13 |
H |
0.129 |
|
|
|
| 14 |
H |
0.129 |
|
|
|
| 15 |
H |
0.129 |
|
|
|
| 16 |
H |
0.129 |
|
|
|
Electric dipole moments
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.809 |
1.107 |
0.000 |
| y |
1.107 |
8.147 |
0.000 |
| z |
0.000 |
0.000 |
6.315 |
<r2> (average value of r
2) Å
2
| <r2> |
284.994 |
| (<r2>)1/2 |
16.882 |
Jump to
S1C1
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -308.522437 |
| Energy at 298.15K | |
| HF Energy | -308.231717 |
| Nuclear repulsion energy | 265.626170 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3771 |
3580 |
24.78 |
|
|
|
| 2 |
A |
3686 |
3499 |
148.90 |
|
|
|
| 3 |
A |
3131 |
2972 |
67.62 |
|
|
|
| 4 |
A |
3121 |
2963 |
32.75 |
|
|
|
| 5 |
A |
3107 |
2950 |
5.45 |
|
|
|
| 6 |
A |
3083 |
2926 |
33.45 |
|
|
|
| 7 |
A |
3076 |
2920 |
77.26 |
|
|
|
| 8 |
A |
3070 |
2914 |
11.98 |
|
|
|
| 9 |
A |
3060 |
2904 |
38.17 |
|
|
|
| 10 |
A |
3038 |
2883 |
81.32 |
|
|
|
| 11 |
A |
1578 |
1498 |
0.67 |
|
|
|
| 12 |
A |
1570 |
1490 |
1.43 |
|
|
|
| 13 |
A |
1541 |
1462 |
3.41 |
|
|
|
| 14 |
A |
1529 |
1452 |
14.21 |
|
|
|
| 15 |
A |
1501 |
1425 |
61.26 |
|
|
|
| 16 |
A |
1489 |
1413 |
14.69 |
|
|
|
| 17 |
A |
1432 |
1359 |
0.75 |
|
|
|
| 18 |
A |
1419 |
1347 |
6.72 |
|
|
|
| 19 |
A |
1407 |
1336 |
4.48 |
|
|
|
| 20 |
A |
1353 |
1285 |
15.86 |
|
|
|
| 21 |
A |
1317 |
1250 |
2.55 |
|
|
|
| 22 |
A |
1288 |
1223 |
38.44 |
|
|
|
| 23 |
A |
1271 |
1206 |
2.44 |
|
|
|
| 24 |
A |
1206 |
1145 |
8.24 |
|
|
|
| 25 |
A |
1164 |
1105 |
7.17 |
|
|
|
| 26 |
A |
1134 |
1076 |
57.85 |
|
|
|
| 27 |
A |
1116 |
1059 |
19.09 |
|
|
|
| 28 |
A |
1054 |
1000 |
15.34 |
|
|
|
| 29 |
A |
1017 |
965 |
74.28 |
|
|
|
| 30 |
A |
957 |
909 |
2.71 |
|
|
|
| 31 |
A |
908 |
862 |
3.31 |
|
|
|
| 32 |
A |
817 |
775 |
1.29 |
|
|
|
| 33 |
A |
812 |
770 |
1.78 |
|
|
|
| 34 |
A |
565 |
537 |
22.18 |
|
|
|
| 35 |
A |
528 |
501 |
91.94 |
|
|
|
| 36 |
A |
453 |
430 |
1.83 |
|
|
|
| 37 |
A |
377 |
358 |
154.27 |
|
|
|
| 38 |
A |
335 |
318 |
3.31 |
|
|
|
| 39 |
A |
260 |
247 |
2.47 |
|
|
|
| 40 |
A |
230 |
218 |
38.62 |
|
|
|
| 41 |
A |
182 |
173 |
5.39 |
|
|
|
| 42 |
A |
84 |
80 |
10.66 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32017.7 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 30391.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.434 |
-0.017 |
0.525 |
| C2 |
-0.726 |
1.251 |
0.062 |
| C3 |
0.676 |
1.033 |
-0.527 |
| C4 |
1.590 |
0.132 |
0.308 |
| O5 |
-1.336 |
-1.013 |
-0.498 |
| O6 |
1.400 |
-1.251 |
0.052 |
| H7 |
-0.986 |
-0.379 |
1.463 |
| H8 |
-2.488 |
0.210 |
0.736 |
| H9 |
-0.664 |
1.924 |
0.928 |
| H10 |
-1.357 |
1.762 |
-0.676 |
| H11 |
0.598 |
0.601 |
-1.531 |
| H12 |
1.151 |
2.014 |
-0.655 |
| H13 |
1.439 |
0.361 |
1.377 |
| H14 |
2.640 |
0.353 |
0.089 |
| H15 |
-1.935 |
-1.740 |
-0.288 |
| H16 |
0.447 |
-1.397 |
-0.083 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
O5 |
O6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
| C1 | | 1.5244 | 2.5805 | 3.0347 | 1.4311 | 3.1272 | 1.1009 | 1.0986 | 2.1264 | 2.1476 | 2.9562 | 3.4929 | 3.0197 | 4.1140 | 1.9700 | 2.4108 |
C2 | 1.5244 | | 1.5358 | 2.5834 | 2.4106 | 3.2834 | 2.1656 | 2.1546 | 1.0985 | 1.0974 | 2.1713 | 2.1492 | 2.6846 | 3.4839 | 3.2451 | 2.8999 | C3 | 2.5805 | 1.5358 | | 1.5310 | 2.8690 | 2.4649 | 2.9524 | 3.5041 | 2.1692 | 2.1651 | 1.0959 | 1.0980 | 2.1587 | 2.1682 | 3.8160 | 2.4808 | C4 | 3.0347 | 2.5834 | 1.5310 | | 3.2426 | 1.4186 | 2.8687 | 4.1005 | 2.9452 | 3.5086 | 2.1409 | 2.1597 | 1.1043 | 1.0958 | 4.0349 | 1.9479 | O5 | 1.4311 | 2.4106 | 2.8690 | 3.2426 | | 2.8007 | 2.0901 | 2.0843 | 3.3330 | 2.7806 | 2.7224 | 3.9207 | 3.6190 | 4.2446 | 0.9654 | 1.8702 | O6 | 3.1272 | 3.2834 | 2.4649 | 1.4186 | 2.8007 | | 2.9062 | 4.2096 | 3.8868 | 4.1488 | 2.5646 | 3.3502 | 2.0869 | 2.0276 | 3.3884 | 0.9738 | H7 | 1.1009 | 2.1656 | 2.9524 | 2.8687 | 2.0901 | 2.9062 | | 1.7701 | 2.3864 | 3.0493 | 3.5265 | 3.8450 | 2.5363 | 3.9464 | 2.4122 | 2.3407 | H8 | 1.0986 | 2.1546 | 3.5041 | 4.1005 | 2.0843 | 4.2096 | 1.7701 | | 2.5102 | 2.3826 | 3.8490 | 4.2930 | 3.9813 | 5.1707 | 2.2705 | 3.4448 | H9 | 2.1264 | 1.0985 | 2.1692 | 2.9452 | 3.3330 | 3.8868 | 2.3864 | 2.5102 | | 1.7551 | 3.0646 | 2.4100 | 2.6578 | 3.7533 | 4.0642 | 3.6447 | H10 | 2.1476 | 1.0974 | 2.1651 | 3.5086 | 2.7806 | 4.1488 | 3.0493 | 2.3826 | 1.7551 | | 2.4302 | 2.5213 | 3.7412 | 4.3072 | 3.5702 | 3.6862 | H11 | 2.9562 | 2.1713 | 1.0959 | 2.1409 | 2.7224 | 2.5646 | 3.5265 | 3.8490 | 3.0646 | 2.4302 | | 1.7526 | 3.0368 | 2.6183 | 3.6663 | 2.4722 | H12 | 3.4929 | 2.1492 | 1.0980 | 2.1597 | 3.9207 | 3.3502 | 3.8450 | 4.2930 | 2.4100 | 2.5213 | 1.7526 | | 2.6359 | 2.3520 | 4.8737 | 3.5301 | H13 | 3.0197 | 2.6846 | 2.1587 | 1.1043 | 3.6190 | 2.0869 | 2.5363 | 3.9813 | 2.6578 | 3.7412 | 3.0368 | 2.6359 | | 1.7617 | 4.3090 | 2.4908 | H14 | 4.1140 | 3.4839 | 2.1682 | 1.0958 | 4.2446 | 2.0276 | 3.9464 | 5.1707 | 3.7533 | 4.3072 | 2.6183 | 2.3520 | 1.7617 | | 5.0455 | 2.8112 | H15 | 1.9700 | 3.2451 | 3.8160 | 4.0349 | 0.9654 | 3.3884 | 2.4122 | 2.2705 | 4.0642 | 3.5702 | 3.6663 | 4.8737 | 4.3090 | 5.0455 | | 2.4156 | H16 | 2.4108 | 2.8999 | 2.4808 | 1.9479 | 1.8702 | 0.9738 | 2.3407 | 3.4448 | 3.6447 | 3.6862 | 2.4722 | 3.5301 | 2.4908 | 2.8112 | 2.4156 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C3 |
C5 |
29.853 |
|
O1 |
C3 |
H15 |
28.305 |
| O1 |
C3 |
H16 |
56.850 |
|
O2 |
C4 |
C6 |
106.602 |
| O2 |
C4 |
H13 |
83.087 |
|
O2 |
C4 |
H14 |
138.830 |
| C3 |
O1 |
H7 |
98.559 |
|
C3 |
C5 |
C6 |
51.522 |
| C3 |
C5 |
H9 |
40.074 |
|
C3 |
C5 |
H10 |
45.035 |
| C4 |
O2 |
H8 |
119.595 |
|
C4 |
C6 |
C5 |
94.748 |
| C4 |
C6 |
H11 |
56.592 |
|
C4 |
C6 |
H12 |
25.597 |
| C5 |
C3 |
H15 |
3.248 |
|
C5 |
C3 |
H16 |
40.104 |
| C5 |
C6 |
H11 |
60.792 |
|
C5 |
C6 |
H12 |
78.640 |
| C6 |
C4 |
H13 |
111.013 |
|
C6 |
C4 |
H14 |
106.788 |
| C6 |
C5 |
H9 |
78.112 |
|
C6 |
C5 |
H10 |
96.033 |
| H9 |
C5 |
H10 |
31.759 |
|
H11 |
C6 |
H12 |
30.999 |
| H13 |
C4 |
H14 |
106.401 |
|
H15 |
C3 |
H16 |
38.190 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.051 |
|
|
|
| 2 |
C |
-0.287 |
|
|
|
| 3 |
C |
-0.339 |
|
|
|
| 4 |
C |
0.016 |
|
|
|
| 5 |
O |
-0.686 |
|
|
|
| 6 |
O |
-0.716 |
|
|
|
| 7 |
H |
0.168 |
|
|
|
| 8 |
H |
0.142 |
|
|
|
| 9 |
H |
0.148 |
|
|
|
| 10 |
H |
0.158 |
|
|
|
| 11 |
H |
0.169 |
|
|
|
| 12 |
H |
0.149 |
|
|
|
| 13 |
H |
0.132 |
|
|
|
| 14 |
H |
0.134 |
|
|
|
| 15 |
H |
0.421 |
|
|
|
| 16 |
H |
0.441 |
|
|
|
Electric dipole moments
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.044 |
0.324 |
-0.122 |
| y |
0.324 |
7.342 |
0.110 |
| z |
-0.122 |
0.110 |
6.626 |
<r2> (average value of r
2) Å
2
| <r2> |
177.833 |
| (<r2>)1/2 |
13.335 |
