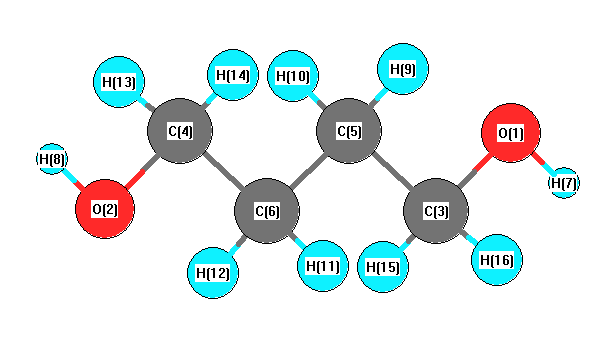
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3641 |
3581 |
0.00 |
|
|
|
| 2 |
Ag |
2970 |
2921 |
0.00 |
|
|
|
| 3 |
Ag |
2891 |
2844 |
0.00 |
|
|
|
| 4 |
Ag |
1507 |
1482 |
0.00 |
|
|
|
| 5 |
Ag |
1473 |
1449 |
0.00 |
|
|
|
| 6 |
Ag |
1433 |
1409 |
0.00 |
|
|
|
| 7 |
Ag |
1352 |
1330 |
0.00 |
|
|
|
| 8 |
Ag |
1249 |
1229 |
0.00 |
|
|
|
| 9 |
Ag |
1070 |
1052 |
0.00 |
|
|
|
| 10 |
Ag |
1049 |
1032 |
0.00 |
|
|
|
| 11 |
Ag |
1014 |
997 |
0.00 |
|
|
|
| 12 |
Ag |
352 |
346 |
0.00 |
|
|
|
| 13 |
Ag |
330 |
325 |
0.00 |
|
|
|
| 14 |
Au |
3030 |
2980 |
74.65 |
|
|
|
| 15 |
Au |
2916 |
2868 |
128.24 |
|
|
|
| 16 |
Au |
1293 |
1271 |
2.95 |
|
|
|
| 17 |
Au |
1187 |
1168 |
1.95 |
|
|
|
| 18 |
Au |
929 |
914 |
2.44 |
|
|
|
| 19 |
Au |
747 |
735 |
0.83 |
|
|
|
| 20 |
Au |
284 |
280 |
237.68 |
|
|
|
| 21 |
Au |
99 |
98 |
14.37 |
|
|
|
| 22 |
Au |
78 |
77 |
3.46 |
|
|
|
| 23 |
Bg |
3007 |
2957 |
0.00 |
|
|
|
| 24 |
Bg |
2917 |
2868 |
0.00 |
|
|
|
| 25 |
Bg |
1287 |
1266 |
0.00 |
|
|
|
| 26 |
Bg |
1255 |
1234 |
0.00 |
|
|
|
| 27 |
Bg |
1152 |
1133 |
0.00 |
|
|
|
| 28 |
Bg |
801 |
788 |
0.00 |
|
|
|
| 29 |
Bg |
282 |
277 |
0.00 |
|
|
|
| 30 |
Bg |
153 |
150 |
0.00 |
|
|
|
| 31 |
Bu |
3641 |
3581 |
12.76 |
|
|
|
| 32 |
Bu |
2979 |
2930 |
61.68 |
|
|
|
| 33 |
Bu |
2892 |
2844 |
126.08 |
|
|
|
| 34 |
Bu |
1511 |
1486 |
7.14 |
|
|
|
| 35 |
Bu |
1482 |
1458 |
3.51 |
|
|
|
| 36 |
Bu |
1433 |
1410 |
17.97 |
|
|
|
| 37 |
Bu |
1285 |
1264 |
89.21 |
|
|
|
| 38 |
Bu |
1187 |
1167 |
29.85 |
|
|
|
| 39 |
Bu |
1057 |
1039 |
197.33 |
|
|
|
| 40 |
Bu |
982 |
966 |
2.35 |
|
|
|
| 41 |
Bu |
509 |
500 |
38.13 |
|
|
|
| 42 |
Bu |
135 |
133 |
5.55 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30420.0 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 29918.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.585 |
|
|
|
| 2 |
O |
-0.585 |
|
|
|
| 3 |
C |
-0.077 |
|
|
|
| 4 |
C |
-0.077 |
|
|
|
| 5 |
C |
-0.299 |
|
|
|
| 6 |
C |
-0.299 |
|
|
|
| 7 |
H |
0.386 |
|
|
|
| 8 |
H |
0.386 |
|
|
|
| 9 |
H |
0.159 |
|
|
|
| 10 |
H |
0.159 |
|
|
|
| 11 |
H |
0.159 |
|
|
|
| 12 |
H |
0.159 |
|
|
|
| 13 |
H |
0.129 |
|
|
|
| 14 |
H |
0.129 |
|
|
|
| 15 |
H |
0.129 |
|
|
|
| 16 |
H |
0.129 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.442 |
6.779 |
0.000 |
| y |
6.779 |
-43.427 |
0.000 |
| z |
0.000 |
0.000 |
-38.198 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
14.371 |
6.779 |
0.000 |
| y |
6.779 |
-11.107 |
0.000 |
| z |
0.000 |
0.000 |
-3.264 |
|
| Polar |
|---|
| 3z2-r2 | -6.528 |
|---|
| x2-y2 | 16.985 |
|---|
| xy | 6.779 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.320 |
1.265 |
0.000 |
| y |
1.265 |
8.661 |
0.000 |
| z |
0.000 |
0.000 |
6.563 |
<r2> (average value of r
2) Å
2
| <r2> |
287.016 |
| (<r2>)1/2 |
16.942 |