return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H12S (1-Pentanethiol)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-595.517315
Energy at 298.15K-595.530244
HF Energy-595.260649
Nuclear repulsion energy286.736535
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3153 2992 37.93      
2 A' 3101 2944 36.64      
3 A' 3076 2920 36.80      
4 A' 3074 2917 29.73      
5 A' 3064 2909 17.96      
6 A' 3051 2896 8.88      
7 A' 2732 2594 26.67      
8 A' 1563 1483 4.35      
9 A' 1553 1474 0.07      
10 A' 1542 1464 1.79      
11 A' 1539 1461 2.07      
12 A' 1535 1457 0.49      
13 A' 1463 1389 2.16      
14 A' 1445 1372 0.07      
15 A' 1413 1341 13.34      
16 A' 1347 1278 11.77      
17 A' 1281 1216 19.98      
18 A' 1163 1104 3.76      
19 A' 1093 1037 0.14      
20 A' 1084 1029 0.67      
21 A' 1050 997 0.47      
22 A' 928 881 0.54      
23 A' 885 840 2.21      
24 A' 757 719 3.66      
25 A' 444 421 1.45      
26 A' 352 334 0.35      
27 A' 251 238 1.27      
28 A' 118 112 1.13      
29 A" 3157 2997 34.81      
30 A" 3148 2988 51.06      
31 A" 3119 2960 34.21      
32 A" 3103 2945 8.43      
33 A" 3082 2925 1.67      
34 A" 1552 1473 6.28      
35 A" 1364 1295 0.58      
36 A" 1363 1294 0.38      
37 A" 1320 1253 0.20      
38 A" 1257 1193 0.16      
39 A" 1114 1057 2.00      
40 A" 991 940 0.07      
41 A" 865 821 1.53      
42 A" 779 739 0.09      
43 A" 750 712 3.93      
44 A" 256 243 0.04      
45 A" 201 191 19.24      
46 A" 149 142 1.59      
47 A" 104 98 2.63      
48 A" 67 64 1.77      

Unscaled Zero Point Vibrational Energy (zpe) 35897.0 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 34073.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.47733 0.02657 0.02577

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 -2.582 1.291 0.000
H2 -3.607 0.844 0.000
C3 -1.470 -0.002 0.000
H4 -1.659 -0.613 0.907
H5 -1.659 -0.613 -0.907
C6 0.000 0.425 0.000
H7 0.194 1.039 -0.865
H8 0.194 1.039 0.865
C9 0.909 -0.804 0.000
H10 0.715 -1.411 0.886
H11 0.715 -1.411 -0.886
C12 2.382 -0.379 0.000
H13 2.588 0.211 0.892
H14 2.588 0.211 -0.892
C15 3.295 -1.607 0.000
H16 3.105 -2.222 0.865
H17 3.105 -2.222 -0.865
H18 4.341 -1.298 0.000

Atom - Atom Distances (Å)
  S1 H2 C3 H4 H5 C6 H7 H8 C9 H10 H11 C12 H13 H14 C15 H16 H17 H18
S11.11821.70562.30222.30222.72342.91832.91834.07114.35394.35395.23685.35655.35656.55266.73986.73987.3916
H21.11822.29872.59632.59633.63143.90293.90294.80734.95504.95506.11226.29116.29117.32447.42947.42948.2321
C31.70562.29871.10991.10991.53062.14472.14472.51012.74662.74663.86974.16054.16055.02785.15765.15765.9540
H42.30222.59631.10991.81412.15723.05012.48292.73032.50493.08054.14804.32694.68605.13375.02845.33136.1074
H52.30222.59631.10991.81412.15722.48293.05012.73033.08052.50494.14804.68604.32695.13375.33135.02846.1074
C62.72343.63141.53062.15722.15721.07831.07831.52832.16042.16042.51352.74592.74593.87114.17034.17034.6708
H72.91833.90292.14473.05012.48291.07831.73012.15753.05622.50482.74673.08312.53364.16724.70074.37094.8386
H82.91833.90292.14472.48293.05011.07831.73012.15752.50483.05622.74672.53363.08314.16724.37094.70074.8386
C94.07114.80732.51012.73032.73031.52832.15752.15751.09161.09161.53282.15562.15562.51762.75322.75323.4679
H104.35394.95502.74662.50493.08052.16043.05622.50481.09161.77222.15122.47823.05002.73482.52353.07133.7347
H114.35394.95502.74663.08052.50492.16042.50483.05621.09161.77222.15123.05002.47822.73483.07132.52353.7347
C125.23686.11223.86974.14804.14802.51352.74672.74671.53282.15122.15121.08931.08931.53052.16032.16032.1647
H135.35656.29114.16054.32694.68602.74593.08312.53362.15562.47823.05001.08931.78382.14502.48753.04512.4794
H145.35656.29114.16054.68604.32692.74592.53363.08312.15563.05002.47821.08931.78382.14503.04512.48752.4794
C156.55267.32445.02785.13375.13373.87114.16724.16722.51762.73482.73481.53052.14502.14501.07791.07791.0911
H166.73987.42945.15765.02845.33134.17034.70074.37092.75322.52353.07132.16032.48753.04511.07791.72941.7692
H176.73987.42945.15765.33135.02844.17034.37094.70072.75323.07132.52352.16033.04512.48751.07791.72941.7692
H187.39168.23215.95406.10746.10744.67084.83864.83863.46793.73473.73472.16472.47942.47941.09111.76921.7692

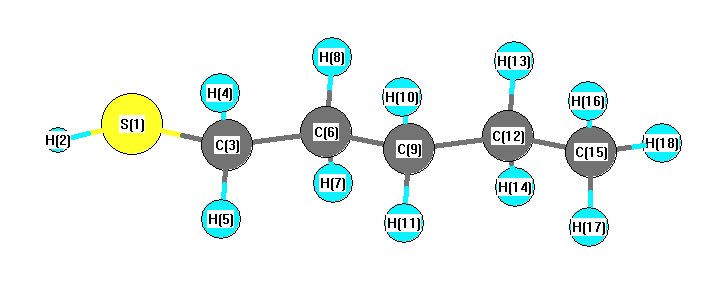
picture of 1-Pentanethiol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C3 H4 107.822 S1 C3 H5 107.822
S1 C3 C6 114.493 H2 S1 C3 107.147
C3 C6 H7 109.355 C3 C6 H8 109.355
C3 C6 C9 110.287 H4 C3 H5 109.615
H4 C3 C6 108.512 H5 C3 C6 108.512
C6 C9 H10 109.981 C6 C9 H11 109.981
C6 C9 C12 110.394 H7 C6 H8 106.690
H7 C6 C9 110.538 H8 C6 C9 110.538
C9 C12 H13 109.425 C9 C12 H14 109.425
C9 C12 C15 110.545 H10 C9 H11 108.535
H10 C9 C12 108.956 H11 C9 C12 108.956
C12 C15 H16 110.630 C12 C15 H17 110.630
C12 C15 H18 110.197 H13 C12 H14 109.927
H13 C12 C15 108.753 H14 C12 C15 108.753
H16 C15 H17 106.683 H16 C15 H18 109.312
H17 C15 H18 109.312
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 S -0.077      
2 H 0.088      
3 C -0.424      
4 H 0.181      
5 H 0.181      
6 C -0.293      
7 H 0.163      
8 H 0.163      
9 C -0.273      
10 H 0.142      
11 H 0.142      
12 C -0.273      
13 H 0.145      
14 H 0.145      
15 C -0.461      
16 H 0.150      
17 H 0.150      
18 H 0.154      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 12.218 -1.773 0.000
y -1.773 10.225 0.000
z 0.000 0.000 8.384


<r2> (average value of r2) Å2
<r2> 412.533
(<r2>)1/2 20.311