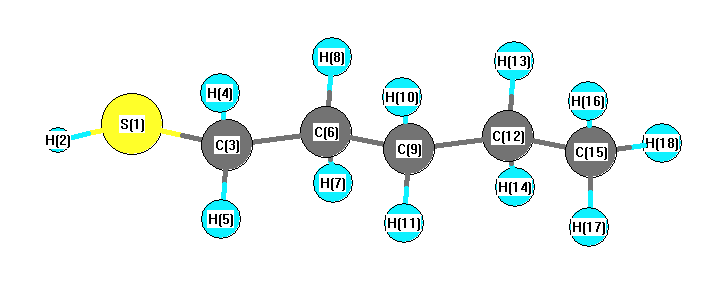
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3148 |
2994 |
34.93 |
|
|
|
| 2 |
A' |
3088 |
2937 |
37.69 |
|
|
|
| 3 |
A' |
3065 |
2915 |
41.19 |
|
|
|
| 4 |
A' |
3063 |
2913 |
27.16 |
|
|
|
| 5 |
A' |
3054 |
2905 |
17.24 |
|
|
|
| 6 |
A' |
3041 |
2892 |
8.06 |
|
|
|
| 7 |
A' |
2736 |
2602 |
25.41 |
|
|
|
| 8 |
A' |
1538 |
1463 |
6.77 |
|
|
|
| 9 |
A' |
1529 |
1454 |
0.28 |
|
|
|
| 10 |
A' |
1517 |
1443 |
2.08 |
|
|
|
| 11 |
A' |
1515 |
1440 |
1.53 |
|
|
|
| 12 |
A' |
1511 |
1437 |
0.44 |
|
|
|
| 13 |
A' |
1443 |
1372 |
4.00 |
|
|
|
| 14 |
A' |
1433 |
1363 |
0.24 |
|
|
|
| 15 |
A' |
1396 |
1328 |
11.77 |
|
|
|
| 16 |
A' |
1327 |
1262 |
12.20 |
|
|
|
| 17 |
A' |
1262 |
1200 |
16.85 |
|
|
|
| 18 |
A' |
1159 |
1102 |
4.44 |
|
|
|
| 19 |
A' |
1099 |
1045 |
0.15 |
|
|
|
| 20 |
A' |
1087 |
1034 |
0.92 |
|
|
|
| 21 |
A' |
1055 |
1003 |
0.61 |
|
|
|
| 22 |
A' |
927 |
882 |
0.89 |
|
|
|
| 23 |
A' |
881 |
838 |
2.28 |
|
|
|
| 24 |
A' |
764 |
727 |
3.64 |
|
|
|
| 25 |
A' |
441 |
420 |
1.39 |
|
|
|
| 26 |
A' |
351 |
334 |
0.38 |
|
|
|
| 27 |
A' |
249 |
237 |
1.31 |
|
|
|
| 28 |
A' |
117 |
112 |
1.20 |
|
|
|
| 29 |
A" |
3147 |
2992 |
35.27 |
|
|
|
| 30 |
A" |
3140 |
2986 |
45.21 |
|
|
|
| 31 |
A" |
3110 |
2958 |
32.22 |
|
|
|
| 32 |
A" |
3094 |
2943 |
6.96 |
|
|
|
| 33 |
A" |
3072 |
2922 |
1.55 |
|
|
|
| 34 |
A" |
1529 |
1454 |
7.87 |
|
|
|
| 35 |
A" |
1350 |
1283 |
0.30 |
|
|
|
| 36 |
A" |
1348 |
1282 |
0.68 |
|
|
|
| 37 |
A" |
1303 |
1239 |
0.28 |
|
|
|
| 38 |
A" |
1237 |
1176 |
0.13 |
|
|
|
| 39 |
A" |
1100 |
1046 |
2.29 |
|
|
|
| 40 |
A" |
977 |
929 |
0.05 |
|
|
|
| 41 |
A" |
853 |
811 |
2.04 |
|
|
|
| 42 |
A" |
769 |
731 |
0.14 |
|
|
|
| 43 |
A" |
743 |
706 |
5.30 |
|
|
|
| 44 |
A" |
256 |
243 |
0.04 |
|
|
|
| 45 |
A" |
203 |
193 |
20.80 |
|
|
|
| 46 |
A" |
151 |
144 |
1.44 |
|
|
|
| 47 |
A" |
102 |
97 |
2.53 |
|
|
|
| 48 |
A" |
67 |
64 |
1.91 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35673.0 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 33925.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
-0.087 |
|
|
|
| 2 |
H |
0.105 |
|
|
|
| 3 |
C |
-0.470 |
|
|
|
| 4 |
H |
0.199 |
|
|
|
| 5 |
H |
0.199 |
|
|
|
| 6 |
C |
-0.327 |
|
|
|
| 7 |
H |
0.181 |
|
|
|
| 8 |
H |
0.181 |
|
|
|
| 9 |
C |
-0.306 |
|
|
|
| 10 |
H |
0.159 |
|
|
|
| 11 |
H |
0.159 |
|
|
|
| 12 |
C |
-0.311 |
|
|
|
| 13 |
H |
0.163 |
|
|
|
| 14 |
H |
0.163 |
|
|
|
| 15 |
C |
-0.514 |
|
|
|
| 16 |
H |
0.168 |
|
|
|
| 17 |
H |
0.168 |
|
|
|
| 18 |
H |
0.172 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.309 |
-1.833 |
0.000 |
1.859 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.315 |
3.661 |
0.000 |
| y |
3.661 |
-51.561 |
0.000 |
| z |
0.000 |
0.000 |
-48.126 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.529 |
3.661 |
0.000 |
| y |
3.661 |
-5.341 |
0.000 |
| z |
0.000 |
0.000 |
-0.188 |
|
| Polar |
|---|
| 3z2-r2 | -0.376 |
|---|
| x2-y2 | 7.246 |
|---|
| xy | 3.661 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.378 |
-1.810 |
0.000 |
| y |
-1.810 |
10.356 |
0.000 |
| z |
0.000 |
0.000 |
8.461 |
<r2> (average value of r
2) Å
2
| <r2> |
409.668 |
| (<r2>)1/2 |
20.240 |