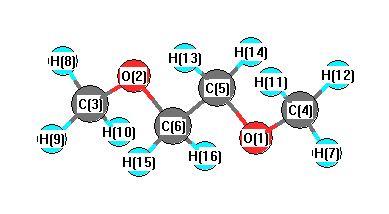
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3148 |
3011 |
0.00 |
|
|
|
| 2 |
Ag |
2999 |
2869 |
0.00 |
|
|
|
| 3 |
Ag |
2987 |
2858 |
0.00 |
|
|
|
| 4 |
Ag |
1556 |
1488 |
0.00 |
|
|
|
| 5 |
Ag |
1530 |
1464 |
0.00 |
|
|
|
| 6 |
Ag |
1505 |
1440 |
0.00 |
|
|
|
| 7 |
Ag |
1463 |
1400 |
0.00 |
|
|
|
| 8 |
Ag |
1255 |
1201 |
0.00 |
|
|
|
| 9 |
Ag |
1197 |
1145 |
0.00 |
|
|
|
| 10 |
Ag |
1097 |
1049 |
0.00 |
|
|
|
| 11 |
Ag |
1042 |
997 |
0.00 |
|
|
|
| 12 |
Ag |
401 |
383 |
0.00 |
|
|
|
| 13 |
Ag |
332 |
317 |
0.00 |
|
|
|
| 14 |
Au |
3045 |
2914 |
7.63 |
|
|
|
| 15 |
Au |
3042 |
2910 |
254.01 |
|
|
|
| 16 |
Au |
1511 |
1445 |
12.15 |
|
|
|
| 17 |
Au |
1250 |
1196 |
12.52 |
|
|
|
| 18 |
Au |
1189 |
1137 |
2.28 |
|
|
|
| 19 |
Au |
844 |
807 |
0.26 |
|
|
|
| 20 |
Au |
244 |
234 |
8.66 |
|
|
|
| 21 |
Au |
102 |
98 |
7.67 |
|
|
|
| 22 |
Au |
70 |
67 |
3.16 |
|
|
|
| 23 |
Bg |
3043 |
2911 |
0.00 |
|
|
|
| 24 |
Bg |
3018 |
2887 |
0.00 |
|
|
|
| 25 |
Bg |
1511 |
1445 |
0.00 |
|
|
|
| 26 |
Bg |
1308 |
1251 |
0.00 |
|
|
|
| 27 |
Bg |
1207 |
1154 |
0.00 |
|
|
|
| 28 |
Bg |
1165 |
1115 |
0.00 |
|
|
|
| 29 |
Bg |
228 |
218 |
0.00 |
|
|
|
| 30 |
Bg |
126 |
120 |
0.00 |
|
|
|
| 31 |
Bu |
3148 |
3011 |
65.23 |
|
|
|
| 32 |
Bu |
3001 |
2871 |
145.47 |
|
|
|
| 33 |
Bu |
2988 |
2859 |
82.16 |
|
|
|
| 34 |
Bu |
1562 |
1495 |
2.19 |
|
|
|
| 35 |
Bu |
1531 |
1465 |
25.25 |
|
|
|
| 36 |
Bu |
1497 |
1433 |
5.21 |
|
|
|
| 37 |
Bu |
1374 |
1315 |
33.36 |
|
|
|
| 38 |
Bu |
1239 |
1186 |
214.40 |
|
|
|
| 39 |
Bu |
1193 |
1141 |
173.76 |
|
|
|
| 40 |
Bu |
995 |
952 |
62.62 |
|
|
|
| 41 |
Bu |
506 |
484 |
3.96 |
|
|
|
| 42 |
Bu |
143 |
137 |
3.99 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31295.5 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 29940.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.466 |
|
|
-0.350 |
| 2 |
O |
-0.466 |
|
|
-0.347 |
| 3 |
C |
-0.251 |
|
|
-0.027 |
| 4 |
C |
-0.251 |
|
|
-0.036 |
| 5 |
C |
-0.072 |
|
|
0.115 |
| 6 |
C |
-0.072 |
|
|
0.085 |
| 7 |
H |
0.181 |
|
|
0.089 |
| 8 |
H |
0.181 |
|
|
0.087 |
| 9 |
H |
0.151 |
|
|
0.050 |
| 10 |
H |
0.151 |
|
|
0.050 |
| 11 |
H |
0.151 |
|
|
0.052 |
| 12 |
H |
0.151 |
|
|
0.052 |
| 13 |
H |
0.154 |
|
|
0.042 |
| 14 |
H |
0.154 |
|
|
0.042 |
| 15 |
H |
0.154 |
|
|
0.048 |
| 16 |
H |
0.154 |
|
|
0.048 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.005 |
-0.010 |
0.000 |
0.011 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.202 |
2.171 |
0.000 |
| y |
2.171 |
-29.836 |
0.000 |
| z |
0.000 |
0.000 |
-37.864 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.352 |
2.171 |
0.000 |
| y |
2.171 |
9.197 |
0.000 |
| z |
0.000 |
0.000 |
-2.846 |
|
| Polar |
|---|
| 3z2-r2 | -5.691 |
|---|
| x2-y2 | -10.366 |
|---|
| xy | 2.171 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
268.527 |
| (<r2>)1/2 |
16.387 |