return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H10O2 (Ethane, 1,2-dimethoxy-)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1Ag
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-308.505930
Energy at 298.15K-308.517629
HF Energy-308.218426
Nuclear repulsion energy252.630087
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3178 3017 0.00      
2 Ag 3034 2879 0.00      
3 Ag 3023 2869 0.00      
4 Ag 1579 1499 0.00      
5 Ag 1556 1477 0.00      
6 Ag 1526 1449 0.00      
7 Ag 1483 1407 0.00      
8 Ag 1265 1201 0.00      
9 Ag 1188 1127 0.00      
10 Ag 1106 1050 0.00      
11 Ag 1035 982 0.00      
12 Ag 402 382 0.00      
13 Ag 337 320 0.00      
14 Au 3082 2925 6.88      
15 Au 3079 2923 241.54      
16 Au 1537 1459 9.63      
17 Au 1261 1197 12.89      
18 Au 1201 1140 2.80      
19 Au 856 813 0.53      
20 Au 249 236 9.07      
21 Au 108 102 7.38      
22 Au 71 68 4.18      
23 Bg 3081 2924 0.00      
24 Bg 3055 2900 0.00      
25 Bg 1537 1459 0.00      
26 Bg 1324 1257 0.00      
27 Bg 1220 1158 0.00      
28 Bg 1180 1120 0.00      
29 Bg 233 221 0.00      
30 Bg 128 121 0.00      
31 Bu 3178 3017 64.61      
32 Bu 3035 2881 137.82      
33 Bu 3024 2871 69.27      
34 Bu 1587 1506 2.01      
35 Bu 1556 1477 20.87      
36 Bu 1518 1441 9.10      
37 Bu 1391 1320 36.97      
38 Bu 1245 1182 132.84      
39 Bu 1186 1126 247.61      
40 Bu 982 932 60.76      
41 Bu 513 487 4.39      
42 Bu 149 142 4.18      

Unscaled Zero Point Vibrational Energy (zpe) 31638.0 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 30030.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.62977 0.04305 0.04156

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 0.435 1.750 0.000
O2 -0.435 -1.750 0.000
C3 0.274 -2.986 0.000
C4 -0.274 2.986 0.000
C5 -0.435 0.622 0.000
C6 0.435 -0.622 0.000
H7 0.474 3.780 0.000
H8 -0.474 -3.780 0.000
H9 0.904 -3.087 0.892
H10 0.904 -3.087 -0.892
H11 -0.904 3.087 0.892
H12 -0.904 3.087 -0.892
H13 -1.077 0.621 0.892
H14 -1.077 0.621 -0.892
H15 1.077 -0.621 0.892
H16 1.077 -0.621 -0.892

Atom - Atom Distances (Å)
  O1 O2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
O13.60604.73821.42511.42462.37182.03115.60424.94034.94032.09222.09222.08712.08712.61232.6123
O23.60601.42514.73822.37181.42465.60422.03112.09222.09224.94034.94032.61232.61232.08712.0871
C34.73821.42515.99673.67712.36926.76911.09101.09671.09676.24986.24983.95283.95282.65222.6522
C41.42514.73825.99672.36923.67711.09106.76916.24986.24981.09671.09672.65222.65223.95283.9528
C51.42462.37183.67712.36921.51893.28644.40274.04314.04312.66262.66261.09821.09822.15072.1507
C62.37181.42462.36923.67711.51894.40273.28642.66262.66264.04314.04312.15072.15071.09821.0982
H72.03115.60426.76911.09103.28644.40277.61996.93826.93821.78171.78173.63093.63094.53064.5306
H85.60422.03111.09106.76914.40273.28647.61991.78171.78176.93826.93824.53064.53063.63093.6309
H94.94032.09221.09676.24984.04312.66266.93821.78171.78406.43296.67574.20324.56602.47243.0485
H104.94032.09221.09676.24984.04312.66266.93821.78171.78406.67576.43294.56604.20323.04852.4724
H112.09224.94036.24981.09672.66264.04311.78176.93826.43296.67571.78402.47243.04854.20324.5660
H122.09224.94036.24981.09672.66264.04311.78176.93826.67576.43291.78403.04852.47244.56604.2032
H132.08712.61233.95282.65221.09822.15073.63094.53064.20324.56602.47243.04851.78312.48533.0588
H142.08712.61233.95282.65221.09822.15073.63094.53064.56604.20323.04852.47241.78313.05882.4853
H152.61232.08712.65223.95282.15071.09824.53063.63092.47243.04854.20324.56602.48533.05881.7831
H162.61232.08712.65223.95282.15071.09824.53063.63093.04852.47244.56604.20323.05882.48531.7831

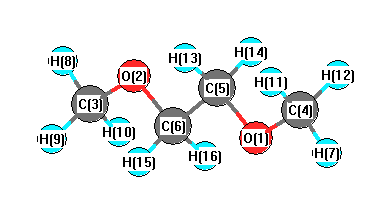
picture of Ethane, 1,2-dimethoxy- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C4 H7 106.900 O1 C4 H11 111.453
O1 C4 H12 111.453 O1 C5 C6 107.334
O1 C5 H13 110.983 O1 C5 H14 110.983
O2 C3 H8 106.900 O2 C3 H9 111.453
O2 C3 H10 111.453 O2 C6 C5 107.334
O2 C6 H15 110.983 O2 C6 H16 110.983
C3 O2 C6 112.479 C4 O1 C5 112.479
C5 C6 H15 109.484 C5 C6 H16 109.484
C6 C5 H13 109.484 C6 C5 H14 109.484
H7 C4 H11 109.063 H7 C4 H12 109.063
H8 C3 H9 109.063 H8 C3 H10 109.063
H9 C3 H10 108.845 H11 C4 H12 108.845
H13 C5 H14 108.553 H15 C6 H16 108.553
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 O -0.537      
2 O -0.537      
3 C -0.180      
4 C -0.180      
5 C -0.012      
6 C -0.012      
7 H 0.167      
8 H 0.167      
9 H 0.139      
10 H 0.139      
11 H 0.139      
12 H 0.139      
13 H 0.142      
14 H 0.142      
15 H 0.142      
16 H 0.142      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 6.663 -0.103 0.000
y -0.103 9.467 0.000
z 0.000 0.000 6.597


<r2> (average value of r2) Å2
<r2> 269.344
(<r2>)1/2 16.412